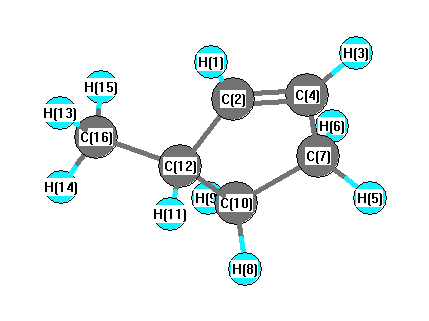
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3170 |
3089 |
42.94 |
|
|
|
| 2 |
A |
3136 |
3056 |
13.74 |
|
|
|
| 3 |
A |
3066 |
2988 |
111.94 |
|
|
|
| 4 |
A |
3059 |
2981 |
23.29 |
|
|
|
| 5 |
A |
3058 |
2980 |
47.62 |
|
|
|
| 6 |
A |
3024 |
2947 |
56.63 |
|
|
|
| 7 |
A |
2995 |
2918 |
36.84 |
|
|
|
| 8 |
A |
2974 |
2898 |
40.76 |
|
|
|
| 9 |
A |
2969 |
2893 |
73.30 |
|
|
|
| 10 |
A |
2938 |
2863 |
44.86 |
|
|
|
| 11 |
A |
1642 |
1600 |
2.22 |
|
|
|
| 12 |
A |
1524 |
1485 |
4.25 |
|
|
|
| 13 |
A |
1520 |
1481 |
6.29 |
|
|
|
| 14 |
A |
1518 |
1479 |
4.89 |
|
|
|
| 15 |
A |
1508 |
1470 |
1.88 |
|
|
|
| 16 |
A |
1427 |
1391 |
4.56 |
|
|
|
| 17 |
A |
1381 |
1346 |
5.67 |
|
|
|
| 18 |
A |
1340 |
1306 |
1.63 |
|
|
|
| 19 |
A |
1327 |
1293 |
0.22 |
|
|
|
| 20 |
A |
1313 |
1279 |
3.59 |
|
|
|
| 21 |
A |
1308 |
1274 |
0.18 |
|
|
|
| 22 |
A |
1226 |
1195 |
1.21 |
|
|
|
| 23 |
A |
1202 |
1171 |
0.33 |
|
|
|
| 24 |
A |
1144 |
1115 |
1.26 |
|
|
|
| 25 |
A |
1121 |
1092 |
3.65 |
|
|
|
| 26 |
A |
1110 |
1082 |
7.20 |
|
|
|
| 27 |
A |
1073 |
1046 |
1.53 |
|
|
|
| 28 |
A |
1002 |
976 |
1.67 |
|
|
|
| 29 |
A |
984 |
958 |
0.44 |
|
|
|
| 30 |
A |
970 |
945 |
1.65 |
|
|
|
| 31 |
A |
943 |
919 |
1.14 |
|
|
|
| 32 |
A |
901 |
878 |
4.64 |
|
|
|
| 33 |
A |
848 |
826 |
1.49 |
|
|
|
| 34 |
A |
821 |
800 |
2.48 |
|
|
|
| 35 |
A |
757 |
738 |
1.60 |
|
|
|
| 36 |
A |
747 |
728 |
44.17 |
|
|
|
| 37 |
A |
592 |
577 |
6.54 |
|
|
|
| 38 |
A |
492 |
480 |
2.53 |
|
|
|
| 39 |
A |
337 |
328 |
0.06 |
|
|
|
| 40 |
A |
282 |
275 |
0.14 |
|
|
|
| 41 |
A |
229 |
223 |
0.03 |
|
|
|
| 42 |
A |
105 |
102 |
0.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31539.1 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 30734.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.146 |
|
|
|
| 2 |
C |
-0.119 |
|
|
|
| 3 |
H |
0.148 |
|
|
|
| 4 |
C |
-0.092 |
|
|
|
| 5 |
H |
0.138 |
|
|
|
| 6 |
H |
0.135 |
|
|
|
| 7 |
C |
-0.327 |
|
|
|
| 8 |
H |
0.127 |
|
|
|
| 9 |
H |
0.139 |
|
|
|
| 10 |
C |
-0.338 |
|
|
|
| 11 |
H |
0.131 |
|
|
|
| 12 |
C |
0.009 |
|
|
|
| 13 |
H |
0.142 |
|
|
|
| 14 |
H |
0.145 |
|
|
|
| 15 |
H |
0.146 |
|
|
|
| 16 |
C |
-0.531 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.007 |
-0.202 |
0.149 |
0.251 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.975 |
-0.328 |
-0.701 |
| y |
-0.328 |
-38.174 |
-0.131 |
| z |
-0.701 |
-0.131 |
-39.650 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.937 |
-0.328 |
-0.701 |
| y |
-0.328 |
0.639 |
-0.131 |
| z |
-0.701 |
-0.131 |
-1.576 |
|
| Polar |
|---|
| 3z2-r2 | -3.151 |
|---|
| x2-y2 | 0.199 |
|---|
| xy | -0.328 |
|---|
| xz | -0.701 |
|---|
| yz | -0.131 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.067 |
-0.359 |
-0.528 |
| y |
-0.359 |
9.829 |
-0.094 |
| z |
-0.528 |
-0.094 |
7.880 |
<r2> (average value of r
2) Å
2
| <r2> |
140.270 |
| (<r2>)1/2 |
11.844 |