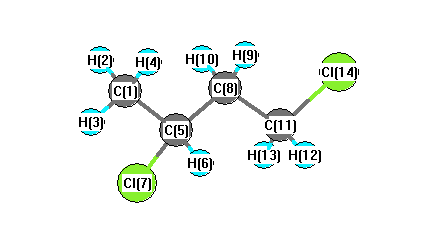
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3169 |
3089 |
10.98 |
|
|
|
| 2 |
A |
3115 |
3036 |
33.17 |
|
|
|
| 3 |
A |
3090 |
3011 |
42.15 |
|
|
|
| 4 |
A |
3079 |
3000 |
11.07 |
|
|
|
| 5 |
A |
3071 |
2993 |
7.55 |
|
|
|
| 6 |
A |
3066 |
2988 |
1.30 |
|
|
|
| 7 |
A |
3007 |
2931 |
24.47 |
|
|
|
| 8 |
A |
2992 |
2916 |
13.64 |
|
|
|
| 9 |
A |
1518 |
1479 |
6.01 |
|
|
|
| 10 |
A |
1507 |
1468 |
14.99 |
|
|
|
| 11 |
A |
1501 |
1463 |
3.56 |
|
|
|
| 12 |
A |
1493 |
1455 |
0.25 |
|
|
|
| 13 |
A |
1436 |
1399 |
9.04 |
|
|
|
| 14 |
A |
1395 |
1360 |
15.37 |
|
|
|
| 15 |
A |
1344 |
1310 |
6.11 |
|
|
|
| 16 |
A |
1324 |
1291 |
8.21 |
|
|
|
| 17 |
A |
1279 |
1247 |
11.53 |
|
|
|
| 18 |
A |
1272 |
1240 |
44.05 |
|
|
|
| 19 |
A |
1172 |
1142 |
12.29 |
|
|
|
| 20 |
A |
1123 |
1094 |
2.13 |
|
|
|
| 21 |
A |
1117 |
1088 |
10.41 |
|
|
|
| 22 |
A |
1064 |
1037 |
2.34 |
|
|
|
| 23 |
A |
1024 |
998 |
15.93 |
|
|
|
| 24 |
A |
966 |
942 |
2.49 |
|
|
|
| 25 |
A |
910 |
887 |
8.67 |
|
|
|
| 26 |
A |
790 |
770 |
7.88 |
|
|
|
| 27 |
A |
689 |
671 |
43.60 |
|
|
|
| 28 |
A |
556 |
542 |
39.07 |
|
|
|
| 29 |
A |
423 |
413 |
3.81 |
|
|
|
| 30 |
A |
386 |
376 |
10.33 |
|
|
|
| 31 |
A |
321 |
313 |
6.25 |
|
|
|
| 32 |
A |
233 |
227 |
0.79 |
|
|
|
| 33 |
A |
226 |
220 |
0.09 |
|
|
|
| 34 |
A |
144 |
141 |
3.29 |
|
|
|
| 35 |
A |
107 |
104 |
1.59 |
|
|
|
| 36 |
A |
69 |
67 |
4.95 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24989.2 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 24352.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.436 |
|
|
|
| 2 |
H |
0.172 |
|
|
|
| 3 |
H |
0.181 |
|
|
|
| 4 |
H |
0.156 |
|
|
|
| 5 |
C |
-0.282 |
|
|
|
| 6 |
H |
0.192 |
|
|
|
| 7 |
Cl |
-0.063 |
|
|
|
| 8 |
C |
-0.154 |
|
|
|
| 9 |
H |
0.164 |
|
|
|
| 10 |
H |
0.176 |
|
|
|
| 11 |
C |
-0.489 |
|
|
|
| 12 |
H |
0.196 |
|
|
|
| 13 |
H |
0.217 |
|
|
|
| 14 |
Cl |
-0.030 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.879 |
-1.901 |
1.413 |
3.023 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-60.293 |
-5.606 |
-0.405 |
| y |
-5.606 |
-52.499 |
0.388 |
| z |
-0.405 |
0.388 |
-49.429 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-9.329 |
-5.606 |
-0.405 |
| y |
-5.606 |
2.362 |
0.388 |
| z |
-0.405 |
0.388 |
6.968 |
|
| Polar |
|---|
| 3z2-r2 | 13.935 |
|---|
| x2-y2 | -7.794 |
|---|
| xy | -5.606 |
|---|
| xz | -0.405 |
|---|
| yz | 0.388 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.184 |
0.958 |
0.434 |
| y |
0.958 |
10.348 |
-0.309 |
| z |
0.434 |
-0.309 |
7.615 |
<r2> (average value of r
2) Å
2
| <r2> |
222.195 |
| (<r2>)1/2 |
14.906 |