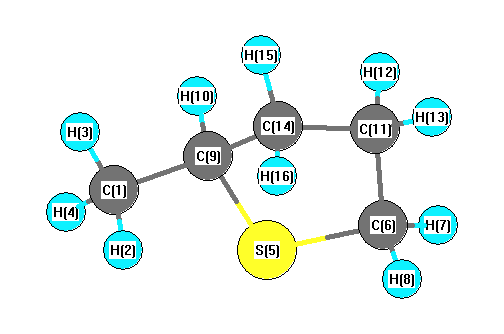
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3111 |
3031 |
33.75 |
|
|
|
| 2 |
A |
3089 |
3011 |
51.51 |
|
|
|
| 3 |
A |
3065 |
2987 |
57.94 |
|
|
|
| 4 |
A |
3057 |
2979 |
56.91 |
|
|
|
| 5 |
A |
3046 |
2968 |
29.72 |
|
|
|
| 6 |
A |
3033 |
2956 |
37.94 |
|
|
|
| 7 |
A |
3022 |
2945 |
6.83 |
|
|
|
| 8 |
A |
2997 |
2921 |
38.60 |
|
|
|
| 9 |
A |
2986 |
2910 |
63.62 |
|
|
|
| 10 |
A |
2979 |
2903 |
25.30 |
|
|
|
| 11 |
A |
1521 |
1482 |
7.61 |
|
|
|
| 12 |
A |
1515 |
1476 |
1.60 |
|
|
|
| 13 |
A |
1509 |
1471 |
9.04 |
|
|
|
| 14 |
A |
1504 |
1466 |
5.54 |
|
|
|
| 15 |
A |
1501 |
1463 |
3.16 |
|
|
|
| 16 |
A |
1432 |
1396 |
6.94 |
|
|
|
| 17 |
A |
1376 |
1341 |
2.82 |
|
|
|
| 18 |
A |
1361 |
1326 |
1.15 |
|
|
|
| 19 |
A |
1330 |
1296 |
0.89 |
|
|
|
| 20 |
A |
1299 |
1265 |
10.12 |
|
|
|
| 21 |
A |
1282 |
1250 |
26.89 |
|
|
|
| 22 |
A |
1232 |
1201 |
5.54 |
|
|
|
| 23 |
A |
1199 |
1169 |
9.50 |
|
|
|
| 24 |
A |
1154 |
1125 |
0.33 |
|
|
|
| 25 |
A |
1110 |
1081 |
0.24 |
|
|
|
| 26 |
A |
1072 |
1045 |
2.53 |
|
|
|
| 27 |
A |
1048 |
1022 |
6.20 |
|
|
|
| 28 |
A |
1029 |
1002 |
1.93 |
|
|
|
| 29 |
A |
981 |
956 |
3.37 |
|
|
|
| 30 |
A |
954 |
930 |
4.11 |
|
|
|
| 31 |
A |
890 |
867 |
1.56 |
|
|
|
| 32 |
A |
867 |
845 |
2.17 |
|
|
|
| 33 |
A |
839 |
818 |
3.12 |
|
|
|
| 34 |
A |
645 |
628 |
3.36 |
|
|
|
| 35 |
A |
587 |
572 |
5.74 |
|
|
|
| 36 |
A |
516 |
503 |
1.06 |
|
|
|
| 37 |
A |
442 |
431 |
0.29 |
|
|
|
| 38 |
A |
401 |
390 |
0.98 |
|
|
|
| 39 |
A |
278 |
271 |
1.57 |
|
|
|
| 40 |
A |
243 |
236 |
0.26 |
|
|
|
| 41 |
A |
227 |
221 |
0.11 |
|
|
|
| 42 |
A |
88 |
86 |
1.64 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30908.2 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 30120.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.453 |
|
|
|
| 2 |
H |
0.157 |
|
|
|
| 3 |
H |
0.141 |
|
|
|
| 4 |
H |
0.161 |
|
|
|
| 5 |
S |
0.207 |
|
|
|
| 6 |
C |
-0.558 |
|
|
|
| 7 |
H |
0.180 |
|
|
|
| 8 |
H |
0.177 |
|
|
|
| 9 |
C |
-0.322 |
|
|
|
| 10 |
H |
0.173 |
|
|
|
| 11 |
C |
-0.198 |
|
|
|
| 12 |
H |
0.142 |
|
|
|
| 13 |
H |
0.149 |
|
|
|
| 14 |
C |
-0.244 |
|
|
|
| 15 |
H |
0.145 |
|
|
|
| 16 |
H |
0.141 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.438 |
-2.370 |
0.061 |
2.411 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.915 |
-0.490 |
-1.039 |
| y |
-0.490 |
-49.672 |
0.546 |
| z |
-1.039 |
0.546 |
-45.846 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.844 |
-0.490 |
-1.039 |
| y |
-0.490 |
-5.791 |
0.546 |
| z |
-1.039 |
0.546 |
-0.053 |
|
| Polar |
|---|
| 3z2-r2 | -0.106 |
|---|
| x2-y2 | 7.757 |
|---|
| xy | -0.490 |
|---|
| xz | -1.039 |
|---|
| yz | 0.546 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.809 |
-0.234 |
-0.340 |
| y |
-0.234 |
11.641 |
-0.026 |
| z |
-0.340 |
-0.026 |
8.963 |
<r2> (average value of r
2) Å
2
| <r2> |
162.145 |
| (<r2>)1/2 |
12.734 |