Jump to
S1C2
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -118.956315 |
| Energy at 298.15K | |
| HF Energy | -118.956315 |
| Nuclear repulsion energy | 178.888599 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
Geometric Data calculated at B3LYP/CEP-121G
Point Group is D4h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
1.034 |
0.000 |
| C2 |
1.034 |
0.000 |
0.000 |
| C3 |
0.000 |
-1.034 |
0.000 |
| C4 |
-1.034 |
0.000 |
0.000 |
| F5 |
0.000 |
2.398 |
0.000 |
| F6 |
2.398 |
0.000 |
0.000 |
| F7 |
0.000 |
-2.398 |
0.000 |
| F8 |
-2.398 |
0.000 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
| C1 | | 1.4621 | 2.0677 | 1.4621 | 1.3637 | 2.6110 | 3.4314 | 2.6110 |
C2 | 1.4621 | | 1.4621 | 2.0677 | 2.6110 | 1.3637 | 2.6110 | 3.4314 | C3 | 2.0677 | 1.4621 | | 1.4621 | 3.4314 | 2.6110 | 1.3637 | 2.6110 | C4 | 1.4621 | 2.0677 | 1.4621 | | 2.6110 | 3.4314 | 2.6110 | 1.3637 | F5 | 1.3637 | 2.6110 | 3.4314 | 2.6110 | | 3.3907 | 4.7952 | 3.3907 | F6 | 2.6110 | 1.3637 | 2.6110 | 3.4314 | 3.3907 | | 3.3907 | 4.7952 | F7 | 3.4314 | 2.6110 | 1.3637 | 2.6110 | 4.7952 | 3.3907 | | 3.3907 | F8 | 2.6110 | 3.4314 | 2.6110 | 1.3637 | 3.3907 | 4.7952 | 3.3907 | |
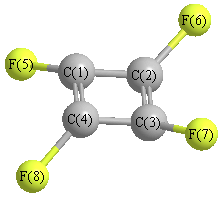 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
90.000 |
|
C1 |
C2 |
F6 |
135.000 |
| C1 |
C4 |
C3 |
90.000 |
|
C1 |
C4 |
F8 |
135.000 |
| C2 |
C1 |
C4 |
90.000 |
|
C2 |
C1 |
F5 |
135.000 |
| C2 |
C3 |
C4 |
90.000 |
|
C2 |
C3 |
F7 |
135.000 |
| C3 |
C2 |
F6 |
135.000 |
|
C3 |
C4 |
F8 |
135.000 |
| C4 |
C1 |
F5 |
135.000 |
|
C4 |
C3 |
F7 |
135.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.315 |
|
|
|
| 2 |
C |
-0.093 |
|
|
|
| 3 |
C |
0.315 |
|
|
|
| 4 |
C |
-0.093 |
|
|
|
| 5 |
F |
-0.075 |
|
|
|
| 6 |
F |
-0.148 |
|
|
|
| 7 |
F |
-0.075 |
|
|
|
| 8 |
F |
-0.148 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-52.630 |
0.000 |
0.000 |
| y |
0.000 |
-41.460 |
0.000 |
| z |
0.000 |
0.000 |
-41.327 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-11.236 |
0.000 |
0.000 |
| y |
0.000 |
5.519 |
0.000 |
| z |
0.000 |
0.000 |
5.718 |
|
| Polar |
|---|
| 3z2-r2 | 11.435 |
|---|
| x2-y2 | -11.170 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
206.249 |
| (<r2>)1/2 |
14.361 |
Jump to
S1C1
Energy calculated at B3LYP/CEP-121G
| | hartrees |
|---|
| Energy at 0K | -119.034266 |
| Energy at 298.15K | -119.033971 |
| HF Energy | -119.034266 |
| Nuclear repulsion energy | 179.392280 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/CEP-121G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
1795 |
1749 |
0.00 |
|
|
|
| 2 |
Ag |
1188 |
1158 |
0.00 |
|
|
|
| 3 |
Ag |
643 |
627 |
0.00 |
|
|
|
| 4 |
Ag |
278 |
271 |
0.00 |
|
|
|
| 5 |
Ag |
175 |
171 |
0.00 |
|
|
|
| 6 |
Au |
1231 |
1200 |
410.79 |
|
|
|
| 7 |
Au |
867 |
845 |
51.23 |
|
|
|
| 8 |
Au |
568 |
553 |
59.80 |
|
|
|
| 9 |
Au |
221 |
215 |
0.58 |
|
|
|
| 10 |
Au |
144 |
141 |
0.06 |
|
|
|
| 11 |
Bg |
1270 |
1237 |
0.00 |
|
|
|
| 12 |
Bg |
712 |
694 |
0.00 |
|
|
|
| 13 |
Bg |
472 |
460 |
0.00 |
|
|
|
| 14 |
Bg |
373 |
364 |
0.00 |
|
|
|
| 15 |
Bu |
1755 |
1710 |
148.91 |
|
|
|
| 16 |
Bu |
907 |
884 |
183.34 |
|
|
|
| 17 |
Bu |
285 |
278 |
9.19 |
|
|
|
| 18 |
Bu |
170 |
166 |
4.37 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6527.4 cm
-1
Scaled (by 0.9745) Zero Point Vibrational Energy (zpe) 6360.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/CEP-121G
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.118 |
0.784 |
0.675 |
| C2 |
-0.118 |
-0.784 |
0.675 |
| C3 |
-0.118 |
-0.784 |
-0.675 |
| C4 |
0.118 |
0.784 |
-0.675 |
| F5 |
-0.118 |
1.715 |
1.649 |
| F6 |
0.118 |
-1.715 |
1.649 |
| F7 |
0.118 |
-1.715 |
-1.649 |
| F8 |
-0.118 |
1.715 |
-1.649 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
| C1 | | 1.5848 | 2.0822 | 1.3505 | 1.3681 | 2.6819 | 3.4128 | 2.5153 |
C2 | 1.5848 | | 1.3505 | 2.0822 | 2.6819 | 1.3681 | 2.5153 | 3.4128 | C3 | 2.0822 | 1.3505 | | 1.5848 | 3.4128 | 2.5153 | 1.3681 | 2.6819 | C4 | 1.3505 | 2.0822 | 1.5848 | | 2.5153 | 3.4128 | 2.6819 | 1.3681 | F5 | 1.3681 | 2.6819 | 3.4128 | 2.5153 | | 3.4383 | 4.7648 | 3.2987 | F6 | 2.6819 | 1.3681 | 2.5153 | 3.4128 | 3.4383 | | 3.2987 | 4.7648 | F7 | 3.4128 | 2.5153 | 1.3681 | 2.6819 | 4.7648 | 3.2987 | | 3.4383 | F8 | 2.5153 | 3.4128 | 2.6819 | 1.3681 | 3.2987 | 4.7648 | 3.4383 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
90.000 |
|
C1 |
C2 |
F6 |
130.373 |
| C1 |
C4 |
C3 |
90.000 |
|
C1 |
C4 |
F8 |
135.395 |
| C2 |
C1 |
C4 |
90.000 |
|
C2 |
C1 |
F5 |
130.373 |
| C2 |
C3 |
C4 |
90.000 |
|
C2 |
C3 |
F7 |
135.395 |
| C3 |
C2 |
F6 |
135.395 |
|
C3 |
C4 |
F8 |
130.373 |
| C4 |
C1 |
F5 |
135.395 |
|
C4 |
C3 |
F7 |
130.373 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/CEP-121G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.129 |
|
|
|
| 2 |
C |
0.129 |
|
|
|
| 3 |
C |
0.129 |
|
|
|
| 4 |
C |
0.129 |
|
|
|
| 5 |
F |
-0.129 |
|
|
|
| 6 |
F |
-0.129 |
|
|
|
| 7 |
F |
-0.129 |
|
|
|
| 8 |
F |
-0.129 |
|
|
|
Electric dipole moments
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.043 |
0.154 |
0.000 |
| y |
0.154 |
-47.298 |
0.000 |
| z |
0.000 |
0.000 |
-47.281 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.247 |
0.154 |
0.000 |
| y |
0.154 |
-3.137 |
0.000 |
| z |
0.000 |
0.000 |
-3.110 |
|
| Polar |
|---|
| 3z2-r2 | -6.220 |
|---|
| x2-y2 | 6.256 |
|---|
| xy | 0.154 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.338 |
-0.076 |
0.000 |
| y |
-0.076 |
6.337 |
0.000 |
| z |
0.000 |
0.000 |
8.248 |
<r2> (average value of r
2) Å
2
| <r2> |
204.499 |
| (<r2>)1/2 |
14.300 |
