Jump to
S1C2
Energy calculated at B3LYP/SDD
| | hartrees |
|---|
| Energy at 0K | -251.861717 |
| Energy at 298.15K | -251.875939 |
| Nuclear repulsion energy | 256.637616 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/SDD
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3540 |
3403 |
1.59 |
|
|
|
| 2 |
A' |
3111 |
2991 |
134.94 |
|
|
|
| 3 |
A' |
3093 |
2973 |
6.91 |
|
|
|
| 4 |
A' |
3083 |
2964 |
80.02 |
|
|
|
| 5 |
A' |
3045 |
2927 |
20.62 |
|
|
|
| 6 |
A' |
3024 |
2907 |
40.64 |
|
|
|
| 7 |
A' |
2916 |
2803 |
188.10 |
|
|
|
| 8 |
A' |
1527 |
1468 |
5.97 |
|
|
|
| 9 |
A' |
1514 |
1455 |
9.33 |
|
|
|
| 10 |
A' |
1508 |
1450 |
11.16 |
|
|
|
| 11 |
A' |
1426 |
1371 |
0.90 |
|
|
|
| 12 |
A' |
1389 |
1336 |
1.76 |
|
|
|
| 13 |
A' |
1316 |
1265 |
0.25 |
|
|
|
| 14 |
A' |
1293 |
1243 |
2.40 |
|
|
|
| 15 |
A' |
1175 |
1130 |
2.80 |
|
|
|
| 16 |
A' |
1077 |
1036 |
1.24 |
|
|
|
| 17 |
A' |
1055 |
1014 |
12.45 |
|
|
|
| 18 |
A' |
905 |
870 |
8.55 |
|
|
|
| 19 |
A' |
873 |
839 |
4.10 |
|
|
|
| 20 |
A' |
818 |
786 |
0.34 |
|
|
|
| 21 |
A' |
677 |
651 |
85.23 |
|
|
|
| 22 |
A' |
531 |
511 |
57.69 |
|
|
|
| 23 |
A' |
428 |
411 |
16.30 |
|
|
|
| 24 |
A' |
376 |
362 |
3.66 |
|
|
|
| 25 |
A' |
232 |
223 |
1.60 |
|
|
|
| 26 |
A" |
3102 |
2982 |
11.32 |
|
|
|
| 27 |
A" |
3088 |
2968 |
116.64 |
|
|
|
| 28 |
A" |
3045 |
2927 |
53.12 |
|
|
|
| 29 |
A" |
2909 |
2796 |
49.79 |
|
|
|
| 30 |
A" |
1515 |
1456 |
0.19 |
|
|
|
| 31 |
A" |
1501 |
1443 |
1.45 |
|
|
|
| 32 |
A" |
1491 |
1434 |
7.93 |
|
|
|
| 33 |
A" |
1382 |
1329 |
0.55 |
|
|
|
| 34 |
A" |
1379 |
1326 |
0.12 |
|
|
|
| 35 |
A" |
1360 |
1307 |
29.71 |
|
|
|
| 36 |
A" |
1299 |
1249 |
2.33 |
|
|
|
| 37 |
A" |
1210 |
1163 |
10.78 |
|
|
|
| 38 |
A" |
1179 |
1134 |
5.57 |
|
|
|
| 39 |
A" |
1147 |
1103 |
11.86 |
|
|
|
| 40 |
A" |
1093 |
1051 |
3.63 |
|
|
|
| 41 |
A" |
988 |
950 |
2.36 |
|
|
|
| 42 |
A" |
888 |
853 |
1.82 |
|
|
|
| 43 |
A" |
830 |
798 |
0.26 |
|
|
|
| 44 |
A" |
448 |
431 |
0.97 |
|
|
|
| 45 |
A" |
238 |
228 |
0.77 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35010.6 cm
-1
Scaled (by 0.9613) Zero Point Vibrational Energy (zpe) 33655.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/SDD
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.647 |
1.333 |
0.000 |
| H2 |
0.590 |
2.431 |
0.000 |
| H3 |
1.717 |
1.070 |
0.000 |
| C4 |
-0.018 |
0.760 |
1.273 |
| C5 |
-0.018 |
0.760 |
-1.273 |
| C6 |
-0.018 |
-0.783 |
-1.238 |
| C7 |
-0.018 |
-0.783 |
1.238 |
| N8 |
-0.674 |
-1.252 |
0.000 |
| H9 |
-0.851 |
-2.256 |
0.000 |
| H10 |
0.507 |
1.110 |
2.174 |
| H11 |
0.507 |
1.110 |
-2.174 |
| H12 |
-1.059 |
1.109 |
1.334 |
| H13 |
-1.059 |
1.109 |
-1.334 |
| H14 |
1.032 |
-1.138 |
-1.317 |
| H15 |
1.032 |
-1.138 |
1.317 |
| H16 |
-0.568 |
-1.182 |
-2.100 |
| H17 |
-0.568 |
-1.182 |
2.100 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.0989 | 1.1021 | 1.5471 | 1.5471 | 2.5403 | 2.5403 | 2.9031 | 3.8891 | 2.1895 | 2.1895 | 2.1767 | 2.1767 | 2.8270 | 2.8270 | 3.4950 | 3.4950 |
H2 | 1.0989 | | 1.7675 | 2.1873 | 2.1873 | 3.4971 | 3.4971 | 3.8934 | 4.9029 | 2.5449 | 2.5449 | 2.4987 | 2.4987 | 3.8300 | 3.8300 | 4.3365 | 4.3365 | H3 | 1.1021 | 1.7675 | | 2.1746 | 2.1746 | 2.8239 | 2.8239 | 3.3325 | 4.2014 | 2.4879 | 2.4879 | 3.0796 | 3.0796 | 2.6603 | 2.6603 | 3.8342 | 3.8342 | C4 | 1.5471 | 2.1873 | 2.1746 | | 2.5470 | 2.9472 | 1.5430 | 2.4694 | 3.3775 | 1.0995 | 3.5045 | 1.0989 | 2.8289 | 3.3786 | 2.1699 | 3.9312 | 2.1811 | C5 | 1.5471 | 2.1873 | 2.1746 | 2.5470 | | 1.5430 | 2.9472 | 2.4694 | 3.3775 | 3.5045 | 1.0995 | 2.8289 | 1.0989 | 2.1699 | 3.3786 | 2.1811 | 3.9312 | C6 | 2.5403 | 3.4971 | 2.8239 | 2.9472 | 1.5430 | | 2.4755 | 1.4771 | 2.0963 | 3.9365 | 2.1759 | 3.3577 | 2.1610 | 1.1119 | 2.7847 | 1.0977 | 3.4061 | C7 | 2.5403 | 3.4971 | 2.8239 | 1.5430 | 2.9472 | 2.4755 | | 1.4771 | 2.0963 | 2.1759 | 3.9365 | 2.1610 | 3.3577 | 2.7847 | 1.1119 | 3.4061 | 1.0977 | N8 | 2.9031 | 3.8934 | 3.3325 | 2.4694 | 2.4694 | 1.4771 | 1.4771 | | 1.0193 | 3.4201 | 3.4201 | 2.7385 | 2.7385 | 2.1582 | 2.1582 | 2.1038 | 2.1038 | H9 | 3.8891 | 4.9029 | 4.2014 | 3.3775 | 3.3775 | 2.0963 | 2.0963 | 1.0193 | | 4.2304 | 4.2304 | 3.6250 | 3.6250 | 2.5552 | 2.5552 | 2.3752 | 2.3752 | H10 | 2.1895 | 2.5449 | 2.4879 | 1.0995 | 3.5045 | 3.9365 | 2.1759 | 3.4201 | 4.2304 | | 4.3472 | 1.7767 | 3.8411 | 4.1847 | 2.4627 | 4.9673 | 2.5330 | H11 | 2.1895 | 2.5449 | 2.4879 | 3.5045 | 1.0995 | 2.1759 | 3.9365 | 3.4201 | 4.2304 | 4.3472 | | 3.8411 | 1.7767 | 2.4627 | 4.1847 | 2.5330 | 4.9673 | H12 | 2.1767 | 2.4987 | 3.0796 | 1.0989 | 2.8289 | 3.3577 | 2.1610 | 2.7385 | 3.6250 | 1.7767 | 3.8411 | | 2.6679 | 4.0554 | 3.0695 | 4.1572 | 2.4653 | H13 | 2.1767 | 2.4987 | 3.0796 | 2.8289 | 1.0989 | 2.1610 | 3.3577 | 2.7385 | 3.6250 | 3.8411 | 1.7767 | 2.6679 | | 3.0695 | 4.0554 | 2.4653 | 4.1572 | H14 | 2.8270 | 3.8300 | 2.6603 | 3.3786 | 2.1699 | 1.1119 | 2.7847 | 2.1582 | 2.5552 | 4.1847 | 2.4627 | 4.0554 | 3.0695 | | 2.6331 | 1.7821 | 3.7729 | H15 | 2.8270 | 3.8300 | 2.6603 | 2.1699 | 3.3786 | 2.7847 | 1.1119 | 2.1582 | 2.5552 | 2.4627 | 4.1847 | 3.0695 | 4.0554 | 2.6331 | | 3.7729 | 1.7821 | H16 | 3.4950 | 4.3365 | 3.8342 | 3.9312 | 2.1811 | 1.0977 | 3.4061 | 2.1038 | 2.3752 | 4.9673 | 2.5330 | 4.1572 | 2.4653 | 1.7821 | 3.7729 | | 4.1999 | H17 | 3.4950 | 4.3365 | 3.8342 | 2.1811 | 3.9312 | 3.4061 | 1.0977 | 2.1038 | 2.3752 | 2.5330 | 4.9673 | 2.4653 | 4.1572 | 3.7729 | 1.7821 | 4.1999 | |
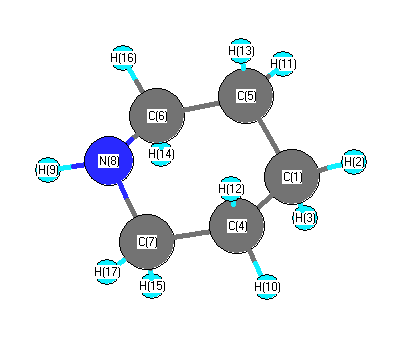 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.590 |
|
C1 |
C4 |
H10 |
110.503 |
| C1 |
C4 |
H12 |
109.539 |
|
C1 |
C5 |
C6 |
110.590 |
| C1 |
C5 |
H11 |
110.503 |
|
C1 |
C5 |
H13 |
109.539 |
| H2 |
C1 |
H3 |
106.839 |
|
H2 |
C1 |
C4 |
110.363 |
| H2 |
C1 |
C5 |
110.363 |
|
H3 |
C1 |
C4 |
109.186 |
| H3 |
C1 |
C5 |
109.186 |
|
C4 |
C1 |
C5 |
110.807 |
| C4 |
C7 |
N8 |
109.681 |
|
C4 |
C7 |
H15 |
108.544 |
| C4 |
C7 |
H17 |
110.225 |
|
C5 |
C6 |
N8 |
109.681 |
| C5 |
C6 |
H14 |
108.544 |
|
C5 |
C6 |
H16 |
110.225 |
| C6 |
C5 |
H11 |
109.715 |
|
C6 |
C5 |
H13 |
108.595 |
| C6 |
N8 |
C7 |
113.846 |
|
C6 |
N8 |
H9 |
112.943 |
| C7 |
C4 |
H10 |
109.715 |
|
C7 |
C4 |
H12 |
108.595 |
| C7 |
N8 |
H9 |
112.943 |
|
N8 |
C6 |
H14 |
112.170 |
| N8 |
C6 |
H16 |
108.687 |
|
N8 |
C7 |
H15 |
112.170 |
| N8 |
C7 |
H17 |
108.687 |
|
H10 |
C4 |
H12 |
107.834 |
| H11 |
C5 |
H13 |
107.834 |
|
H14 |
C6 |
H16 |
107.509 |
| H15 |
C7 |
H17 |
107.509 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/SDD
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.356 |
|
|
|
| 2 |
H |
0.190 |
|
|
|
| 3 |
H |
0.167 |
|
|
|
| 4 |
C |
-0.315 |
|
|
|
| 5 |
C |
-0.315 |
|
|
|
| 6 |
C |
-0.357 |
|
|
|
| 7 |
C |
-0.357 |
|
|
|
| 8 |
N |
-0.307 |
|
|
|
| 9 |
H |
0.253 |
|
|
|
| 10 |
H |
0.167 |
|
|
|
| 11 |
H |
0.167 |
|
|
|
| 12 |
H |
0.186 |
|
|
|
| 13 |
H |
0.186 |
|
|
|
| 14 |
H |
0.154 |
|
|
|
| 15 |
H |
0.154 |
|
|
|
| 16 |
H |
0.192 |
|
|
|
| 17 |
H |
0.192 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.825 |
-0.515 |
0.000 |
0.972 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.702 |
-0.031 |
0.000 |
| y |
-0.031 |
-35.845 |
0.000 |
| z |
0.000 |
0.000 |
-37.404 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.078 |
-0.031 |
0.000 |
| y |
-0.031 |
3.208 |
0.000 |
| z |
0.000 |
0.000 |
0.869 |
|
| Polar |
|---|
| 3z2-r2 | 1.739 |
|---|
| x2-y2 | -4.857 |
|---|
| xy | -0.031 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.476 |
0.400 |
0.000 |
| y |
0.400 |
8.598 |
0.000 |
| z |
0.000 |
0.000 |
9.035 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |
Jump to
S1C1
Energy calculated at B3LYP/SDD
| | hartrees |
|---|
| Energy at 0K | -251.861043 |
| Energy at 298.15K | -251.875233 |
| Nuclear repulsion energy | 256.032270 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/SDD
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3514 |
3378 |
1.84 |
|
|
|
| 2 |
A' |
3109 |
2989 |
108.82 |
|
|
|
| 3 |
A' |
3092 |
2972 |
75.81 |
|
|
|
| 4 |
A' |
3081 |
2961 |
60.20 |
|
|
|
| 5 |
A' |
3037 |
2920 |
45.91 |
|
|
|
| 6 |
A' |
3027 |
2910 |
32.62 |
|
|
|
| 7 |
A' |
3023 |
2906 |
39.78 |
|
|
|
| 8 |
A' |
1520 |
1461 |
6.97 |
|
|
|
| 9 |
A' |
1510 |
1451 |
15.30 |
|
|
|
| 10 |
A' |
1505 |
1446 |
4.67 |
|
|
|
| 11 |
A' |
1401 |
1347 |
0.49 |
|
|
|
| 12 |
A' |
1381 |
1327 |
2.54 |
|
|
|
| 13 |
A' |
1342 |
1290 |
0.04 |
|
|
|
| 14 |
A' |
1281 |
1232 |
6.03 |
|
|
|
| 15 |
A' |
1197 |
1151 |
1.71 |
|
|
|
| 16 |
A' |
1057 |
1016 |
3.99 |
|
|
|
| 17 |
A' |
1021 |
982 |
6.55 |
|
|
|
| 18 |
A' |
908 |
873 |
4.44 |
|
|
|
| 19 |
A' |
862 |
828 |
25.24 |
|
|
|
| 20 |
A' |
813 |
782 |
4.90 |
|
|
|
| 21 |
A' |
665 |
639 |
166.00 |
|
|
|
| 22 |
A' |
552 |
531 |
7.08 |
|
|
|
| 23 |
A' |
433 |
417 |
2.63 |
|
|
|
| 24 |
A' |
375 |
361 |
10.17 |
|
|
|
| 25 |
A' |
228 |
220 |
9.25 |
|
|
|
| 26 |
A" |
3105 |
2985 |
32.95 |
|
|
|
| 27 |
A" |
3080 |
2961 |
96.90 |
|
|
|
| 28 |
A" |
3034 |
2916 |
29.08 |
|
|
|
| 29 |
A" |
3023 |
2906 |
52.04 |
|
|
|
| 30 |
A" |
1507 |
1449 |
0.01 |
|
|
|
| 31 |
A" |
1502 |
1444 |
10.15 |
|
|
|
| 32 |
A" |
1486 |
1429 |
11.24 |
|
|
|
| 33 |
A" |
1389 |
1336 |
0.41 |
|
|
|
| 34 |
A" |
1376 |
1322 |
1.94 |
|
|
|
| 35 |
A" |
1370 |
1317 |
1.65 |
|
|
|
| 36 |
A" |
1305 |
1254 |
0.94 |
|
|
|
| 37 |
A" |
1224 |
1177 |
18.40 |
|
|
|
| 38 |
A" |
1135 |
1091 |
1.91 |
|
|
|
| 39 |
A" |
1130 |
1086 |
12.28 |
|
|
|
| 40 |
A" |
1092 |
1050 |
5.32 |
|
|
|
| 41 |
A" |
960 |
923 |
8.40 |
|
|
|
| 42 |
A" |
877 |
844 |
1.46 |
|
|
|
| 43 |
A" |
818 |
786 |
0.00 |
|
|
|
| 44 |
A" |
455 |
437 |
1.07 |
|
|
|
| 45 |
A" |
226 |
217 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35013.9 cm
-1
Scaled (by 0.9613) Zero Point Vibrational Energy (zpe) 33658.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/SDD
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.651 |
1.331 |
0.000 |
| H2 |
0.594 |
2.428 |
0.000 |
| H3 |
1.721 |
1.068 |
0.000 |
| C4 |
-0.011 |
0.756 |
1.277 |
| C5 |
-0.011 |
0.756 |
-1.277 |
| C6 |
-0.011 |
-0.795 |
-1.237 |
| C7 |
-0.011 |
-0.795 |
1.237 |
| N8 |
-0.605 |
-1.353 |
0.000 |
| H9 |
-1.624 |
-1.273 |
0.000 |
| H10 |
0.519 |
1.108 |
2.175 |
| H11 |
0.519 |
1.108 |
-2.175 |
| H12 |
-1.049 |
1.119 |
1.348 |
| H13 |
-1.049 |
1.119 |
-1.348 |
| H14 |
1.027 |
-1.156 |
-1.301 |
| H15 |
1.027 |
-1.156 |
1.301 |
| H16 |
-0.550 |
-1.207 |
-2.099 |
| H17 |
-0.550 |
-1.207 |
2.099 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.0990 | 1.1019 | 1.5483 | 1.5483 | 2.5468 | 2.5468 | 2.9634 | 3.4570 | 2.1899 | 2.1899 | 2.1796 | 2.1796 | 2.8314 | 2.8314 | 3.5054 | 3.5054 |
H2 | 1.0990 | | 1.7667 | 2.1887 | 2.1887 | 3.5050 | 3.5050 | 3.9671 | 4.3143 | 2.5449 | 2.5449 | 2.4959 | 2.4959 | 3.8374 | 3.8374 | 4.3507 | 4.3507 | H3 | 1.1019 | 1.7667 | | 2.1737 | 2.1737 | 2.8282 | 2.8282 | 3.3576 | 4.0823 | 2.4851 | 2.4851 | 3.0806 | 3.0806 | 2.6680 | 2.6680 | 3.8391 | 3.8391 | C4 | 1.5483 | 2.1887 | 2.1737 | | 2.5534 | 2.9537 | 1.5520 | 2.5366 | 2.8895 | 1.1002 | 3.5094 | 1.1017 | 2.8460 | 3.3732 | 2.1756 | 3.9425 | 2.1959 | C5 | 1.5483 | 2.1887 | 2.1737 | 2.5534 | | 1.5520 | 2.9537 | 2.5366 | 2.8895 | 3.5094 | 1.1002 | 2.8460 | 1.1017 | 2.1756 | 3.3732 | 2.1959 | 3.9425 | C6 | 2.5468 | 3.5050 | 2.8282 | 2.9537 | 1.5520 | | 2.4735 | 1.4815 | 2.0880 | 3.9423 | 2.1871 | 3.3798 | 2.1800 | 1.1006 | 2.7655 | 1.0975 | 3.4044 | C7 | 2.5468 | 3.5050 | 2.8282 | 1.5520 | 2.9537 | 2.4735 | | 1.4815 | 2.0880 | 2.1871 | 3.9423 | 2.1800 | 3.3798 | 2.7655 | 1.1006 | 3.4044 | 1.0975 | N8 | 2.9634 | 3.9671 | 3.3576 | 2.5366 | 2.5366 | 1.4815 | 1.4815 | | 1.0216 | 3.4718 | 3.4718 | 2.8505 | 2.8505 | 2.0970 | 2.0970 | 2.1052 | 2.1052 | H9 | 3.4570 | 4.3143 | 4.0823 | 2.8895 | 2.8895 | 2.0880 | 2.0880 | 1.0216 | | 3.8716 | 3.8716 | 2.8049 | 2.8049 | 2.9554 | 2.9554 | 2.3589 | 2.3589 | H10 | 2.1899 | 2.5449 | 2.4851 | 1.1002 | 3.5094 | 3.9423 | 2.1871 | 3.4718 | 3.8716 | | 4.3493 | 1.7718 | 3.8560 | 4.1792 | 2.4794 | 4.9770 | 2.5512 | H11 | 2.1899 | 2.5449 | 2.4851 | 3.5094 | 1.1002 | 2.1871 | 3.9423 | 3.4718 | 3.8716 | 4.3493 | | 3.8560 | 1.7718 | 2.4794 | 4.1792 | 2.5512 | 4.9770 | H12 | 2.1796 | 2.4959 | 3.0806 | 1.1017 | 2.8460 | 3.3798 | 2.1800 | 2.8505 | 2.8049 | 1.7718 | 3.8560 | | 2.6968 | 4.0622 | 3.0795 | 4.1885 | 2.4940 | H13 | 2.1796 | 2.4959 | 3.0806 | 2.8460 | 1.1017 | 2.1800 | 3.3798 | 2.8505 | 2.8049 | 3.8560 | 1.7718 | 2.6968 | | 3.0795 | 4.0622 | 2.4940 | 4.1885 | H14 | 2.8314 | 3.8374 | 2.6680 | 3.3732 | 2.1756 | 1.1006 | 2.7655 | 2.0970 | 2.9554 | 4.1792 | 2.4794 | 4.0622 | 3.0795 | | 2.6023 | 1.7685 | 3.7489 | H15 | 2.8314 | 3.8374 | 2.6680 | 2.1756 | 3.3732 | 2.7655 | 1.1006 | 2.0970 | 2.9554 | 2.4794 | 4.1792 | 3.0795 | 4.0622 | 2.6023 | | 3.7489 | 1.7685 | H16 | 3.5054 | 4.3507 | 3.8391 | 3.9425 | 2.1959 | 1.0975 | 3.4044 | 2.1052 | 2.3589 | 4.9770 | 2.5512 | 4.1885 | 2.4940 | 1.7685 | 3.7489 | | 4.1987 | H17 | 3.5054 | 4.3507 | 3.8391 | 2.1959 | 3.9425 | 3.4044 | 1.0975 | 2.1052 | 2.3589 | 2.5512 | 4.9770 | 2.4940 | 4.1885 | 3.7489 | 1.7685 | 4.1987 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.463 |
|
C1 |
C4 |
H10 |
110.412 |
| C1 |
C4 |
H12 |
109.516 |
|
C1 |
C5 |
C6 |
110.463 |
| C1 |
C5 |
H11 |
110.412 |
|
C1 |
C5 |
H13 |
109.516 |
| H2 |
C1 |
H3 |
106.778 |
|
H2 |
C1 |
C4 |
110.380 |
| H2 |
C1 |
C5 |
110.380 |
|
H3 |
C1 |
C4 |
109.046 |
| H3 |
C1 |
C5 |
109.046 |
|
C4 |
C1 |
C5 |
111.097 |
| C4 |
C7 |
N8 |
113.464 |
|
C4 |
C7 |
H15 |
109.019 |
| C4 |
C7 |
H17 |
110.777 |
|
C5 |
C6 |
N8 |
113.464 |
| C5 |
C6 |
H14 |
109.019 |
|
C5 |
C6 |
H16 |
110.777 |
| C6 |
C5 |
H11 |
109.934 |
|
C6 |
C5 |
H13 |
109.292 |
| C6 |
N8 |
C7 |
113.189 |
|
C6 |
N8 |
H9 |
111.744 |
| C7 |
C4 |
H10 |
109.934 |
|
C7 |
C4 |
H12 |
109.292 |
| C7 |
N8 |
H9 |
111.744 |
|
N8 |
C6 |
H14 |
107.700 |
| N8 |
C6 |
H16 |
108.512 |
|
N8 |
C7 |
H15 |
107.700 |
| N8 |
C7 |
H17 |
108.512 |
|
H10 |
C4 |
H12 |
107.158 |
| H11 |
C5 |
H13 |
107.158 |
|
H14 |
C6 |
H16 |
107.135 |
| H15 |
C7 |
H17 |
107.135 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/SDD
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.349 |
|
|
|
| 2 |
H |
0.191 |
|
|
|
| 3 |
H |
0.171 |
|
|
|
| 4 |
C |
-0.328 |
|
|
|
| 5 |
C |
-0.328 |
|
|
|
| 6 |
C |
-0.392 |
|
|
|
| 7 |
C |
-0.392 |
|
|
|
| 8 |
N |
-0.263 |
|
|
|
| 9 |
H |
0.239 |
|
|
|
| 10 |
H |
0.173 |
|
|
|
| 11 |
H |
0.173 |
|
|
|
| 12 |
H |
0.170 |
|
|
|
| 13 |
H |
0.170 |
|
|
|
| 14 |
H |
0.184 |
|
|
|
| 15 |
H |
0.184 |
|
|
|
| 16 |
H |
0.198 |
|
|
|
| 17 |
H |
0.198 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.323 |
1.402 |
0.000 |
1.438 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.117 |
-0.094 |
0.000 |
| y |
-0.094 |
-43.927 |
0.000 |
| z |
0.000 |
0.000 |
-37.215 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.454 |
-0.094 |
0.000 |
| y |
-0.094 |
-7.261 |
0.000 |
| z |
0.000 |
0.000 |
2.807 |
|
| Polar |
|---|
| 3z2-r2 | 5.615 |
|---|
| x2-y2 | 7.810 |
|---|
| xy | -0.094 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.808 |
0.318 |
0.000 |
| y |
0.318 |
8.104 |
0.000 |
| z |
0.000 |
0.000 |
9.015 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |
