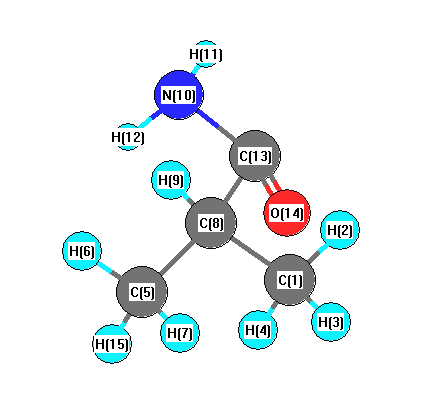
Vibrational Frequencies calculated at B3LYP/SDD
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3756 |
3610 |
26.72 |
|
|
|
| 2 |
A |
3601 |
3462 |
35.80 |
|
|
|
| 3 |
A |
3154 |
3032 |
51.13 |
|
|
|
| 4 |
A |
3148 |
3026 |
4.05 |
|
|
|
| 5 |
A |
3133 |
3012 |
59.33 |
|
|
|
| 6 |
A |
3128 |
3007 |
24.98 |
|
|
|
| 7 |
A |
3049 |
2931 |
26.75 |
|
|
|
| 8 |
A |
3042 |
2925 |
41.94 |
|
|
|
| 9 |
A |
3041 |
2923 |
34.03 |
|
|
|
| 10 |
A |
1688 |
1622 |
292.44 |
|
|
|
| 11 |
A |
1646 |
1582 |
63.67 |
|
|
|
| 12 |
A |
1537 |
1477 |
21.79 |
|
|
|
| 13 |
A |
1522 |
1463 |
17.22 |
|
|
|
| 14 |
A |
1518 |
1459 |
3.86 |
|
|
|
| 15 |
A |
1509 |
1451 |
3.55 |
|
|
|
| 16 |
A |
1444 |
1388 |
42.45 |
|
|
|
| 17 |
A |
1423 |
1368 |
12.25 |
|
|
|
| 18 |
A |
1414 |
1360 |
40.26 |
|
|
|
| 19 |
A |
1337 |
1286 |
0.28 |
|
|
|
| 20 |
A |
1300 |
1249 |
116.91 |
|
|
|
| 21 |
A |
1209 |
1163 |
8.72 |
|
|
|
| 22 |
A |
1137 |
1093 |
15.87 |
|
|
|
| 23 |
A |
1134 |
1090 |
0.78 |
|
|
|
| 24 |
A |
1075 |
1033 |
7.24 |
|
|
|
| 25 |
A |
988 |
950 |
0.18 |
|
|
|
| 26 |
A |
952 |
916 |
6.60 |
|
|
|
| 27 |
A |
924 |
888 |
3.35 |
|
|
|
| 28 |
A |
764 |
734 |
35.44 |
|
|
|
| 29 |
A |
756 |
727 |
1.99 |
|
|
|
| 30 |
A |
615 |
591 |
131.34 |
|
|
|
| 31 |
A |
609 |
585 |
4.77 |
|
|
|
| 32 |
A |
522 |
502 |
159.53 |
|
|
|
| 33 |
A |
473 |
455 |
2.38 |
|
|
|
| 34 |
A |
326 |
314 |
1.03 |
|
|
|
| 35 |
A |
285 |
274 |
7.54 |
|
|
|
| 36 |
A |
235 |
225 |
0.88 |
|
|
|
| 37 |
A |
234 |
225 |
4.97 |
|
|
|
| 38 |
A |
206 |
198 |
0.40 |
|
|
|
| 39 |
A |
30 |
29 |
9.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28930.8 cm
-1
Scaled (by 0.9613) Zero Point Vibrational Energy (zpe) 27811.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/SDD
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.639 |
|
|
|
| 2 |
H |
0.199 |
|
|
|
| 3 |
H |
0.230 |
|
|
|
| 4 |
H |
0.195 |
|
|
|
| 5 |
C |
-0.639 |
|
|
|
| 6 |
H |
0.199 |
|
|
|
| 7 |
H |
0.230 |
|
|
|
| 8 |
C |
-0.109 |
|
|
|
| 9 |
H |
0.166 |
|
|
|
| 10 |
N |
-0.628 |
|
|
|
| 11 |
H |
0.332 |
|
|
|
| 12 |
H |
0.311 |
|
|
|
| 13 |
C |
0.257 |
|
|
|
| 14 |
O |
-0.300 |
|
|
|
| 15 |
H |
0.195 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.009 |
0.038 |
-4.103 |
4.103 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.347 |
-0.051 |
5.413 |
| y |
-0.051 |
-38.146 |
0.015 |
| z |
5.413 |
0.015 |
-39.571 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.511 |
-0.051 |
5.413 |
| y |
-0.051 |
-2.187 |
0.015 |
| z |
5.413 |
0.015 |
-4.324 |
|
| Polar |
|---|
| 3z2-r2 | -8.649 |
|---|
| x2-y2 | 5.798 |
|---|
| xy | -0.051 |
|---|
| xz | 5.413 |
|---|
| yz | 0.015 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.313 |
-0.003 |
0.347 |
| y |
-0.003 |
6.489 |
-0.013 |
| z |
0.347 |
-0.013 |
7.824 |
<r2> (average value of r
2) Å
2
| <r2> |
176.307 |
| (<r2>)1/2 |
13.278 |