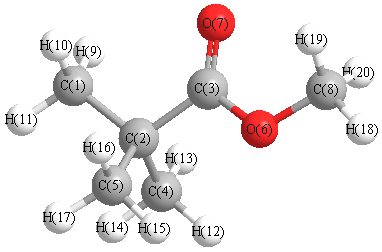
Vibrational Frequencies calculated at B3LYP/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3156 |
3045 |
15.80 |
|
|
|
| 2 |
A |
3128 |
3019 |
35.00 |
|
|
|
| 3 |
A |
3127 |
3017 |
23.29 |
|
|
|
| 4 |
A |
3124 |
3015 |
2.89 |
|
|
|
| 5 |
A |
3124 |
3014 |
15.82 |
|
|
|
| 6 |
A |
3116 |
3007 |
32.84 |
|
|
|
| 7 |
A |
3101 |
2993 |
23.06 |
|
|
|
| 8 |
A |
3099 |
2991 |
10.81 |
|
|
|
| 9 |
A |
3050 |
2943 |
29.72 |
|
|
|
| 10 |
A |
3045 |
2939 |
24.87 |
|
|
|
| 11 |
A |
3036 |
2930 |
11.17 |
|
|
|
| 12 |
A |
3034 |
2928 |
28.19 |
|
|
|
| 13 |
A |
1812 |
1749 |
192.29 |
|
|
|
| 14 |
A |
1522 |
1468 |
20.84 |
|
|
|
| 15 |
A |
1502 |
1450 |
9.89 |
|
|
|
| 16 |
A |
1498 |
1446 |
4.71 |
|
|
|
| 17 |
A |
1497 |
1445 |
4.64 |
|
|
|
| 18 |
A |
1490 |
1438 |
0.04 |
|
|
|
| 19 |
A |
1483 |
1432 |
0.14 |
|
|
|
| 20 |
A |
1480 |
1428 |
6.11 |
|
|
|
| 21 |
A |
1476 |
1424 |
0.07 |
|
|
|
| 22 |
A |
1471 |
1419 |
7.03 |
|
|
|
| 23 |
A |
1431 |
1381 |
8.85 |
|
|
|
| 24 |
A |
1401 |
1352 |
3.10 |
|
|
|
| 25 |
A |
1395 |
1346 |
3.09 |
|
|
|
| 26 |
A |
1306 |
1260 |
61.98 |
|
|
|
| 27 |
A |
1252 |
1209 |
4.08 |
|
|
|
| 28 |
A |
1229 |
1186 |
2.31 |
|
|
|
| 29 |
A |
1212 |
1170 |
100.46 |
|
|
|
| 30 |
A |
1183 |
1142 |
241.55 |
|
|
|
| 31 |
A |
1179 |
1137 |
1.23 |
|
|
|
| 32 |
A |
1059 |
1022 |
13.45 |
|
|
|
| 33 |
A |
1050 |
1013 |
0.23 |
|
|
|
| 34 |
A |
1012 |
976 |
5.24 |
|
|
|
| 35 |
A |
968 |
934 |
0.04 |
|
|
|
| 36 |
A |
947 |
914 |
2.24 |
|
|
|
| 37 |
A |
941 |
908 |
1.22 |
|
|
|
| 38 |
A |
872 |
841 |
11.74 |
|
|
|
| 39 |
A |
795 |
767 |
1.59 |
|
|
|
| 40 |
A |
778 |
751 |
4.92 |
|
|
|
| 41 |
A |
587 |
566 |
2.33 |
|
|
|
| 42 |
A |
494 |
476 |
2.39 |
|
|
|
| 43 |
A |
377 |
364 |
0.90 |
|
|
|
| 44 |
A |
361 |
349 |
10.52 |
|
|
|
| 45 |
A |
338 |
327 |
0.39 |
|
|
|
| 46 |
A |
318 |
307 |
6.70 |
|
|
|
| 47 |
A |
285 |
275 |
2.64 |
|
|
|
| 48 |
A |
254 |
245 |
0.09 |
|
|
|
| 49 |
A |
247 |
238 |
1.00 |
|
|
|
| 50 |
A |
199 |
192 |
0.15 |
|
|
|
| 51 |
A |
198 |
191 |
1.46 |
|
|
|
| 52 |
A |
130 |
125 |
1.45 |
|
|
|
| 53 |
A |
108 |
104 |
0.48 |
|
|
|
| 54 |
A |
38 |
36 |
0.68 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38157.0 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 36821.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/6-31G(2df,p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.398 |
|
|
|
| 2 |
C |
0.129 |
|
|
|
| 3 |
C |
0.454 |
|
|
|
| 4 |
C |
-0.409 |
|
|
|
| 5 |
C |
-0.409 |
|
|
|
| 6 |
O |
-0.241 |
|
|
|
| 7 |
O |
-0.370 |
|
|
|
| 8 |
C |
-0.239 |
|
|
|
| 9 |
H |
0.123 |
|
|
|
| 10 |
H |
0.123 |
|
|
|
| 11 |
H |
0.105 |
|
|
|
| 12 |
H |
0.129 |
|
|
|
| 13 |
H |
0.119 |
|
|
|
| 14 |
H |
0.114 |
|
|
|
| 15 |
H |
0.129 |
|
|
|
| 16 |
H |
0.119 |
|
|
|
| 17 |
H |
0.114 |
|
|
|
| 18 |
H |
0.131 |
|
|
|
| 19 |
H |
0.139 |
|
|
|
| 20 |
H |
0.139 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.137 |
-1.709 |
0.002 |
1.714 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.741 |
-1.703 |
0.001 |
| y |
-1.703 |
-54.295 |
0.000 |
| z |
0.001 |
0.000 |
-48.460 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
7.636 |
-1.703 |
0.001 |
| y |
-1.703 |
-8.195 |
0.000 |
| z |
0.001 |
0.000 |
0.558 |
|
| Polar |
|---|
| 3z2-r2 | 1.116 |
|---|
| x2-y2 | 10.554 |
|---|
| xy | -1.703 |
|---|
| xz | 0.001 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.061 |
-0.185 |
-0.000 |
| y |
-0.185 |
10.365 |
-0.001 |
| z |
-0.000 |
-0.001 |
9.371 |
<r2> (average value of r
2) Å
2
| <r2> |
297.382 |
| (<r2>)1/2 |
17.245 |