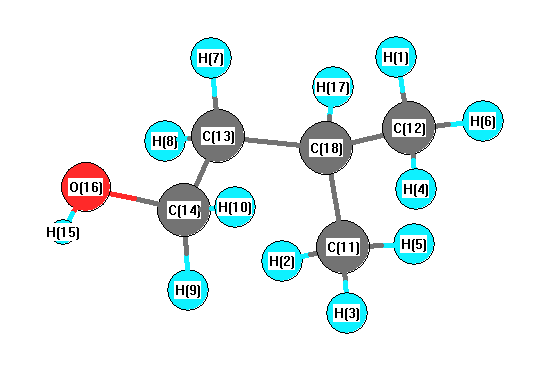
Vibrational Frequencies calculated at B3LYP/TZVP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3820 |
3687 |
23.79 |
|
|
|
| 2 |
A' |
3089 |
2982 |
58.77 |
|
|
|
| 3 |
A' |
3088 |
2981 |
79.21 |
|
|
|
| 4 |
A' |
3031 |
2926 |
13.18 |
|
|
|
| 5 |
A' |
3027 |
2922 |
46.41 |
|
|
|
| 6 |
A' |
3006 |
2902 |
11.22 |
|
|
|
| 7 |
A' |
2990 |
2887 |
40.47 |
|
|
|
| 8 |
A' |
1531 |
1478 |
1.22 |
|
|
|
| 9 |
A' |
1514 |
1462 |
8.78 |
|
|
|
| 10 |
A' |
1512 |
1460 |
12.23 |
|
|
|
| 11 |
A' |
1493 |
1441 |
2.69 |
|
|
|
| 12 |
A' |
1457 |
1407 |
1.84 |
|
|
|
| 13 |
A' |
1429 |
1380 |
4.67 |
|
|
|
| 14 |
A' |
1385 |
1337 |
0.75 |
|
|
|
| 15 |
A' |
1339 |
1293 |
21.64 |
|
|
|
| 16 |
A' |
1242 |
1199 |
28.86 |
|
|
|
| 17 |
A' |
1205 |
1164 |
4.03 |
|
|
|
| 18 |
A' |
1128 |
1089 |
5.55 |
|
|
|
| 19 |
A' |
1044 |
1008 |
93.63 |
|
|
|
| 20 |
A' |
992 |
958 |
8.11 |
|
|
|
| 21 |
A' |
954 |
921 |
13.15 |
|
|
|
| 22 |
A' |
795 |
767 |
5.48 |
|
|
|
| 23 |
A' |
559 |
539 |
1.62 |
|
|
|
| 24 |
A' |
406 |
392 |
6.72 |
|
|
|
| 25 |
A' |
327 |
315 |
5.53 |
|
|
|
| 26 |
A' |
245 |
236 |
0.10 |
|
|
|
| 27 |
A' |
207 |
200 |
2.12 |
|
|
|
| 28 |
A" |
3087 |
2980 |
26.32 |
|
|
|
| 29 |
A" |
3082 |
2975 |
0.39 |
|
|
|
| 30 |
A" |
3067 |
2961 |
47.53 |
|
|
|
| 31 |
A" |
3024 |
2920 |
59.75 |
|
|
|
| 32 |
A" |
3016 |
2912 |
6.07 |
|
|
|
| 33 |
A" |
1502 |
1450 |
1.35 |
|
|
|
| 34 |
A" |
1491 |
1439 |
0.53 |
|
|
|
| 35 |
A" |
1409 |
1360 |
8.70 |
|
|
|
| 36 |
A" |
1376 |
1328 |
1.55 |
|
|
|
| 37 |
A" |
1327 |
1282 |
0.12 |
|
|
|
| 38 |
A" |
1253 |
1210 |
0.02 |
|
|
|
| 39 |
A" |
1177 |
1136 |
5.27 |
|
|
|
| 40 |
A" |
1004 |
969 |
0.04 |
|
|
|
| 41 |
A" |
968 |
935 |
0.01 |
|
|
|
| 42 |
A" |
934 |
901 |
0.44 |
|
|
|
| 43 |
A" |
801 |
774 |
0.04 |
|
|
|
| 44 |
A" |
368 |
355 |
0.00 |
|
|
|
| 45 |
A" |
268 |
259 |
114.95 |
|
|
|
| 46 |
A" |
227 |
219 |
2.97 |
|
|
|
| 47 |
A" |
102 |
98 |
0.00 |
|
|
|
| 48 |
A" |
65 |
63 |
11.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36179.8 cm
-1
Scaled (by 0.9654) Zero Point Vibrational Energy (zpe) 34927.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/TZVP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.121 |
|
|
|
| 2 |
H |
0.121 |
|
|
|
| 3 |
H |
0.112 |
|
|
|
| 4 |
H |
0.112 |
|
|
|
| 5 |
H |
0.120 |
|
|
|
| 6 |
H |
0.120 |
|
|
|
| 7 |
H |
0.120 |
|
|
|
| 8 |
H |
0.120 |
|
|
|
| 9 |
H |
0.104 |
|
|
|
| 10 |
H |
0.104 |
|
|
|
| 11 |
C |
-0.393 |
|
|
|
| 12 |
C |
-0.393 |
|
|
|
| 13 |
C |
-0.169 |
|
|
|
| 14 |
C |
-0.164 |
|
|
|
| 15 |
H |
0.261 |
|
|
|
| 16 |
O |
-0.378 |
|
|
|
| 17 |
H |
0.129 |
|
|
|
| 18 |
C |
-0.046 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.290 |
-1.525 |
0.000 |
1.552 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.266 |
2.985 |
0.000 |
| y |
2.985 |
-44.940 |
0.000 |
| z |
0.000 |
0.000 |
-41.017 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
7.712 |
2.985 |
0.000 |
| y |
2.985 |
-6.798 |
0.000 |
| z |
0.000 |
0.000 |
-0.914 |
|
| Polar |
|---|
| 3z2-r2 | -1.828 |
|---|
| x2-y2 | 9.674 |
|---|
| xy | 2.985 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.224 |
-0.794 |
0.000 |
| y |
-0.794 |
9.472 |
0.000 |
| z |
0.000 |
0.000 |
9.388 |
<r2> (average value of r
2) Å
2
| <r2> |
225.192 |
| (<r2>)1/2 |
15.006 |