Jump to
S1C2
Energy calculated at B3LYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -187.985666 |
| Energy at 298.15K | -187.989549 |
| HF Energy | -187.985666 |
| Nuclear repulsion energy | 101.766643 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3685 |
3545 |
12.67 |
|
|
|
| 2 |
A |
3580 |
3444 |
0.00 |
|
|
|
| 3 |
A |
2372 |
2282 |
0.00 |
|
|
|
| 4 |
A |
1705 |
1640 |
0.00 |
|
|
|
| 5 |
A |
1183 |
1138 |
6.65 |
|
|
|
| 6 |
A |
847 |
815 |
0.00 |
|
|
|
| 7 |
A |
544 |
523 |
0.00 |
|
|
|
| 8 |
A |
476 |
458 |
191.32 |
|
|
|
| 9 |
A |
404 |
389 |
244.10 |
|
|
|
| 10 |
A |
180 |
173 |
27.63 |
|
|
|
| 11 |
B |
3685 |
3545 |
12.70 |
|
|
|
| 12 |
B |
3587 |
3451 |
3.40 |
|
|
|
| 13 |
B |
1713 |
1648 |
39.71 |
|
|
|
| 14 |
B |
1442 |
1387 |
120.92 |
|
|
|
| 15 |
B |
1183 |
1138 |
6.68 |
|
|
|
| 16 |
B |
476 |
458 |
191.04 |
|
|
|
| 17 |
B |
404 |
389 |
243.51 |
|
|
|
| 18 |
B |
180 |
173 |
27.63 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13824.3 cm
-1
Scaled (by 0.962) Zero Point Vibrational Energy (zpe) 13299.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/6-31G
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.023 |
0.609 |
-0.029 |
| C2 |
-0.023 |
-0.609 |
-0.029 |
| N3 |
0.023 |
1.975 |
-0.020 |
| N4 |
-0.023 |
-1.975 |
-0.020 |
| H5 |
-0.386 |
2.461 |
0.771 |
| H6 |
0.798 |
2.466 |
-0.454 |
| H7 |
0.386 |
-2.461 |
0.771 |
| H8 |
-0.798 |
-2.466 |
-0.454 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2189 | 1.3656 | 2.5840 | 2.0585 | 2.0566 | 3.1932 | 3.2111 |
C2 | 1.2189 | | 2.5840 | 1.3656 | 3.1932 | 3.2111 | 2.0585 | 2.0566 | N3 | 1.3656 | 2.5840 | | 3.9494 | 1.0147 | 1.0149 | 4.5201 | 4.5368 | N4 | 2.5840 | 1.3656 | 3.9494 | | 4.5201 | 4.5368 | 1.0147 | 1.0149 | H5 | 2.0585 | 3.1932 | 1.0147 | 4.5201 | | 1.7034 | 4.9824 | 5.0937 | H6 | 2.0566 | 3.2111 | 1.0149 | 4.5368 | 1.7034 | | 5.0937 | 5.1841 | H7 | 3.1932 | 2.0585 | 4.5201 | 1.0147 | 4.9824 | 5.0937 | | 1.7034 | H8 | 3.2111 | 2.0566 | 4.5368 | 1.0149 | 5.0937 | 5.1841 | 1.7034 | |
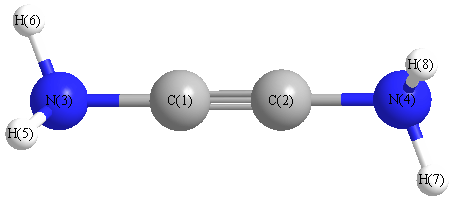 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N4 |
177.772 |
|
C1 |
N3 |
H5 |
118.985 |
| C1 |
N3 |
H6 |
118.783 |
|
C2 |
C1 |
N3 |
177.772 |
| C2 |
N4 |
H7 |
118.985 |
|
C2 |
N4 |
H8 |
118.783 |
| H5 |
N3 |
H6 |
114.130 |
|
H7 |
N4 |
H8 |
114.130 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.128 |
|
|
|
| 2 |
C |
0.128 |
|
|
|
| 3 |
N |
-0.765 |
|
|
|
| 4 |
N |
-0.765 |
|
|
|
| 5 |
H |
0.319 |
|
|
|
| 6 |
H |
0.319 |
|
|
|
| 7 |
H |
0.319 |
|
|
|
| 8 |
H |
0.319 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.007 |
0.007 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.473 |
-0.015 |
0.000 |
| y |
-0.015 |
-10.656 |
0.000 |
| z |
0.000 |
0.000 |
-23.481 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.404 |
-0.015 |
0.000 |
| y |
-0.015 |
12.821 |
0.000 |
| z |
0.000 |
0.000 |
-6.417 |
|
| Polar |
|---|
| 3z2-r2 | -12.833 |
|---|
| x2-y2 | -12.817 |
|---|
| xy | -0.015 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.230 |
-0.003 |
0.000 |
| y |
-0.003 |
9.068 |
0.000 |
| z |
0.000 |
0.000 |
2.229 |
<r2> (average value of r
2) Å
2
| <r2> |
97.559 |
| (<r2>)1/2 |
9.877 |
Jump to
S1C1
Energy calculated at B3LYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -187.985654 |
| Energy at 298.15K | -187.989522 |
| HF Energy | -187.985654 |
| Nuclear repulsion energy | 101.769240 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3581 |
3445 |
0.00 |
|
|
|
| 2 |
A1 |
2372 |
2282 |
0.00 |
|
|
|
| 3 |
A1 |
1704 |
1639 |
0.00 |
|
|
|
| 4 |
A1 |
848 |
815 |
0.00 |
|
|
|
| 5 |
B1 |
541 |
520 |
0.00 |
|
|
|
| 6 |
B2 |
3588 |
3452 |
3.41 |
|
|
|
| 7 |
B2 |
1712 |
1647 |
39.76 |
|
|
|
| 8 |
B2 |
1442 |
1388 |
120.95 |
|
|
|
| 9 |
E |
3686 |
3546 |
12.72 |
|
|
|
| 9 |
E |
3686 |
3546 |
12.72 |
|
|
|
| 10 |
E |
1182 |
1138 |
6.63 |
|
|
|
| 10 |
E |
1182 |
1138 |
6.63 |
|
|
|
| 11 |
E |
476 |
458 |
186.87 |
|
|
|
| 11 |
E |
476 |
458 |
186.87 |
|
|
|
| 12 |
E |
402 |
387 |
247.95 |
|
|
|
| 12 |
E |
402 |
387 |
247.95 |
|
|
|
| 13 |
E |
177 |
171 |
27.87 |
|
|
|
| 13 |
E |
177 |
171 |
27.87 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13817.2 cm
-1
Scaled (by 0.962) Zero Point Vibrational Energy (zpe) 13292.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/6-31G
Point Group is D2d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.611 |
| C2 |
0.000 |
0.000 |
-0.611 |
| N3 |
0.000 |
0.000 |
1.954 |
| N4 |
0.000 |
0.000 |
-1.954 |
| H5 |
0.000 |
0.859 |
2.482 |
| H6 |
0.000 |
-0.859 |
2.482 |
| H7 |
0.859 |
0.000 |
-2.482 |
| H8 |
-0.859 |
0.000 |
-2.482 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2226 | 1.3431 | 2.5657 | 2.0588 | 2.0588 | 3.2107 | 3.2107 |
C2 | 1.2226 | | 2.5657 | 1.3431 | 3.2107 | 3.2107 | 2.0588 | 2.0588 | N3 | 1.3431 | 2.5657 | | 3.9088 | 1.0080 | 1.0080 | 4.5192 | 4.5192 | N4 | 2.5657 | 1.3431 | 3.9088 | | 4.5192 | 4.5192 | 1.0080 | 1.0080 | H5 | 2.0588 | 3.2107 | 1.0080 | 4.5192 | | 1.7172 | 5.1112 | 5.1112 | H6 | 2.0588 | 3.2107 | 1.0080 | 4.5192 | 1.7172 | | 5.1112 | 5.1112 | H7 | 3.2107 | 2.0588 | 4.5192 | 1.0080 | 5.1112 | 5.1112 | | 1.7172 | H8 | 3.2107 | 2.0588 | 4.5192 | 1.0080 | 5.1112 | 5.1112 | 1.7172 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N4 |
180.000 |
|
C1 |
N3 |
H5 |
121.591 |
| C1 |
N3 |
H6 |
121.591 |
|
C2 |
C1 |
N3 |
180.000 |
| C2 |
N4 |
H7 |
121.591 |
|
C2 |
N4 |
H8 |
121.591 |
| H5 |
N3 |
H6 |
116.818 |
|
H7 |
N4 |
H8 |
116.818 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.127 |
|
|
|
| 2 |
C |
0.127 |
|
|
|
| 3 |
N |
-0.765 |
|
|
|
| 4 |
N |
-0.765 |
|
|
|
| 5 |
H |
0.319 |
|
|
|
| 6 |
H |
0.319 |
|
|
|
| 7 |
H |
0.319 |
|
|
|
| 8 |
H |
0.319 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.476 |
0.000 |
0.000 |
| y |
0.000 |
-23.476 |
0.000 |
| z |
0.000 |
0.000 |
-10.660 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.408 |
0.000 |
0.000 |
| y |
0.000 |
-6.408 |
0.000 |
| z |
0.000 |
0.000 |
12.816 |
|
| Polar |
|---|
| 3z2-r2 | 25.631 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.229 |
0.000 |
0.000 |
| y |
0.000 |
2.229 |
0.000 |
| z |
0.000 |
0.000 |
9.067 |
<r2> (average value of r
2) Å
2
| <r2> |
97.554 |
| (<r2>)1/2 |
9.877 |
