return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H13N (cyclohexanamine)

using model chemistry: ROHF/cc-pVQZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at ROHF/cc-pVQZ
 hartrees
Energy at 0K-289.353484
Energy at 298.15K 
HF Energy-289.353484
Nuclear repulsion energy331.565024
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at ROHF/cc-pVQZ
Rotational Constants (cm-1) from geometry optimized at ROHF/cc-pVQZ
See section I.F.4 to change rotational constant units
Geometric Data calculated at ROHF/cc-pVQZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.876 0.016 0.285
C2 1.180 -1.248 -0.214
C3 -0.302 -1.265 0.158
C4 -1.030 -0.014 -0.327
C5 -0.324 1.248 0.178
C6 1.156 1.272 -0.202
N7 -2.435 -0.096 0.035
H8 2.910 0.028 -0.039
H9 1.892 0.012 1.372
H10 1.278 -1.307 -1.295
H11 1.670 -2.128 0.187
H12 -0.789 -2.141 -0.252
H13 -0.401 -1.330 1.240
H14 -0.990 -0.008 -1.413
H15 -0.420 1.291 1.261
H16 -0.826 2.127 -0.214
H17 1.629 2.159 0.204
H18 1.248 1.340 -1.283
H19 -2.945 0.689 -0.310
H20 -2.550 -0.112 1.027

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 N7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20
C11.52722.52952.96922.52291.52714.31951.08431.08712.14582.15693.46992.81193.33092.80163.46452.15802.14644.90354.4894
C21.52721.52772.53362.94042.52053.80302.15672.14731.08701.08412.16192.14922.77243.34473.92663.46202.80174.55864.0925
C32.52951.52771.52602.51282.94772.43623.46762.81362.14632.15291.08271.08872.12642.78693.45223.93113.35613.32042.6726
C42.96922.53361.52601.53162.53891.45383.95043.37962.81653.46752.14122.14061.08692.14432.15383.47492.81712.04052.0384
C52.52292.94042.51281.53161.52782.50663.46322.80383.35573.92123.44702.78932.13331.08831.08572.15552.14782.72402.7433
C61.52712.52052.94772.53891.52783.84992.15682.14642.80333.46103.92793.35762.77622.15062.15861.08421.08714.14334.1425
N74.31953.80302.43621.45382.50663.84995.34724.53004.12624.58392.64132.66742.04822.73622.75504.65114.16710.99760.9985
H81.08432.15673.46763.95043.46322.15685.34721.74102.45342.49754.29273.80034.13473.79214.28912.49812.45615.89845.5653
H91.08712.14732.81363.37962.80382.14644.53001.74103.03802.45693.80242.65994.00762.64463.79142.45823.03815.16544.4570
H102.14581.08702.14632.81653.35572.80334.12622.45343.03801.73912.46033.04032.61634.02094.16973.79202.64734.77374.6342
H112.15691.08412.15293.46753.92123.46104.58392.49752.45691.73912.49802.45663.75954.14974.94994.28753.79115.43034.7526
H123.46992.16191.08272.14123.44703.92792.64134.29273.80242.46032.49801.74192.43653.76904.26784.95384.16243.55832.9761
H132.81192.14921.08872.14062.78933.35762.66743.80032.65993.04032.45661.74193.02232.62203.77454.16784.02693.59932.4802
H143.33092.77242.12641.08692.13332.77622.04824.13474.00762.61633.75952.43653.02233.02702.45383.76402.61542.35052.8983
H152.80163.34472.78692.14431.08832.15062.73623.79212.64464.02094.14973.76902.62203.02701.74302.46423.04263.03402.5613
H163.46453.92663.45222.15381.08572.15862.75504.28913.79144.16974.94994.26783.77452.45381.74302.49122.46242.56213.0860
H172.15803.46203.93113.47492.15551.08424.65112.49812.45823.79204.28754.95384.16783.76402.46422.49121.73954.83204.8274
H182.14642.80173.35612.81712.14781.08714.16712.45613.03812.64733.79114.16244.02692.61543.04262.46241.73954.35304.6766
H194.90354.55863.32042.04052.72404.14330.99765.89845.16544.77375.43033.55833.59932.35053.03402.56214.83204.35301.6079
H204.48944.09252.67262.03842.74334.14250.99855.56534.45704.63424.75262.97612.48022.89832.56133.08604.82744.67661.6079

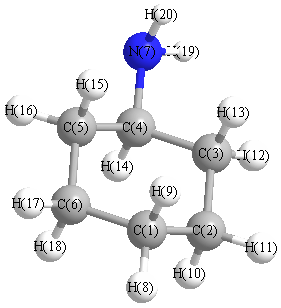
picture of cyclohexanamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 111.788 C1 C2 H10 109.172
C1 C2 H11 110.223 C1 C6 C5 111.353
C1 C6 H17 110.316 C1 C6 H18 109.223
C2 C1 C6 111.223 C2 C1 H8 110.188
C2 C1 H9 109.282 C2 C3 C4 112.129
C2 C3 H12 110.670 C2 C3 H13 109.307
C3 C2 H10 109.182 C3 C2 H11 109.872
C3 C4 C5 110.535 C3 C4 N7 109.657
C3 C4 H14 107.747 C4 C3 H12 109.138
C4 C3 H13 108.750 C4 C5 C6 112.169
C4 C5 H15 108.683 C4 C5 H16 109.575
C4 N7 H19 111.329 C4 N7 H20 111.084
C5 C4 N7 114.171 C5 C4 H14 107.903
C5 C6 H17 110.062 C5 C6 H18 109.284
C6 C1 H8 110.211 C6 C1 H9 109.223
C6 C5 H15 109.432 C6 C5 H16 110.215
N7 C4 H14 106.546 H8 C1 H9 106.594
H10 C2 H11 106.451 H12 C3 H13 106.680
H15 C5 H16 106.599 H17 C6 H18 106.472
H19 N7 H20 107.322
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at ROHF/cc-pVQZ Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.160      
2 C -0.153      
3 C -0.166      
4 C 0.086      
5 C -0.145      
6 C -0.165      
7 N -0.430      
8 H 0.098      
9 H 0.061      
10 H 0.062      
11 H 0.098      
12 H 0.113      
13 H 0.050      
14 H 0.096      
15 H 0.050      
16 H 0.080      
17 H 0.098      
18 H 0.061      
19 H 0.135      
20 H 0.133      


Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section VII.A.3)
  x y z Total
  0.580 1.044 0.433 1.270
CHELPG        
AIM        
ESP        


Electric Quadrupole moment
Quadrupole components in D Å
Primitive
 xyz
x -48.387 -3.791 -1.492
y -3.791 -46.711 -0.731
z -1.492 -0.731 -44.456
Traceless
 xyz
x -2.804 -3.791 -1.492
y -3.791 -0.290 -0.731
z -1.492 -0.731 3.093
Polar
3z2-r26.187
x2-y2-1.676
xy-3.791
xz-1.492
yz-0.731


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 0.000 0.000 0.000
y 0.000 0.000 0.000
z 0.000 0.000 0.000


<r2> (average value of r2) Å2
<r2> 231.522
(<r2>)1/2 15.216