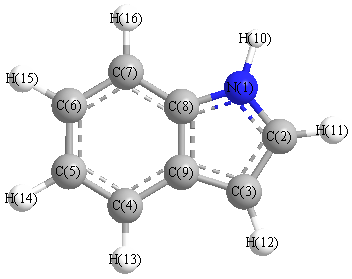
Vibrational Frequencies calculated at B97D3/6-311+G(3df,2p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3627 |
3580 |
53.54 |
|
|
|
| 2 |
A' |
3206 |
3165 |
4.37 |
|
|
|
| 3 |
A' |
3187 |
3145 |
2.34 |
|
|
|
| 4 |
A' |
3136 |
3096 |
22.23 |
|
|
|
| 5 |
A' |
3125 |
3084 |
39.39 |
|
|
|
| 6 |
A' |
3114 |
3074 |
3.95 |
|
|
|
| 7 |
A' |
3107 |
3067 |
1.94 |
|
|
|
| 8 |
A' |
1618 |
1597 |
2.86 |
|
|
|
| 9 |
A' |
1575 |
1554 |
1.09 |
|
|
|
| 10 |
A' |
1509 |
1490 |
4.66 |
|
|
|
| 11 |
A' |
1489 |
1470 |
1.47 |
|
|
|
| 12 |
A' |
1446 |
1427 |
17.33 |
|
|
|
| 13 |
A' |
1411 |
1393 |
8.13 |
|
|
|
| 14 |
A' |
1354 |
1336 |
20.70 |
|
|
|
| 15 |
A' |
1346 |
1329 |
9.97 |
|
|
|
| 16 |
A' |
1272 |
1255 |
21.81 |
|
|
|
| 17 |
A' |
1245 |
1228 |
9.61 |
|
|
|
| 18 |
A' |
1197 |
1181 |
2.65 |
|
|
|
| 19 |
A' |
1153 |
1138 |
1.35 |
|
|
|
| 20 |
A' |
1118 |
1104 |
0.18 |
|
|
|
| 21 |
A' |
1086 |
1071 |
20.88 |
|
|
|
| 22 |
A' |
1064 |
1050 |
8.13 |
|
|
|
| 23 |
A' |
1012 |
999 |
6.91 |
|
|
|
| 24 |
A' |
899 |
887 |
6.25 |
|
|
|
| 25 |
A' |
866 |
855 |
0.34 |
|
|
|
| 26 |
A' |
768 |
758 |
1.60 |
|
|
|
| 27 |
A' |
610 |
602 |
1.13 |
|
|
|
| 28 |
A' |
545 |
538 |
0.05 |
|
|
|
| 29 |
A' |
398 |
392 |
3.98 |
|
|
|
| 30 |
A" |
950 |
938 |
0.03 |
|
|
|
| 31 |
A" |
912 |
900 |
1.87 |
|
|
|
| 32 |
A" |
830 |
819 |
4.45 |
|
|
|
| 33 |
A" |
824 |
813 |
0.10 |
|
|
|
| 34 |
A" |
737 |
728 |
45.25 |
|
|
|
| 35 |
A" |
724 |
715 |
48.69 |
|
|
|
| 36 |
A" |
693 |
684 |
37.90 |
|
|
|
| 37 |
A" |
599 |
591 |
6.58 |
|
|
|
| 38 |
A" |
564 |
557 |
0.64 |
|
|
|
| 39 |
A" |
414 |
409 |
4.32 |
|
|
|
| 40 |
A" |
369 |
364 |
58.31 |
|
|
|
| 41 |
A" |
233 |
230 |
0.03 |
|
|
|
| 42 |
A" |
201 |
198 |
11.70 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27765.7 cm
-1
Scaled (by 0.987) Zero Point Vibrational Energy (zpe) 27404.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/6-311+G(3df,2p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.791 |
|
|
|
| 2 |
C |
0.136 |
|
|
|
| 3 |
C |
-0.336 |
|
|
|
| 4 |
C |
-0.367 |
|
|
|
| 5 |
C |
-0.342 |
|
|
|
| 6 |
C |
-0.110 |
|
|
|
| 7 |
C |
-0.419 |
|
|
|
| 8 |
C |
-0.065 |
|
|
|
| 9 |
C |
1.484 |
|
|
|
| 10 |
H |
0.176 |
|
|
|
| 11 |
H |
0.123 |
|
|
|
| 12 |
H |
0.120 |
|
|
|
| 13 |
H |
0.096 |
|
|
|
| 14 |
H |
0.095 |
|
|
|
| 15 |
H |
0.095 |
|
|
|
| 16 |
H |
0.104 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.909 |
1.942 |
0.000 |
2.144 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.778 |
-2.050 |
0.000 |
| y |
-2.050 |
-43.019 |
0.000 |
| z |
0.000 |
0.000 |
-57.929 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.696 |
-2.050 |
0.000 |
| y |
-2.050 |
9.835 |
0.000 |
| z |
0.000 |
0.000 |
-12.531 |
|
| Polar |
|---|
| 3z2-r2 | -25.061 |
|---|
| x2-y2 | -4.759 |
|---|
| xy | -2.050 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
20.993 |
-1.649 |
0.000 |
| y |
-1.649 |
16.671 |
0.000 |
| z |
0.000 |
0.000 |
8.984 |
<r2> (average value of r
2) Å
2
| <r2> |
283.479 |
| (<r2>)1/2 |
16.837 |