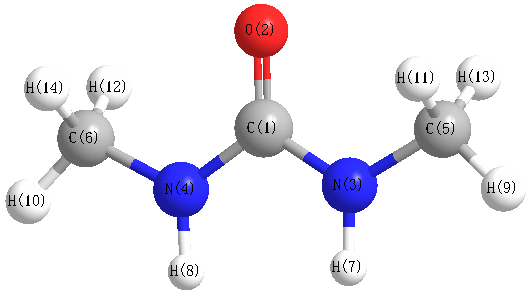
Vibrational Frequencies calculated at B97D3/6-311+G(3df,2p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3549 |
3503 |
10.29 |
|
|
|
| 2 |
A |
3085 |
3045 |
15.13 |
|
|
|
| 3 |
A |
3037 |
2998 |
7.98 |
|
|
|
| 4 |
A |
2956 |
2917 |
7.73 |
|
|
|
| 5 |
A |
1714 |
1691 |
249.47 |
|
|
|
| 6 |
A |
1497 |
1478 |
15.65 |
|
|
|
| 7 |
A |
1464 |
1445 |
0.80 |
|
|
|
| 8 |
A |
1442 |
1423 |
3.99 |
|
|
|
| 9 |
A |
1420 |
1401 |
10.74 |
|
|
|
| 10 |
A |
1170 |
1155 |
0.09 |
|
|
|
| 11 |
A |
1147 |
1132 |
1.47 |
|
|
|
| 12 |
A |
1119 |
1104 |
5.80 |
|
|
|
| 13 |
A |
899 |
887 |
1.93 |
|
|
|
| 14 |
A |
509 |
502 |
38.79 |
|
|
|
| 15 |
A |
397 |
392 |
46.84 |
|
|
|
| 16 |
A |
216 |
213 |
4.42 |
|
|
|
| 17 |
A |
171 |
169 |
0.14 |
|
|
|
| 18 |
A |
85 |
84 |
0.14 |
|
|
|
| 19 |
B |
3544 |
3498 |
5.18 |
|
|
|
| 20 |
B |
3085 |
3045 |
4.88 |
|
|
|
| 21 |
B |
3037 |
2998 |
57.57 |
|
|
|
| 22 |
B |
2954 |
2916 |
140.72 |
|
|
|
| 23 |
B |
1518 |
1498 |
275.75 |
|
|
|
| 24 |
B |
1480 |
1461 |
7.06 |
|
|
|
| 25 |
B |
1453 |
1434 |
81.46 |
|
|
|
| 26 |
B |
1421 |
1403 |
17.34 |
|
|
|
| 27 |
B |
1226 |
1210 |
242.30 |
|
|
|
| 28 |
B |
1130 |
1115 |
41.66 |
|
|
|
| 29 |
B |
1116 |
1102 |
0.09 |
|
|
|
| 30 |
B |
1011 |
998 |
1.65 |
|
|
|
| 31 |
B |
744 |
734 |
7.93 |
|
|
|
| 32 |
B |
710 |
700 |
20.99 |
|
|
|
| 33 |
B |
468 |
462 |
104.54 |
|
|
|
| 34 |
B |
312 |
308 |
40.53 |
|
|
|
| 35 |
B |
113 |
112 |
9.75 |
|
|
|
| 36 |
B |
97 |
96 |
0.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25647.0 cm
-1
Scaled (by 0.987) Zero Point Vibrational Energy (zpe) 25313.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/6-311+G(3df,2p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
1.210 |
|
|
|
| 2 |
O |
-0.749 |
|
|
|
| 3 |
N |
-0.794 |
|
|
|
| 4 |
N |
-0.794 |
|
|
|
| 5 |
C |
0.053 |
|
|
|
| 6 |
C |
0.053 |
|
|
|
| 7 |
H |
0.177 |
|
|
|
| 8 |
H |
0.177 |
|
|
|
| 9 |
H |
0.107 |
|
|
|
| 10 |
H |
0.107 |
|
|
|
| 11 |
H |
0.123 |
|
|
|
| 12 |
H |
0.123 |
|
|
|
| 13 |
H |
0.104 |
|
|
|
| 14 |
H |
0.104 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.647 |
3.647 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.015 |
-1.722 |
0.000 |
| y |
-1.722 |
-30.717 |
0.000 |
| z |
0.000 |
0.000 |
-37.824 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.744 |
-1.722 |
0.000 |
| y |
-1.722 |
7.203 |
0.000 |
| z |
0.000 |
0.000 |
-3.458 |
|
| Polar |
|---|
| 3z2-r2 | -6.916 |
|---|
| x2-y2 | -7.298 |
|---|
| xy | -1.722 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.996 |
-0.520 |
0.000 |
| y |
-0.520 |
11.860 |
0.000 |
| z |
0.000 |
0.000 |
8.977 |
<r2> (average value of r
2) Å
2
| <r2> |
194.071 |
| (<r2>)1/2 |
13.931 |