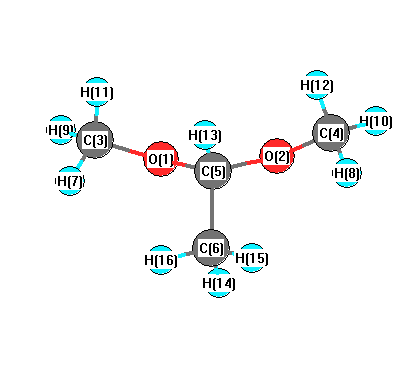
Vibrational Frequencies calculated at B97D3/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3108 |
3056 |
17.18 |
|
|
|
| 2 |
A' |
3083 |
3030 |
32.54 |
|
|
|
| 3 |
A' |
3012 |
2961 |
19.17 |
|
|
|
| 4 |
A' |
2973 |
2922 |
126.36 |
|
|
|
| 5 |
A' |
2929 |
2879 |
64.64 |
|
|
|
| 6 |
A' |
2845 |
2796 |
69.54 |
|
|
|
| 7 |
A' |
1549 |
1523 |
2.83 |
|
|
|
| 8 |
A' |
1542 |
1515 |
3.72 |
|
|
|
| 9 |
A' |
1518 |
1493 |
14.99 |
|
|
|
| 10 |
A' |
1476 |
1451 |
1.18 |
|
|
|
| 11 |
A' |
1421 |
1397 |
21.36 |
|
|
|
| 12 |
A' |
1367 |
1344 |
27.30 |
|
|
|
| 13 |
A' |
1192 |
1172 |
2.95 |
|
|
|
| 14 |
A' |
1140 |
1120 |
3.13 |
|
|
|
| 15 |
A' |
1117 |
1098 |
22.78 |
|
|
|
| 16 |
A' |
1064 |
1046 |
30.57 |
|
|
|
| 17 |
A' |
978 |
962 |
33.15 |
|
|
|
| 18 |
A' |
811 |
797 |
8.41 |
|
|
|
| 19 |
A' |
500 |
491 |
8.02 |
|
|
|
| 20 |
A' |
410 |
403 |
11.92 |
|
|
|
| 21 |
A' |
234 |
230 |
3.35 |
|
|
|
| 22 |
A' |
187 |
184 |
1.32 |
|
|
|
| 23 |
A' |
67 |
66 |
5.72 |
|
|
|
| 24 |
A" |
3083 |
3030 |
18.26 |
|
|
|
| 25 |
A" |
3073 |
3021 |
42.45 |
|
|
|
| 26 |
A" |
2972 |
2921 |
19.13 |
|
|
|
| 27 |
A" |
2927 |
2877 |
59.66 |
|
|
|
| 28 |
A" |
1549 |
1523 |
18.83 |
|
|
|
| 29 |
A" |
1522 |
1496 |
0.16 |
|
|
|
| 30 |
A" |
1519 |
1494 |
0.30 |
|
|
|
| 31 |
A" |
1480 |
1455 |
0.04 |
|
|
|
| 32 |
A" |
1351 |
1328 |
69.12 |
|
|
|
| 33 |
A" |
1156 |
1136 |
17.50 |
|
|
|
| 34 |
A" |
1136 |
1116 |
9.62 |
|
|
|
| 35 |
A" |
1119 |
1100 |
155.19 |
|
|
|
| 36 |
A" |
1002 |
985 |
201.32 |
|
|
|
| 37 |
A" |
860 |
845 |
8.25 |
|
|
|
| 38 |
A" |
447 |
440 |
3.48 |
|
|
|
| 39 |
A" |
326 |
321 |
2.14 |
|
|
|
| 40 |
A" |
238 |
234 |
0.11 |
|
|
|
| 41 |
A" |
209 |
205 |
0.03 |
|
|
|
| 42 |
A" |
61 |
60 |
0.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30276.3 cm
-1
Scaled (by 0.983) Zero Point Vibrational Energy (zpe) 29761.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.462 |
|
|
|
| 2 |
O |
-0.462 |
|
|
|
| 3 |
C |
-0.354 |
|
|
|
| 4 |
C |
-0.354 |
|
|
|
| 5 |
C |
0.285 |
|
|
|
| 6 |
C |
-0.606 |
|
|
|
| 7 |
H |
0.185 |
|
|
|
| 8 |
H |
0.185 |
|
|
|
| 9 |
H |
0.221 |
|
|
|
| 10 |
H |
0.221 |
|
|
|
| 11 |
H |
0.182 |
|
|
|
| 12 |
H |
0.182 |
|
|
|
| 13 |
H |
0.161 |
|
|
|
| 14 |
H |
0.229 |
|
|
|
| 15 |
H |
0.194 |
|
|
|
| 16 |
H |
0.194 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.912 |
2.601 |
0.000 |
2.756 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.448 |
-1.369 |
0.000 |
| y |
-1.369 |
-39.664 |
0.000 |
| z |
0.000 |
0.000 |
-30.685 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.274 |
-1.369 |
0.000 |
| y |
-1.369 |
-5.098 |
0.000 |
| z |
0.000 |
0.000 |
8.371 |
|
| Polar |
|---|
| 3z2-r2 | 16.743 |
|---|
| x2-y2 | 1.216 |
|---|
| xy | -1.369 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.700 |
-0.051 |
0.000 |
| y |
-0.051 |
6.203 |
0.000 |
| z |
0.000 |
0.000 |
9.501 |
<r2> (average value of r
2) Å
2
| <r2> |
206.151 |
| (<r2>)1/2 |
14.358 |