Jump to
S1C2
Energy calculated at B97D3/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -377.354639 |
| Energy at 298.15K | |
| HF Energy | -377.354639 |
| Nuclear repulsion energy | 188.798762 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B97D3/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3054 |
3028 |
38.05 |
49.06 |
0.73 |
0.85 |
| 2 |
A |
3005 |
2979 |
41.29 |
114.21 |
0.31 |
0.48 |
| 3 |
A |
2989 |
2964 |
23.48 |
126.95 |
0.08 |
0.15 |
| 4 |
A |
1438 |
1425 |
1.98 |
9.47 |
0.72 |
0.84 |
| 5 |
A |
1405 |
1393 |
14.32 |
3.82 |
0.55 |
0.71 |
| 6 |
A |
1357 |
1346 |
23.51 |
5.87 |
0.74 |
0.85 |
| 7 |
A |
1294 |
1283 |
22.58 |
3.06 |
0.74 |
0.85 |
| 8 |
A |
1223 |
1213 |
9.02 |
8.56 |
0.73 |
0.85 |
| 9 |
A |
1115 |
1106 |
91.01 |
4.98 |
0.70 |
0.82 |
| 10 |
A |
1097 |
1088 |
6.04 |
1.88 |
0.40 |
0.57 |
| 11 |
A |
1067 |
1058 |
260.07 |
0.07 |
0.30 |
0.47 |
| 12 |
A |
1049 |
1040 |
13.06 |
5.97 |
0.58 |
0.73 |
| 13 |
A |
878 |
871 |
38.60 |
4.67 |
0.37 |
0.54 |
| 14 |
A |
557 |
552 |
3.20 |
2.11 |
0.44 |
0.61 |
| 15 |
A |
460 |
456 |
16.85 |
0.88 |
0.74 |
0.85 |
| 16 |
A |
412 |
408 |
4.07 |
1.86 |
0.55 |
0.71 |
| 17 |
A |
235 |
233 |
7.11 |
0.08 |
0.60 |
0.75 |
| 18 |
A |
113 |
112 |
6.74 |
0.02 |
0.73 |
0.84 |
Unscaled Zero Point Vibrational Energy (zpe) 11373.8 cm
-1
Scaled (by 0.9915) Zero Point Vibrational Energy (zpe) 11277.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B97D3/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.782 |
-0.586 |
-0.290 |
| C2 |
0.470 |
0.022 |
0.329 |
| F3 |
-1.902 |
0.107 |
0.155 |
| F4 |
1.543 |
-0.777 |
-0.005 |
| F5 |
0.696 |
1.275 |
-0.182 |
| H6 |
-0.714 |
-0.517 |
-1.392 |
| H7 |
-0.860 |
-1.646 |
0.016 |
| H8 |
0.412 |
0.103 |
1.432 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5230 | 1.3902 | 2.3499 | 2.3785 | 1.1068 | 1.1067 | 2.2054 |
C2 | 1.5230 | | 2.3806 | 1.3782 | 1.3717 | 2.1575 | 2.1566 | 1.1081 | F3 | 1.3902 | 2.3806 | | 3.5603 | 2.8684 | 2.0479 | 2.0446 | 2.6433 | F4 | 2.3499 | 1.3782 | 3.5603 | | 2.2263 | 2.6616 | 2.5557 | 2.0298 | F5 | 2.3785 | 1.3717 | 2.8684 | 2.2263 | | 2.5814 | 3.3158 | 2.0149 | H6 | 1.1068 | 2.1575 | 2.0479 | 2.6616 | 2.5814 | | 1.8110 | 3.1031 | H7 | 1.1067 | 2.1566 | 2.0446 | 2.5557 | 3.3158 | 1.8110 | | 2.5852 | H8 | 2.2054 | 1.1081 | 2.6433 | 2.0298 | 2.0149 | 3.1031 | 2.5852 | |
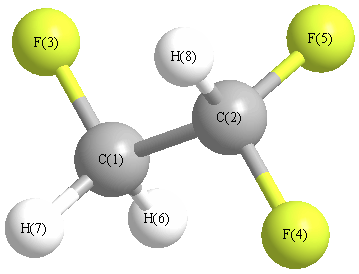 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.084 |
|
C1 |
C2 |
F5 |
110.394 |
| C1 |
C2 |
H8 |
112.952 |
|
C2 |
C1 |
F3 |
109.521 |
| C2 |
C1 |
H6 |
109.225 |
|
C2 |
C1 |
H7 |
109.163 |
| F3 |
C1 |
H6 |
109.684 |
|
F3 |
C1 |
H7 |
109.425 |
| F4 |
C2 |
F5 |
108.111 |
|
F4 |
C2 |
H8 |
108.971 |
| F5 |
C2 |
H8 |
108.216 |
|
H6 |
C1 |
H7 |
109.808 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.212 |
|
|
|
| 2 |
C |
0.335 |
|
|
|
| 3 |
F |
-0.230 |
|
|
|
| 4 |
F |
-0.203 |
|
|
|
| 5 |
F |
-0.194 |
|
|
|
| 6 |
H |
0.034 |
|
|
|
| 7 |
H |
0.027 |
|
|
|
| 8 |
H |
0.018 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.234 |
-1.279 |
0.309 |
1.336 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.426 |
1.313 |
1.181 |
| y |
1.313 |
-27.278 |
0.468 |
| z |
1.181 |
0.468 |
-24.602 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.486 |
1.313 |
1.181 |
| y |
1.313 |
0.736 |
0.468 |
| z |
1.181 |
0.468 |
4.750 |
|
| Polar |
|---|
| 3z2-r2 | 9.500 |
|---|
| x2-y2 | -4.148 |
|---|
| xy | 1.313 |
|---|
| xz | 1.181 |
|---|
| yz | 0.468 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.639 |
-0.002 |
0.040 |
| y |
-0.002 |
3.634 |
0.065 |
| z |
0.040 |
0.065 |
3.543 |
<r2> (average value of r
2) Å
2
| <r2> |
112.973 |
| (<r2>)1/2 |
10.629 |
Jump to
S1C1
Energy calculated at B97D3/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -377.352158 |
| Energy at 298.15K | |
| HF Energy | -377.352158 |
| Nuclear repulsion energy | 191.390783 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B97D3/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
2981 |
2956 |
64.87 |
153.36 |
0.02 |
0.04 |
| 2 |
A' |
2961 |
2936 |
28.62 |
65.43 |
0.67 |
0.80 |
| 3 |
A' |
1433 |
1421 |
4.65 |
8.55 |
0.72 |
0.84 |
| 4 |
A' |
1389 |
1377 |
16.97 |
3.14 |
0.44 |
0.61 |
| 5 |
A' |
1366 |
1354 |
37.70 |
4.85 |
0.60 |
0.75 |
| 6 |
A' |
1142 |
1132 |
124.51 |
3.85 |
0.44 |
0.61 |
| 7 |
A' |
1063 |
1054 |
25.11 |
3.07 |
0.72 |
0.84 |
| 8 |
A' |
843 |
836 |
34.90 |
6.29 |
0.20 |
0.33 |
| 9 |
A' |
736 |
729 |
46.85 |
3.17 |
0.55 |
0.71 |
| 10 |
A' |
499 |
495 |
9.69 |
1.77 |
0.75 |
0.86 |
| 11 |
A' |
225 |
224 |
1.71 |
0.18 |
0.43 |
0.60 |
| 12 |
A" |
3034 |
3008 |
33.57 |
66.93 |
0.75 |
0.86 |
| 13 |
A" |
1348 |
1336 |
31.20 |
2.15 |
0.75 |
0.86 |
| 14 |
A" |
1243 |
1233 |
11.84 |
15.58 |
0.75 |
0.86 |
| 15 |
A" |
1094 |
1085 |
100.59 |
1.09 |
0.75 |
0.86 |
| 16 |
A" |
921 |
913 |
76.67 |
3.20 |
0.75 |
0.86 |
| 17 |
A" |
358 |
355 |
0.13 |
0.31 |
0.75 |
0.86 |
| 18 |
A" |
109 |
108 |
1.62 |
0.03 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 11371.8 cm
-1
Scaled (by 0.9915) Zero Point Vibrational Energy (zpe) 11275.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B97D3/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.737 |
-0.841 |
0.000 |
| C2 |
0.358 |
0.633 |
0.000 |
| F3 |
-0.390 |
-1.647 |
0.000 |
| F4 |
-0.390 |
0.940 |
1.110 |
| F5 |
-0.390 |
0.940 |
-1.110 |
| H6 |
1.342 |
-1.055 |
-0.904 |
| H7 |
1.342 |
-1.055 |
0.904 |
| H8 |
1.265 |
1.275 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5222 | 1.3854 | 2.3822 | 2.3822 | 1.1080 | 1.1080 | 2.1808 |
C2 | 1.5222 | | 2.3993 | 1.3731 | 1.3731 | 2.1524 | 2.1524 | 1.1112 | F3 | 1.3854 | 2.3993 | | 2.8151 | 2.8151 | 2.0410 | 2.0410 | 3.3577 | F4 | 2.3822 | 1.3731 | 2.8151 | | 2.2209 | 3.3214 | 2.6490 | 2.0203 | F5 | 2.3822 | 1.3731 | 2.8151 | 2.2209 | | 2.6490 | 3.3214 | 2.0203 | H6 | 1.1080 | 2.1524 | 2.0410 | 3.3214 | 2.6490 | | 1.8076 | 2.4998 | H7 | 1.1080 | 2.1524 | 2.0410 | 2.6490 | 3.3214 | 1.8076 | | 2.4998 | H8 | 2.1808 | 1.1112 | 3.3577 | 2.0203 | 2.0203 | 2.4998 | 2.4998 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.619 |
|
C1 |
C2 |
F5 |
110.619 |
| C1 |
C2 |
H8 |
110.849 |
|
C2 |
C1 |
F3 |
111.123 |
| C2 |
C1 |
H6 |
108.813 |
|
C2 |
C1 |
H7 |
108.813 |
| F3 |
C1 |
H6 |
109.378 |
|
F3 |
C1 |
H7 |
109.378 |
| F4 |
C2 |
F5 |
107.935 |
|
F4 |
C2 |
H8 |
108.362 |
| F5 |
C2 |
H8 |
108.362 |
|
H6 |
C1 |
H7 |
109.310 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.221 |
|
|
|
| 2 |
C |
0.340 |
|
|
|
| 3 |
F |
-0.221 |
|
|
|
| 4 |
F |
-0.196 |
|
|
|
| 5 |
F |
-0.196 |
|
|
|
| 6 |
H |
0.025 |
|
|
|
| 7 |
H |
0.025 |
|
|
|
| 8 |
H |
0.003 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.784 |
0.042 |
0.000 |
2.785 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.863 |
-0.261 |
0.000 |
| y |
-0.261 |
-28.409 |
0.000 |
| z |
0.000 |
0.000 |
-27.917 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.300 |
-0.261 |
0.000 |
| y |
-0.261 |
-2.019 |
0.000 |
| z |
0.000 |
0.000 |
-1.281 |
|
| Polar |
|---|
| 3z2-r2 | -2.563 |
|---|
| x2-y2 | 3.546 |
|---|
| xy | -0.261 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.617 |
0.000 |
0.000 |
| y |
0.000 |
3.656 |
0.000 |
| z |
0.000 |
0.000 |
3.589 |
<r2> (average value of r
2) Å
2
| <r2> |
104.869 |
| (<r2>)1/2 |
10.241 |
