Jump to
S1C2
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -5224.211044 |
| Energy at 298.15K | -5224.204851 |
| HF Energy | -5218.018163 |
| Nuclear repulsion energy | 418.586137 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3290 |
3290 |
0.00 |
|
|
|
| 2 |
Ag |
1627 |
1627 |
0.00 |
|
|
|
| 3 |
Ag |
1434 |
1434 |
0.00 |
|
|
|
| 4 |
Ag |
1130 |
1130 |
0.00 |
|
|
|
| 5 |
Ag |
740 |
740 |
0.00 |
|
|
|
| 6 |
Ag |
207 |
207 |
0.00 |
|
|
|
| 7 |
Au |
3377 |
3377 |
2.62 |
|
|
|
| 8 |
Au |
1216 |
1216 |
2.96 |
|
|
|
| 9 |
Au |
811 |
811 |
3.41 |
|
|
|
| 10 |
Au |
112 |
112 |
5.83 |
|
|
|
| 11 |
Bg |
3356 |
3356 |
0.00 |
|
|
|
| 12 |
Bg |
1407 |
1407 |
0.00 |
|
|
|
| 13 |
Bg |
1036 |
1036 |
0.00 |
|
|
|
| 14 |
Bu |
3295 |
3295 |
15.21 |
|
|
|
| 15 |
Bu |
1618 |
1618 |
9.08 |
|
|
|
| 16 |
Bu |
1341 |
1341 |
91.59 |
|
|
|
| 17 |
Bu |
654 |
654 |
98.68 |
|
|
|
| 18 |
Bu |
199 |
199 |
7.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13425.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13425.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.492 |
0.575 |
0.000 |
| C2 |
-0.492 |
-0.575 |
0.000 |
| Br3 |
-0.492 |
2.255 |
0.000 |
| Br4 |
0.492 |
-2.255 |
0.000 |
| H5 |
1.110 |
0.581 |
0.885 |
| H6 |
1.110 |
0.581 |
-0.885 |
| H7 |
-1.110 |
-0.581 |
0.885 |
| H8 |
-1.110 |
-0.581 |
-0.885 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5134 | 1.9475 | 2.8295 | 1.0796 | 1.0796 | 2.1649 | 2.1649 |
C2 | 1.5134 | | 2.8295 | 1.9475 | 2.1649 | 2.1649 | 1.0796 | 1.0796 | Br3 | 1.9475 | 2.8295 | | 4.6160 | 2.4810 | 2.4810 | 3.0340 | 3.0340 | Br4 | 2.8295 | 1.9475 | 4.6160 | | 3.0340 | 3.0340 | 2.4810 | 2.4810 | H5 | 1.0796 | 2.1649 | 2.4810 | 3.0340 | | 1.7703 | 2.5062 | 3.0683 | H6 | 1.0796 | 2.1649 | 2.4810 | 3.0340 | 1.7703 | | 3.0683 | 2.5062 | H7 | 2.1649 | 1.0796 | 3.0340 | 2.4810 | 2.5062 | 3.0683 | | 1.7703 | H8 | 2.1649 | 1.0796 | 3.0340 | 2.4810 | 3.0683 | 2.5062 | 1.7703 | |
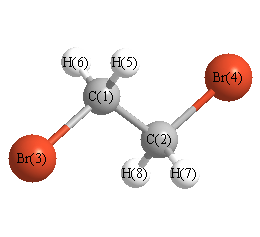 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
109.040 |
|
C1 |
C2 |
H7 |
112.129 |
| C1 |
C2 |
H8 |
112.129 |
|
C2 |
C1 |
Br3 |
109.040 |
| C2 |
C1 |
H5 |
112.129 |
|
C2 |
C1 |
H6 |
112.129 |
| Br3 |
C1 |
H5 |
106.543 |
|
Br3 |
C1 |
H6 |
106.543 |
| Br4 |
C2 |
H7 |
106.543 |
|
Br4 |
C2 |
H8 |
106.543 |
| H5 |
C1 |
H6 |
110.139 |
|
H7 |
C2 |
H8 |
110.139 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -5224.197774 |
| Energy at 298.15K | -5224.191773 |
| HF Energy | -5218.014527 |
| Nuclear repulsion energy | 453.445071 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3342 |
3342 |
0.27 |
105.57 |
0.75 |
0.86 |
| 2 |
A |
3277 |
3277 |
27.50 |
182.95 |
0.04 |
0.07 |
| 3 |
A |
1613 |
1613 |
0.01 |
6.29 |
0.75 |
0.86 |
| 4 |
A |
1445 |
1445 |
37.32 |
2.43 |
0.54 |
0.70 |
| 5 |
A |
1313 |
1313 |
2.42 |
14.08 |
0.74 |
0.85 |
| 6 |
A |
1107 |
1107 |
0.46 |
4.25 |
0.71 |
0.83 |
| 7 |
A |
990 |
990 |
9.63 |
7.13 |
0.47 |
0.64 |
| 8 |
A |
608 |
608 |
16.43 |
24.38 |
0.10 |
0.18 |
| 9 |
A |
244 |
244 |
1.22 |
1.28 |
0.38 |
0.55 |
| 10 |
A |
86 |
86 |
0.36 |
1.31 |
0.72 |
0.84 |
| 11 |
B |
3354 |
3354 |
3.87 |
26.52 |
0.75 |
0.86 |
| 12 |
B |
3266 |
3266 |
3.64 |
48.99 |
0.75 |
0.86 |
| 13 |
B |
1604 |
1604 |
9.09 |
16.84 |
0.75 |
0.86 |
| 14 |
B |
1413 |
1413 |
86.51 |
1.74 |
0.75 |
0.86 |
| 15 |
B |
1231 |
1231 |
2.63 |
5.54 |
0.75 |
0.86 |
| 16 |
B |
928 |
928 |
18.14 |
1.54 |
0.75 |
0.86 |
| 17 |
B |
646 |
646 |
23.49 |
16.69 |
0.75 |
0.86 |
| 18 |
B |
387 |
387 |
8.07 |
3.14 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13427.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13427.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.310 |
0.692 |
1.177 |
| C2 |
-0.310 |
-0.692 |
1.177 |
| Br3 |
-0.310 |
1.792 |
-0.293 |
| Br4 |
0.310 |
-1.792 |
-0.293 |
| H5 |
0.030 |
1.213 |
2.083 |
| H6 |
1.387 |
0.648 |
1.104 |
| H7 |
-0.030 |
-1.213 |
2.083 |
| H8 |
-1.387 |
-0.648 |
1.104 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5161 | 1.9379 | 2.8857 | 1.0829 | 1.0802 | 2.1368 | 2.1636 |
C2 | 1.5161 | | 2.8857 | 1.9379 | 2.1368 | 2.1636 | 1.0829 | 1.0802 | Br3 | 1.9379 | 2.8857 | | 3.6374 | 2.4688 | 2.4783 | 3.8412 | 3.0102 | Br4 | 2.8857 | 1.9379 | 3.6374 | | 3.8412 | 3.0102 | 2.4688 | 2.4783 | H5 | 1.0829 | 2.1368 | 2.4688 | 3.8412 | | 1.7668 | 2.4273 | 2.5358 | H6 | 1.0802 | 2.1636 | 2.4783 | 3.0102 | 1.7668 | | 2.5358 | 3.0620 | H7 | 2.1368 | 1.0829 | 3.8412 | 2.4688 | 2.4273 | 2.5358 | | 1.7668 | H8 | 2.1636 | 1.0802 | 3.0102 | 2.4783 | 2.5358 | 3.0620 | 1.7668 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
112.758 |
|
C1 |
C2 |
H7 |
109.478 |
| C1 |
C2 |
H8 |
111.788 |
|
C2 |
C1 |
Br3 |
112.758 |
| C2 |
C1 |
H5 |
109.478 |
|
C2 |
C1 |
H6 |
111.788 |
| Br3 |
C1 |
H5 |
106.143 |
|
Br3 |
C1 |
H6 |
106.936 |
| Br4 |
C2 |
H7 |
106.143 |
|
Br4 |
C2 |
H8 |
106.936 |
| H5 |
C1 |
H6 |
109.538 |
|
H7 |
C2 |
H8 |
109.538 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
