Jump to
S1C2
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -158.080727 |
| Energy at 298.15K | -158.073949 |
| HF Energy | -157.297080 |
| Nuclear repulsion energy | 130.591267 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3248 |
3248 |
0.00 |
166.36 |
0.66 |
0.80 |
| 2 |
Ag |
3183 |
3183 |
0.00 |
331.10 |
0.03 |
0.05 |
| 3 |
Ag |
3169 |
3169 |
0.00 |
40.78 |
0.52 |
0.69 |
| 4 |
Ag |
1642 |
1642 |
0.00 |
9.79 |
0.75 |
0.86 |
| 5 |
Ag |
1624 |
1624 |
0.00 |
42.71 |
0.74 |
0.85 |
| 6 |
Ag |
1556 |
1556 |
0.00 |
3.00 |
0.67 |
0.80 |
| 7 |
Ag |
1532 |
1532 |
0.00 |
1.09 |
0.73 |
0.84 |
| 8 |
Ag |
1265 |
1265 |
0.00 |
5.36 |
0.26 |
0.41 |
| 9 |
Ag |
1131 |
1131 |
0.00 |
15.29 |
0.62 |
0.77 |
| 10 |
Ag |
898 |
898 |
0.00 |
14.82 |
0.28 |
0.44 |
| 11 |
Ag |
450 |
450 |
0.00 |
2.98 |
0.16 |
0.28 |
| 12 |
Au |
3250 |
3250 |
177.86 |
0.00 |
0.65 |
0.79 |
| 13 |
Au |
3212 |
3212 |
16.69 |
0.00 |
0.41 |
0.58 |
| 14 |
Au |
1636 |
1636 |
10.17 |
0.00 |
0.00 |
0.00 |
| 15 |
Au |
1402 |
1402 |
0.29 |
0.00 |
0.00 |
0.00 |
| 16 |
Au |
1034 |
1034 |
0.60 |
0.00 |
0.00 |
0.00 |
| 17 |
Au |
783 |
783 |
2.15 |
0.00 |
0.00 |
0.00 |
| 18 |
Au |
244 |
244 |
0.00 |
0.00 |
0.00 |
0.00 |
| 19 |
Au |
130 |
130 |
0.01 |
0.00 |
0.00 |
0.00 |
| 20 |
Bg |
3243 |
3243 |
0.00 |
77.08 |
0.75 |
0.86 |
| 21 |
Bg |
3191 |
3191 |
0.00 |
176.86 |
0.75 |
0.86 |
| 22 |
Bg |
1635 |
1635 |
0.00 |
33.42 |
0.75 |
0.86 |
| 23 |
Bg |
1448 |
1448 |
0.00 |
21.81 |
0.75 |
0.86 |
| 24 |
Bg |
1316 |
1316 |
0.00 |
1.41 |
0.75 |
0.86 |
| 25 |
Bg |
870 |
870 |
0.00 |
0.35 |
0.75 |
0.86 |
| 26 |
Bg |
281 |
281 |
0.00 |
0.02 |
0.75 |
0.86 |
| 27 |
Bu |
3249 |
3249 |
118.77 |
0.00 |
0.75 |
0.86 |
| 28 |
Bu |
3179 |
3179 |
92.65 |
0.00 |
0.00 |
0.00 |
| 29 |
Bu |
3176 |
3176 |
58.08 |
0.00 |
0.00 |
0.00 |
| 30 |
Bu |
1648 |
1648 |
3.71 |
0.00 |
0.75 |
0.86 |
| 31 |
Bu |
1627 |
1627 |
1.17 |
0.00 |
0.75 |
0.86 |
| 32 |
Bu |
1552 |
1552 |
2.76 |
0.00 |
0.00 |
0.00 |
| 33 |
Bu |
1440 |
1440 |
4.82 |
0.00 |
0.00 |
0.00 |
| 34 |
Bu |
1090 |
1090 |
0.63 |
0.00 |
0.00 |
0.00 |
| 35 |
Bu |
1047 |
1047 |
3.60 |
0.00 |
0.00 |
0.00 |
| 36 |
Bu |
278 |
278 |
0.02 |
0.00 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 30829.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 30829.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.422 |
0.639 |
0.000 |
| C2 |
0.422 |
-0.639 |
0.000 |
| C3 |
0.422 |
1.916 |
0.000 |
| C4 |
-0.422 |
-1.916 |
0.000 |
| H5 |
-1.077 |
0.638 |
0.872 |
| H6 |
-1.077 |
0.638 |
-0.872 |
| H7 |
1.077 |
-0.638 |
0.872 |
| H8 |
1.077 |
-0.638 |
-0.872 |
| H9 |
-0.206 |
2.805 |
0.000 |
| H10 |
1.063 |
1.964 |
0.879 |
| H11 |
1.063 |
1.964 |
-0.879 |
| H12 |
0.206 |
-2.805 |
0.000 |
| H13 |
-1.063 |
-1.964 |
0.879 |
| H14 |
-1.063 |
-1.964 |
-0.879 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
| C1 | | 1.5317 | 1.5305 | 2.5555 | 1.0909 | 1.0909 | 2.1537 | 2.1537 | 2.1768 | 2.1757 | 2.1757 | 3.5014 | 2.8216 | 2.8216 |
C2 | 1.5317 | | 2.5555 | 1.5305 | 2.1537 | 2.1537 | 1.0909 | 1.0909 | 3.5014 | 2.8216 | 2.8216 | 2.1768 | 2.1757 | 2.1757 | C3 | 1.5305 | 2.5555 | | 3.9242 | 2.1543 | 2.1543 | 2.7774 | 2.7774 | 1.0886 | 1.0893 | 1.0893 | 4.7265 | 4.2468 | 4.2468 | C4 | 2.5555 | 1.5305 | 3.9242 | | 2.7774 | 2.7774 | 2.1543 | 2.1543 | 4.7265 | 4.2468 | 4.2468 | 1.0886 | 1.0893 | 1.0893 | H5 | 1.0909 | 2.1537 | 2.1543 | 2.7774 | | 1.7443 | 2.5036 | 3.0513 | 2.4933 | 2.5179 | 3.0670 | 3.7768 | 2.6021 | 3.1364 | H6 | 1.0909 | 2.1537 | 2.1543 | 2.7774 | 1.7443 | | 3.0513 | 2.5036 | 2.4933 | 3.0670 | 2.5179 | 3.7768 | 3.1364 | 2.6021 | H7 | 2.1537 | 1.0909 | 2.7774 | 2.1543 | 2.5036 | 3.0513 | | 1.7443 | 3.7768 | 2.6021 | 3.1364 | 2.4933 | 2.5179 | 3.0670 | H8 | 2.1537 | 1.0909 | 2.7774 | 2.1543 | 3.0513 | 2.5036 | 1.7443 | | 3.7768 | 3.1364 | 2.6021 | 2.4933 | 3.0670 | 2.5179 | H9 | 2.1768 | 3.5014 | 1.0886 | 4.7265 | 2.4933 | 2.4933 | 3.7768 | 3.7768 | | 1.7585 | 1.7585 | 5.6259 | 4.9250 | 4.9250 | H10 | 2.1757 | 2.8216 | 1.0893 | 4.2468 | 2.5179 | 3.0670 | 2.6021 | 3.1364 | 1.7585 | | 1.7580 | 4.9250 | 4.4670 | 4.8004 | H11 | 2.1757 | 2.8216 | 1.0893 | 4.2468 | 3.0670 | 2.5179 | 3.1364 | 2.6021 | 1.7585 | 1.7580 | | 4.9250 | 4.8004 | 4.4670 | H12 | 3.5014 | 2.1768 | 4.7265 | 1.0886 | 3.7768 | 3.7768 | 2.4933 | 2.4933 | 5.6259 | 4.9250 | 4.9250 | | 1.7585 | 1.7585 | H13 | 2.8216 | 2.1757 | 4.2468 | 1.0893 | 2.6021 | 3.1364 | 2.5179 | 3.0670 | 4.9250 | 4.4670 | 4.8004 | 1.7585 | | 1.7580 | H14 | 2.8216 | 2.1757 | 4.2468 | 1.0893 | 3.1364 | 2.6021 | 3.0670 | 2.5179 | 4.9250 | 4.8004 | 4.4670 | 1.7585 | 1.7580 | |
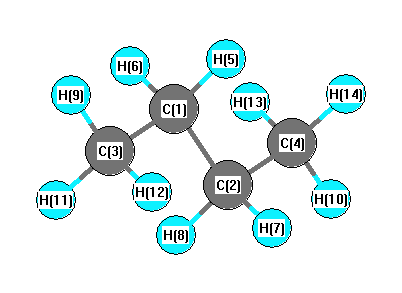 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
113.134 |
|
C1 |
C2 |
H7 |
109.257 |
| C1 |
C2 |
H8 |
109.257 |
|
C1 |
C3 |
H9 |
111.318 |
| C1 |
C3 |
H11 |
111.181 |
|
C1 |
C3 |
H12 |
30.835 |
| C2 |
C1 |
C3 |
113.134 |
|
C2 |
C1 |
H5 |
109.257 |
| C2 |
C1 |
H6 |
109.257 |
|
C2 |
C4 |
H10 |
17.227 |
| C2 |
C4 |
H13 |
111.181 |
|
C2 |
C4 |
H14 |
111.181 |
| C3 |
C1 |
H5 |
109.395 |
|
C3 |
C1 |
H6 |
109.395 |
| C4 |
C2 |
H7 |
109.395 |
|
C4 |
C2 |
H8 |
109.395 |
| H5 |
C1 |
H6 |
106.165 |
|
H7 |
C2 |
H8 |
106.165 |
| H9 |
C3 |
H11 |
107.693 |
|
H9 |
C3 |
H12 |
142.152 |
| H10 |
C4 |
H13 |
94.537 |
|
H10 |
C4 |
H14 |
114.402 |
| H11 |
C3 |
H12 |
94.057 |
|
H13 |
C4 |
H14 |
107.596 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -158.079196 |
| Energy at 298.15K | -158.072462 |
| Nuclear repulsion energy | 132.311211 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3263 |
3263 |
61.54 |
62.26 |
0.74 |
0.85 |
| 2 |
A |
3246 |
3246 |
0.48 |
140.56 |
0.63 |
0.77 |
| 3 |
A |
3209 |
3209 |
37.38 |
163.65 |
0.36 |
0.52 |
| 4 |
A |
3187 |
3187 |
12.95 |
290.79 |
0.07 |
0.13 |
| 5 |
A |
3176 |
3176 |
41.42 |
63.84 |
0.47 |
0.64 |
| 6 |
A |
1648 |
1648 |
0.10 |
2.74 |
0.73 |
0.85 |
| 7 |
A |
1644 |
1644 |
3.60 |
10.01 |
0.75 |
0.86 |
| 8 |
A |
1625 |
1625 |
0.24 |
25.84 |
0.74 |
0.85 |
| 9 |
A |
1553 |
1553 |
2.15 |
3.16 |
0.74 |
0.85 |
| 10 |
A |
1517 |
1517 |
0.30 |
2.06 |
0.71 |
0.83 |
| 11 |
A |
1428 |
1428 |
0.28 |
14.72 |
0.72 |
0.84 |
| 12 |
A |
1302 |
1302 |
0.12 |
1.00 |
0.21 |
0.35 |
| 13 |
A |
1156 |
1156 |
0.07 |
6.86 |
0.62 |
0.77 |
| 14 |
A |
1074 |
1074 |
0.18 |
3.13 |
0.23 |
0.37 |
| 15 |
A |
890 |
890 |
0.11 |
7.50 |
0.18 |
0.30 |
| 16 |
A |
846 |
846 |
0.99 |
7.01 |
0.13 |
0.24 |
| 17 |
A |
343 |
343 |
0.01 |
0.44 |
0.05 |
0.09 |
| 18 |
A |
280 |
280 |
0.03 |
0.08 |
0.20 |
0.33 |
| 19 |
A |
119 |
119 |
0.00 |
0.07 |
0.75 |
0.86 |
| 20 |
B |
3254 |
3254 |
92.16 |
8.70 |
0.75 |
0.86 |
| 21 |
B |
3249 |
3249 |
127.51 |
0.36 |
0.75 |
0.86 |
| 22 |
B |
3211 |
3211 |
32.14 |
53.91 |
0.75 |
0.86 |
| 23 |
B |
3182 |
3182 |
30.29 |
1.97 |
0.75 |
0.86 |
| 24 |
B |
3172 |
3172 |
31.76 |
30.52 |
0.75 |
0.86 |
| 25 |
B |
1643 |
1643 |
7.12 |
6.25 |
0.75 |
0.86 |
| 26 |
B |
1635 |
1635 |
4.91 |
2.37 |
0.75 |
0.86 |
| 27 |
B |
1622 |
1622 |
0.35 |
36.46 |
0.75 |
0.86 |
| 28 |
B |
1558 |
1558 |
4.77 |
3.04 |
0.75 |
0.86 |
| 29 |
B |
1494 |
1494 |
4.59 |
0.77 |
0.75 |
0.86 |
| 30 |
B |
1399 |
1399 |
0.47 |
4.41 |
0.75 |
0.86 |
| 31 |
B |
1253 |
1253 |
1.91 |
1.97 |
0.75 |
0.86 |
| 32 |
B |
1045 |
1045 |
0.95 |
0.32 |
0.75 |
0.86 |
| 33 |
B |
1025 |
1025 |
1.48 |
8.97 |
0.75 |
0.86 |
| 34 |
B |
804 |
804 |
2.51 |
0.24 |
0.75 |
0.86 |
| 35 |
B |
461 |
461 |
0.18 |
0.22 |
0.75 |
0.86 |
| 36 |
B |
230 |
230 |
0.02 |
0.01 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 30869.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 30869.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is C2h
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
