Jump to
S1C2
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -155.665695 |
| Energy at 298.15K | -155.660043 |
| Nuclear repulsion energy | 104.140929 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
Geometric Data calculated at CBS-Q
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -155.660670 |
| Energy at 298.15K | -155.666514 |
| HF Energy | -154.911980 |
| Nuclear repulsion energy | 105.190002 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3403 |
3403 |
9.61 |
|
|
|
| 2 |
A |
3332 |
3332 |
5.75 |
|
|
|
| 3 |
A |
3313 |
3313 |
32.58 |
|
|
|
| 4 |
A |
1857 |
1857 |
1.26 |
|
|
|
| 5 |
A |
1589 |
1589 |
9.05 |
|
|
|
| 6 |
A |
1449 |
1449 |
0.02 |
|
|
|
| 7 |
A |
1154 |
1154 |
0.44 |
|
|
|
| 8 |
A |
1113 |
1113 |
1.60 |
|
|
|
| 9 |
A |
1063 |
1063 |
9.12 |
|
|
|
| 10 |
A |
940 |
940 |
0.15 |
|
|
|
| 11 |
A |
824 |
824 |
5.94 |
|
|
|
| 12 |
A |
296 |
296 |
0.00 |
|
|
|
| 13 |
A |
176 |
176 |
0.06 |
|
|
|
| 14 |
B |
3401 |
3401 |
29.54 |
|
|
|
| 15 |
B |
3323 |
3323 |
18.23 |
|
|
|
| 16 |
B |
3304 |
3304 |
7.33 |
|
|
|
| 17 |
B |
1857 |
1857 |
4.23 |
|
|
|
| 18 |
B |
1559 |
1559 |
0.67 |
|
|
|
| 19 |
B |
1418 |
1418 |
1.38 |
|
|
|
| 20 |
B |
1192 |
1192 |
5.71 |
|
|
|
| 21 |
B |
1135 |
1135 |
23.80 |
|
|
|
| 22 |
B |
1064 |
1064 |
80.60 |
|
|
|
| 23 |
B |
673 |
673 |
7.63 |
|
|
|
| 24 |
B |
498 |
498 |
12.39 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19965.8 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 19965.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.118 |
0.731 |
0.539 |
| C2 |
-0.118 |
-0.731 |
0.539 |
| C3 |
-0.118 |
1.544 |
-0.479 |
| C4 |
0.118 |
-1.544 |
-0.479 |
| H5 |
0.496 |
1.146 |
1.463 |
| H6 |
-0.496 |
-1.146 |
1.463 |
| H7 |
0.091 |
2.600 |
-0.415 |
| H8 |
-0.537 |
1.188 |
-1.407 |
| H9 |
-0.091 |
-2.600 |
-0.415 |
| H10 |
0.537 |
-1.188 |
-1.407 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4799 | 1.3239 | 2.4916 | 1.0816 | 2.1796 | 2.0984 | 2.1037 | 3.4703 | 2.7648 |
C2 | 1.4799 | | 2.4916 | 1.3239 | 2.1796 | 1.0816 | 3.4703 | 2.7648 | 2.0984 | 2.1037 | C3 | 1.3239 | 2.4916 | | 3.0962 | 2.0755 | 3.3388 | 1.0784 | 1.0788 | 4.1439 | 2.9584 | C4 | 2.4916 | 1.3239 | 3.0962 | | 3.3388 | 2.0755 | 4.1439 | 2.9584 | 1.0784 | 1.0788 | H5 | 1.0816 | 2.1796 | 2.0755 | 3.3388 | | 2.4962 | 2.4093 | 3.0510 | 4.2306 | 3.6997 | H6 | 2.1796 | 1.0816 | 3.3388 | 2.0755 | 2.4962 | | 4.2306 | 3.6997 | 2.4093 | 3.0510 | H7 | 2.0984 | 3.4703 | 1.0784 | 4.1439 | 2.4093 | 4.2306 | | 1.8365 | 5.2025 | 3.9409 | H8 | 2.1037 | 2.7648 | 1.0788 | 2.9584 | 3.0510 | 3.6997 | 1.8365 | | 3.9409 | 2.6076 | H9 | 3.4703 | 2.0984 | 4.1439 | 1.0784 | 4.2306 | 2.4093 | 5.2025 | 3.9409 | | 1.8365 | H10 | 2.7648 | 2.1037 | 2.9584 | 1.0788 | 3.6997 | 3.0510 | 3.9409 | 2.6076 | 1.8365 | |
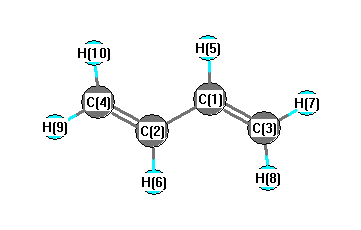 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
125.313 |
|
C1 |
C2 |
H6 |
115.746 |
| C1 |
C3 |
H7 |
121.397 |
|
C1 |
C3 |
H8 |
121.883 |
| C2 |
C1 |
C3 |
125.313 |
|
C2 |
C1 |
H5 |
115.746 |
| C2 |
C4 |
H9 |
121.397 |
|
C2 |
C4 |
H10 |
121.883 |
| C3 |
C1 |
H5 |
118.931 |
|
C4 |
C2 |
H6 |
118.931 |
| H7 |
C3 |
H8 |
116.716 |
|
H9 |
C4 |
H10 |
116.716 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
