Jump to
S1C2
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -612.734202 |
| Energy at 298.15K | -612.728602 |
| HF Energy | -611.812313 |
| Nuclear repulsion energy | 143.627632 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3348 |
3348 |
1.27 |
|
|
|
| 2 |
A |
3265 |
3265 |
12.44 |
|
|
|
| 3 |
A |
3187 |
3187 |
58.40 |
|
|
|
| 4 |
A |
2033 |
2033 |
230.53 |
|
|
|
| 5 |
A |
1590 |
1590 |
9.88 |
|
|
|
| 6 |
A |
1537 |
1537 |
14.29 |
|
|
|
| 7 |
A |
1402 |
1402 |
34.11 |
|
|
|
| 8 |
A |
1311 |
1311 |
5.16 |
|
|
|
| 9 |
A |
1142 |
1142 |
7.78 |
|
|
|
| 10 |
A |
1126 |
1126 |
27.07 |
|
|
|
| 11 |
A |
881 |
881 |
38.62 |
|
|
|
| 12 |
A |
767 |
767 |
22.62 |
|
|
|
| 13 |
A |
494 |
494 |
14.89 |
|
|
|
| 14 |
A |
308 |
308 |
14.14 |
|
|
|
| 15 |
A |
58 |
58 |
21.83 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11223.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 11223.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.027 |
0.657 |
0.128 |
| C2 |
1.168 |
-0.343 |
0.155 |
| Cl3 |
-1.550 |
-0.160 |
-0.069 |
| O4 |
2.259 |
-0.050 |
-0.188 |
| H5 |
-0.001 |
1.200 |
1.066 |
| H6 |
0.169 |
1.351 |
-0.688 |
| H7 |
0.945 |
-1.314 |
0.609 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.5178 | 1.7878 | 2.3622 | 1.0841 | 1.0806 | 2.2269 |
C2 | 1.5178 | | 2.7342 | 1.1801 | 2.1395 | 2.1402 | 1.0946 | Cl3 | 1.7878 | 2.7342 | | 3.8126 | 2.3539 | 2.3714 | 2.8318 | O4 | 2.3622 | 1.1801 | 3.8126 | | 2.8705 | 2.5653 | 1.9894 | H5 | 1.0841 | 2.1395 | 2.3539 | 2.8705 | | 1.7687 | 2.7244 | H6 | 1.0806 | 2.1402 | 2.3714 | 2.5653 | 1.7687 | | 3.0639 | H7 | 2.2269 | 1.0946 | 2.8318 | 1.9894 | 2.7244 | 3.0639 | |
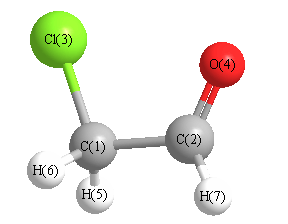 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O4 |
121.726 |
|
C1 |
C2 |
H7 |
116.013 |
| C2 |
C1 |
Cl3 |
111.352 |
|
C2 |
C1 |
H5 |
109.503 |
| C2 |
C1 |
H6 |
109.765 |
|
Cl3 |
C1 |
H5 |
107.562 |
| Cl3 |
C1 |
H6 |
109.032 |
|
O4 |
C2 |
H7 |
121.952 |
| H5 |
C1 |
H6 |
109.588 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -612.732462 |
| Energy at 298.15K | -612.736242 |
| HF Energy | -611.810302 |
| Nuclear repulsion energy | 147.598024 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3244 |
3244 |
24.24 |
|
|
|
| 2 |
A' |
3132 |
3132 |
91.72 |
|
|
|
| 3 |
A' |
2051 |
2051 |
199.78 |
|
|
|
| 4 |
A' |
1588 |
1588 |
19.69 |
|
|
|
| 5 |
A' |
1533 |
1533 |
27.00 |
|
|
|
| 6 |
A' |
1455 |
1455 |
20.50 |
|
|
|
| 7 |
A' |
1025 |
1025 |
4.57 |
|
|
|
| 8 |
A' |
834 |
834 |
9.29 |
|
|
|
| 9 |
A' |
681 |
681 |
65.92 |
|
|
|
| 10 |
A' |
233 |
233 |
3.45 |
|
|
|
| 11 |
A" |
3298 |
3298 |
3.18 |
|
|
|
| 12 |
A" |
1334 |
1334 |
0.89 |
|
|
|
| 13 |
A" |
1145 |
1145 |
2.26 |
|
|
|
| 14 |
A" |
767 |
767 |
0.58 |
|
|
|
| 15 |
A" |
167 |
167 |
2.45 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11242.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 11242.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.915 |
0.000 |
| C2 |
1.366 |
0.262 |
0.000 |
| Cl3 |
-1.346 |
-0.235 |
0.000 |
| O4 |
1.585 |
-0.895 |
0.000 |
| H5 |
-0.087 |
1.549 |
0.877 |
| H6 |
-0.087 |
1.549 |
-0.877 |
| H7 |
2.182 |
0.998 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.5141 | 1.7705 | 2.4052 | 1.0855 | 1.0855 | 2.1839 |
C2 | 1.5141 | | 2.7575 | 1.1770 | 2.1297 | 2.1297 | 1.0989 | Cl3 | 1.7705 | 2.7575 | | 3.0039 | 2.3531 | 2.3531 | 3.7377 | O4 | 2.4052 | 1.1770 | 3.0039 | | 3.0874 | 3.0874 | 1.9847 | H5 | 1.0855 | 2.1297 | 2.3531 | 3.0874 | | 1.7540 | 2.4943 | H6 | 1.0855 | 2.1297 | 2.3531 | 3.0874 | 1.7540 | | 2.4943 | H7 | 2.1839 | 1.0989 | 3.7377 | 1.9847 | 2.4943 | 2.4943 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O4 |
126.233 |
|
C1 |
C2 |
H7 |
112.416 |
| C2 |
C1 |
Cl3 |
113.950 |
|
C2 |
C1 |
H5 |
108.900 |
| C2 |
C1 |
H6 |
108.900 |
|
Cl3 |
C1 |
H5 |
108.568 |
| Cl3 |
C1 |
H6 |
108.568 |
|
O4 |
C2 |
H7 |
121.351 |
| H5 |
C1 |
H6 |
107.784 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
