return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH(C2H5)2 (diethylamine)

using model chemistry: CBS-Q

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CBS-Q
 hartrees
Energy at 0K-213.339468
Energy at 298.15K-213.331560
HF Energy-212.312669
Nuclear repulsion energy188.956375
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3742 3742 0.49      
2 A' 3272 3272 78.84      
3 A' 3249 3249 119.57      
4 A' 3215 3215 41.60      
5 A' 3184 3184 23.51      
6 A' 3110 3110 182.14      
7 A' 1668 1668 0.17      
8 A' 1635 1635 2.32      
9 A' 1627 1627 8.58      
10 A' 1575 1575 2.27      
11 A' 1528 1528 0.20      
12 A' 1441 1441 6.01      
13 A' 1338 1338 2.35      
14 A' 1274 1274 21.01      
15 A' 1129 1129 6.16      
16 A' 968 968 5.71      
17 A' 891 891 7.64      
18 A' 838 838 80.60      
19 A' 452 452 0.52      
20 A' 277 277 1.54      
21 A' 201 201 0.92      
22 A' 123 123 1.68      
23 A" 3271 3271 12.17      
24 A" 3248 3248 31.03      
25 A" 3208 3208 1.30      
26 A" 3183 3183 40.80      
27 A" 3100 3100 5.46      
28 A" 1666 1666 42.15      
29 A" 1654 1654 0.15      
30 A" 1632 1632 3.29      
31 A" 1622 1622 7.18      
32 A" 1549 1549 10.43      
33 A" 1488 1488 30.78      
34 A" 1399 1399 2.77      
35 A" 1256 1256 51.58      
36 A" 1212 1212 8.44      
37 A" 1150 1150 0.58      
38 A" 1005 1005 0.48      
39 A" 864 864 0.44      
40 A" 456 456 0.34      
41 A" 278 278 0.62      
42 A" 118 118 1.16      

Unscaled Zero Point Vibrational Energy (zpe) 35047.3 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 35047.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CBS-Q
ABC
0.58919 0.07015 0.06609

See section I.F.4 to change rotational constant units
Geometric Data calculated at CBS-Q

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.029 -0.273 0.000
C2 0.015 0.519 1.213
C3 0.015 0.519 -1.213
C4 0.015 -0.375 2.446
C5 0.015 -0.375 -2.446
H6 -0.760 -0.888 0.000
H7 -0.840 1.204 1.253
H8 0.908 1.140 1.214
H9 -0.840 1.204 -1.253
H10 0.908 1.140 -1.214
H11 0.033 0.221 3.355
H12 -0.878 -0.999 2.479
H13 0.881 -1.030 2.442
H14 0.033 0.221 -3.355
H15 -0.878 -0.999 -2.479
H16 0.881 -1.030 -2.442

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
N11.44891.44892.44782.44781.00012.12282.06002.12282.06003.39102.73742.69513.39102.73742.6951
C21.44892.42661.52283.76672.01281.09631.08792.69882.66022.16242.16862.15904.57804.09064.0635
C31.44892.42663.76671.52282.01282.69882.66021.09631.08794.57804.09064.06352.16242.16862.1590
C42.44781.52283.76674.89122.61592.15592.14754.11184.06071.08751.08921.08625.83115.04325.0067
C52.44783.76671.52284.89122.61594.11184.06072.15592.14755.83115.04325.00671.08751.08921.0862
H61.00012.01282.01282.61592.61592.43992.89292.43992.89293.62122.48402.94543.62122.48402.9454
H72.12281.09632.69882.15594.11182.43991.74982.50633.02462.47952.52153.06104.79214.33394.6486
H82.06001.08792.66022.14754.06072.89291.74983.02462.42842.48863.06012.49404.74214.62644.2520
H92.12282.69881.09634.11182.15592.43992.50633.02461.74984.79214.33394.64862.47952.52153.0610
H102.06002.66021.08794.06072.14752.89293.02462.42841.74984.74214.62644.25202.48863.06012.4940
H113.39102.16244.57801.08755.83113.62122.47952.48864.79214.74211.75661.76576.70986.02915.9906
H122.73742.16864.09061.08925.04322.48402.52153.06014.33394.62641.75661.75946.02914.95755.2256
H132.69512.15904.06351.08625.00672.94543.06102.49404.64864.25201.76571.75945.99065.22564.8837
H143.39104.57802.16245.83111.08753.62124.79214.74212.47952.48866.70986.02915.99061.75661.7657
H152.73744.09062.16865.04321.08922.48404.33394.62642.52153.06016.02914.95755.22561.75661.7594
H162.69514.06352.15905.00671.08622.94544.64864.25203.06102.49405.99065.22564.88371.76571.7594

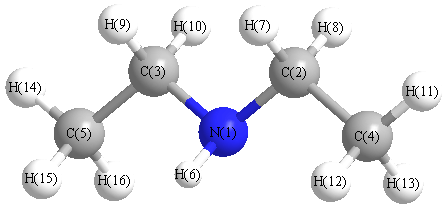
picture of diethylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C4 110.892 N1 C2 H7 112.298
N1 C2 H8 107.743 N1 C3 C5 110.892
N1 C3 H9 112.298 N1 C3 H10 107.743
C2 N1 C3 113.736 C2 N1 H6 109.180
C2 C4 H11 110.766 C2 C4 H12 111.163
C2 C4 H13 110.578 C3 N1 H6 109.180
C3 C5 H14 110.766 C3 C5 H15 111.163
C3 C5 H16 110.578 C4 C2 H7 109.735
C4 C2 H8 109.557 C5 C3 H8 151.121
C5 C3 H10 109.557 H7 C2 H8 106.473
H9 C3 H10 106.473 H11 C4 H12 107.609
H11 C4 H13 108.649 H12 C4 H13 107.962
H14 C5 H15 107.609 H14 C5 H16 108.649
H15 C5 H16 107.962
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability