Jump to
S1C2
S1C3
Vibrational Frequencies calculated at CBS-Q
Geometric Data calculated at CBS-Q
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at CBS-Q
Geometric Data calculated at CBS-Q
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -210.036944 |
| Energy at 298.15K | -210.030939 |
| HF Energy | -209.095805 |
| Nuclear repulsion energy | 135.145252 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4073 |
4073 |
63.21 |
|
|
|
| 2 |
A |
3831 |
3831 |
2.16 |
|
|
|
| 3 |
A |
3745 |
3745 |
0.80 |
|
|
|
| 4 |
A |
3261 |
3261 |
68.44 |
|
|
|
| 5 |
A |
3246 |
3246 |
41.94 |
|
|
|
| 6 |
A |
3157 |
3157 |
99.12 |
|
|
|
| 7 |
A |
3139 |
3139 |
71.95 |
|
|
|
| 8 |
A |
1843 |
1843 |
38.19 |
|
|
|
| 9 |
A |
1655 |
1655 |
0.16 |
|
|
|
| 10 |
A |
1638 |
1638 |
3.19 |
|
|
|
| 11 |
A |
1588 |
1588 |
67.62 |
|
|
|
| 12 |
A |
1543 |
1543 |
31.34 |
|
|
|
| 13 |
A |
1501 |
1501 |
10.03 |
|
|
|
| 14 |
A |
1450 |
1450 |
3.45 |
|
|
|
| 15 |
A |
1366 |
1366 |
37.88 |
|
|
|
| 16 |
A |
1287 |
1287 |
12.57 |
|
|
|
| 17 |
A |
1225 |
1225 |
88.89 |
|
|
|
| 18 |
A |
1160 |
1160 |
19.46 |
|
|
|
| 19 |
A |
1096 |
1096 |
27.69 |
|
|
|
| 20 |
A |
1023 |
1023 |
82.96 |
|
|
|
| 21 |
A |
941 |
941 |
24.34 |
|
|
|
| 22 |
A |
909 |
909 |
83.98 |
|
|
|
| 23 |
A |
573 |
573 |
53.54 |
|
|
|
| 24 |
A |
523 |
523 |
160.22 |
|
|
|
| 25 |
A |
347 |
347 |
1.43 |
|
|
|
| 26 |
A |
291 |
291 |
11.98 |
|
|
|
| 27 |
A |
201 |
201 |
5.61 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23304.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 23304.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.379 |
-0.555 |
0.120 |
| C2 |
-0.636 |
0.637 |
-0.278 |
| C3 |
0.780 |
0.543 |
0.277 |
| O4 |
1.425 |
-0.609 |
-0.173 |
| H5 |
-1.660 |
-0.502 |
1.079 |
| H6 |
-2.205 |
-0.674 |
-0.429 |
| H7 |
-1.102 |
1.572 |
0.045 |
| H8 |
-0.582 |
0.652 |
-1.363 |
| H9 |
1.372 |
1.394 |
-0.047 |
| H10 |
0.747 |
0.564 |
1.371 |
| H11 |
0.811 |
-1.325 |
-0.106 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| N1 | | 1.4599 | 2.4275 | 2.8191 | 1.0008 | 0.9992 | 2.1464 | 2.0715 | 3.3757 | 2.7089 | 2.3318 |
C2 | 1.4599 | | 1.5238 | 2.4099 | 2.0457 | 2.0507 | 1.0936 | 1.0872 | 2.1582 | 2.1529 | 2.4436 | C3 | 2.4275 | 1.5238 | | 1.3947 | 2.7728 | 3.3006 | 2.1579 | 2.1357 | 1.0862 | 1.0944 | 1.9072 | O4 | 2.8191 | 2.4099 | 1.3947 | | 3.3307 | 3.6395 | 3.3449 | 2.6524 | 2.0074 | 2.0544 | 0.9456 | H5 | 1.0008 | 2.0457 | 2.7728 | 3.3307 | | 1.6124 | 2.3830 | 2.9078 | 3.7491 | 2.6486 | 2.8610 | H6 | 0.9992 | 2.0507 | 3.3006 | 3.6395 | 1.6124 | | 2.5466 | 2.2946 | 4.1498 | 3.6728 | 3.1026 | H7 | 2.1464 | 1.0936 | 2.1579 | 3.3449 | 2.3830 | 2.5466 | | 1.7607 | 2.4827 | 2.4885 | 3.4749 | H8 | 2.0715 | 1.0872 | 2.1357 | 2.6524 | 2.9078 | 2.2946 | 1.7607 | | 2.4705 | 3.0417 | 2.7262 | H9 | 3.3757 | 2.1582 | 1.0862 | 2.0074 | 3.7491 | 4.1498 | 2.4827 | 2.4705 | | 1.7582 | 2.7769 | H10 | 2.7089 | 2.1529 | 1.0944 | 2.0544 | 2.6486 | 3.6728 | 2.4885 | 3.0417 | 1.7582 | | 2.3988 | H11 | 2.3318 | 2.4436 | 1.9072 | 0.9456 | 2.8610 | 3.1026 | 3.4749 | 2.7262 | 2.7769 | 2.3988 | |
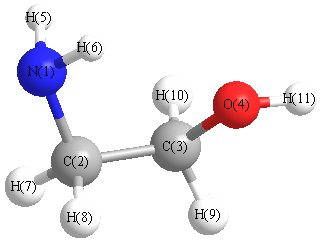 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
108.875 |
|
N1 |
C2 |
H7 |
113.621 |
| N1 |
C2 |
H8 |
107.943 |
|
C2 |
N1 |
H5 |
111.102 |
| C2 |
N1 |
H6 |
111.629 |
|
C2 |
C3 |
O4 |
111.253 |
| C2 |
C3 |
H9 |
110.444 |
|
C2 |
C3 |
H10 |
109.541 |
| C3 |
C2 |
H7 |
109.978 |
|
C3 |
C2 |
H8 |
108.613 |
| C3 |
O4 |
H11 |
107.613 |
|
O4 |
C3 |
H9 |
107.370 |
| O4 |
C3 |
H10 |
110.672 |
|
H5 |
N1 |
H6 |
107.455 |
| H7 |
C2 |
H8 |
107.679 |
|
H9 |
C3 |
H10 |
107.473 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
