Jump to
S1C2
Vibrational Frequencies calculated at CBS-Q
Geometric Data calculated at CBS-Q
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -235.321141 |
| Energy at 298.15K | -235.335535 |
| Nuclear repulsion energy | 255.109745 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3274 |
3274 |
108.56 |
|
|
|
| 2 |
A' |
3248 |
3248 |
56.74 |
|
|
|
| 3 |
A' |
3245 |
3245 |
54.46 |
|
|
|
| 4 |
A' |
3239 |
3239 |
8.96 |
|
|
|
| 5 |
A' |
3223 |
3223 |
23.30 |
|
|
|
| 6 |
A' |
3204 |
3204 |
22.27 |
|
|
|
| 7 |
A' |
3178 |
3178 |
35.61 |
|
|
|
| 8 |
A' |
3171 |
3171 |
33.51 |
|
|
|
| 9 |
A' |
1656 |
1656 |
0.91 |
|
|
|
| 10 |
A' |
1640 |
1640 |
1.66 |
|
|
|
| 11 |
A' |
1629 |
1629 |
4.21 |
|
|
|
| 12 |
A' |
1617 |
1617 |
0.10 |
|
|
|
| 13 |
A' |
1558 |
1558 |
0.88 |
|
|
|
| 14 |
A' |
1545 |
1545 |
6.31 |
|
|
|
| 15 |
A' |
1452 |
1452 |
3.54 |
|
|
|
| 16 |
A' |
1421 |
1421 |
1.41 |
|
|
|
| 17 |
A' |
1342 |
1342 |
3.26 |
|
|
|
| 18 |
A' |
1314 |
1314 |
0.50 |
|
|
|
| 19 |
A' |
1086 |
1086 |
0.30 |
|
|
|
| 20 |
A' |
1060 |
1060 |
0.29 |
|
|
|
| 21 |
A' |
1024 |
1024 |
0.22 |
|
|
|
| 22 |
A' |
989 |
989 |
0.22 |
|
|
|
| 23 |
A' |
938 |
938 |
0.47 |
|
|
|
| 24 |
A' |
759 |
759 |
0.66 |
|
|
|
| 25 |
A' |
650 |
650 |
0.80 |
|
|
|
| 26 |
A' |
419 |
419 |
0.31 |
|
|
|
| 27 |
A' |
371 |
371 |
0.00 |
|
|
|
| 28 |
A' |
149 |
149 |
0.00 |
|
|
|
| 29 |
A" |
3250 |
3250 |
27.34 |
|
|
|
| 30 |
A" |
3243 |
3243 |
67.03 |
|
|
|
| 31 |
A" |
3233 |
3233 |
7.13 |
|
|
|
| 32 |
A" |
3201 |
3201 |
92.30 |
|
|
|
| 33 |
A" |
1646 |
1646 |
4.52 |
|
|
|
| 34 |
A" |
1622 |
1622 |
0.13 |
|
|
|
| 35 |
A" |
1617 |
1617 |
1.79 |
|
|
|
| 36 |
A" |
1407 |
1407 |
0.00 |
|
|
|
| 37 |
A" |
1379 |
1379 |
0.39 |
|
|
|
| 38 |
A" |
1335 |
1335 |
0.11 |
|
|
|
| 39 |
A" |
1303 |
1303 |
1.83 |
|
|
|
| 40 |
A" |
1184 |
1184 |
0.04 |
|
|
|
| 41 |
A" |
1071 |
1071 |
0.00 |
|
|
|
| 42 |
A" |
1007 |
1007 |
1.72 |
|
|
|
| 43 |
A" |
954 |
954 |
0.05 |
|
|
|
| 44 |
A" |
848 |
848 |
0.09 |
|
|
|
| 45 |
A" |
414 |
414 |
0.03 |
|
|
|
| 46 |
A" |
321 |
321 |
0.01 |
|
|
|
| 47 |
A" |
292 |
292 |
0.00 |
|
|
|
| 48 |
A" |
244 |
244 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38984.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 38984.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.089 |
0.403 |
0.000 |
| C2 |
0.329 |
-1.740 |
0.000 |
| C3 |
-0.089 |
-0.716 |
1.082 |
| C4 |
-0.089 |
-0.716 |
-1.082 |
| C5 |
1.219 |
1.198 |
0.000 |
| C6 |
-1.289 |
1.345 |
0.000 |
| H7 |
1.398 |
-1.933 |
0.000 |
| H8 |
-0.188 |
-2.695 |
0.000 |
| H9 |
0.579 |
-0.582 |
1.930 |
| H10 |
0.579 |
-0.582 |
-1.930 |
| H11 |
-1.089 |
-0.913 |
1.463 |
| H12 |
-1.089 |
-0.913 |
-1.463 |
| H13 |
2.089 |
0.543 |
0.000 |
| H14 |
-2.227 |
0.791 |
0.000 |
| H15 |
1.286 |
1.835 |
-0.881 |
| H16 |
1.286 |
1.835 |
0.881 |
| H17 |
-1.283 |
1.988 |
0.880 |
| H18 |
-1.283 |
1.988 |
-0.880 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 2.1827 | 1.5562 | 1.5562 | 1.5305 | 1.5259 | 2.7690 | 3.0992 | 2.2670 | 2.2670 | 2.2069 | 2.2069 | 2.1822 | 2.1726 | 2.1724 | 2.1724 | 2.1710 | 2.1710 |
C2 | 2.1827 | | 1.5467 | 1.5467 | 3.0693 | 3.4833 | 1.0865 | 1.0862 | 2.2642 | 2.2642 | 2.1982 | 2.1982 | 2.8823 | 3.5967 | 3.8045 | 3.8045 | 4.1554 | 4.1554 | C3 | 1.5562 | 1.5467 | | 2.1634 | 2.5579 | 2.6189 | 2.2051 | 2.2575 | 1.0878 | 3.0875 | 1.0879 | 2.7409 | 2.7381 | 2.8305 | 3.5003 | 2.9054 | 2.9628 | 3.5476 | C4 | 1.5562 | 1.5467 | 2.1634 | | 2.5579 | 2.6189 | 2.2051 | 2.2575 | 3.0875 | 1.0878 | 2.7409 | 1.0879 | 2.7381 | 2.8305 | 2.9054 | 3.5003 | 3.5476 | 2.9628 | C5 | 1.5305 | 3.0693 | 2.5579 | 2.5579 | | 2.5123 | 3.1361 | 4.1392 | 2.7018 | 2.7018 | 3.4526 | 3.4526 | 1.0887 | 3.4692 | 1.0895 | 1.0895 | 2.7673 | 2.7673 | C6 | 1.5259 | 3.4833 | 2.6189 | 2.6189 | 2.5123 | | 4.2388 | 4.1873 | 3.3057 | 3.3057 | 2.6980 | 2.6980 | 3.4717 | 1.0887 | 2.7654 | 2.7654 | 1.0899 | 1.0899 | H7 | 2.7690 | 1.0865 | 2.2051 | 2.2051 | 3.1361 | 4.2388 | | 1.7596 | 2.4939 | 2.4939 | 3.0600 | 3.0600 | 2.5707 | 4.5344 | 3.8719 | 3.8719 | 4.8309 | 4.8309 | H8 | 3.0992 | 1.0862 | 2.2575 | 2.2575 | 4.1392 | 4.1873 | 1.7596 | | 2.9626 | 2.9626 | 2.4749 | 2.4749 | 3.9584 | 4.0386 | 4.8450 | 4.8450 | 4.8891 | 4.8891 | H9 | 2.2670 | 2.2642 | 1.0878 | 3.0875 | 2.7018 | 3.3057 | 2.4939 | 2.9626 | | 3.8592 | 1.7634 | 3.7947 | 2.6958 | 3.6718 | 3.7738 | 2.7283 | 3.3428 | 4.2387 | H10 | 2.2670 | 2.2642 | 3.0875 | 1.0878 | 2.7018 | 3.3057 | 2.4939 | 2.9626 | 3.8592 | | 3.7947 | 1.7634 | 2.6958 | 3.6718 | 2.7283 | 3.7738 | 4.2387 | 3.3428 | H11 | 2.2069 | 2.1982 | 1.0879 | 2.7409 | 3.4526 | 2.6980 | 3.0600 | 2.4749 | 1.7634 | 3.7947 | | 2.9254 | 3.7889 | 2.5179 | 4.3227 | 3.6787 | 2.9654 | 3.7340 | H12 | 2.2069 | 2.1982 | 2.7409 | 1.0879 | 3.4526 | 2.6980 | 3.0600 | 2.4749 | 3.7947 | 1.7634 | 2.9254 | | 3.7889 | 2.5179 | 3.6787 | 4.3227 | 3.7340 | 2.9654 | H13 | 2.1822 | 2.8823 | 2.7381 | 2.7381 | 1.0887 | 3.4717 | 2.5707 | 3.9584 | 2.6958 | 2.6958 | 3.7889 | 3.7889 | | 4.3223 | 1.7580 | 1.7580 | 3.7724 | 3.7724 | H14 | 2.1726 | 3.5967 | 2.8305 | 2.8305 | 3.4692 | 1.0887 | 4.5344 | 4.0386 | 3.6718 | 3.6718 | 2.5179 | 2.5179 | 4.3223 | | 3.7687 | 3.7687 | 1.7596 | 1.7596 | H15 | 2.1724 | 3.8045 | 3.5003 | 2.9054 | 1.0895 | 2.7654 | 3.8719 | 4.8450 | 3.7738 | 2.7283 | 4.3227 | 3.6787 | 1.7580 | 3.7687 | | 1.7614 | 3.1182 | 2.5735 | H16 | 2.1724 | 3.8045 | 2.9054 | 3.5003 | 1.0895 | 2.7654 | 3.8719 | 4.8450 | 2.7283 | 3.7738 | 3.6787 | 4.3227 | 1.7580 | 3.7687 | 1.7614 | | 2.5735 | 3.1182 | H17 | 2.1710 | 4.1554 | 2.9628 | 3.5476 | 2.7673 | 1.0899 | 4.8309 | 4.8891 | 3.3428 | 4.2387 | 2.9654 | 3.7340 | 3.7724 | 1.7596 | 3.1182 | 2.5735 | | 1.7601 | H18 | 2.1710 | 4.1554 | 3.5476 | 2.9628 | 2.7673 | 1.0899 | 4.8309 | 4.8891 | 4.2387 | 3.3428 | 3.7340 | 2.9654 | 3.7724 | 1.7596 | 2.5735 | 3.1182 | 1.7601 | |
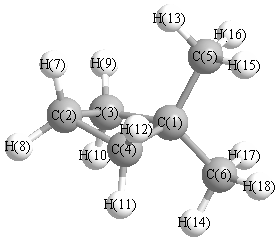 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C2 |
89.407 |
|
C1 |
C3 |
H9 |
116.949 |
| C1 |
C3 |
H11 |
111.944 |
|
C1 |
C4 |
C2 |
89.407 |
| C1 |
C4 |
H10 |
116.949 |
|
C1 |
C4 |
H12 |
111.944 |
| C1 |
C5 |
H13 |
111.733 |
|
C1 |
C5 |
H15 |
110.900 |
| C1 |
C5 |
H16 |
110.900 |
|
C1 |
C6 |
H14 |
111.298 |
| C1 |
C6 |
H17 |
111.095 |
|
C1 |
C6 |
H18 |
111.095 |
| C2 |
C3 |
H9 |
117.437 |
|
C2 |
C3 |
H11 |
111.913 |
| C2 |
C4 |
H10 |
117.437 |
|
C2 |
C4 |
H12 |
111.913 |
| C3 |
C1 |
C4 |
88.072 |
|
C3 |
C1 |
C5 |
111.927 |
| C3 |
C1 |
C6 |
116.359 |
|
C3 |
C2 |
C4 |
88.748 |
| C3 |
C2 |
H7 |
112.560 |
|
C3 |
C2 |
H8 |
116.966 |
| C4 |
C1 |
C5 |
111.927 |
|
C4 |
C1 |
C6 |
116.359 |
| C4 |
C2 |
H7 |
112.560 |
|
C4 |
C2 |
H8 |
116.966 |
| C5 |
C1 |
C6 |
110.563 |
|
H7 |
C2 |
H8 |
108.162 |
| H9 |
C3 |
H11 |
108.290 |
|
H10 |
C4 |
H12 |
108.290 |
| H13 |
C5 |
H15 |
107.629 |
|
H13 |
C5 |
H16 |
107.629 |
| H14 |
C6 |
H17 |
107.744 |
|
H14 |
C6 |
H18 |
107.744 |
| H15 |
C5 |
H16 |
107.879 |
|
H17 |
C6 |
H18 |
107.697 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
