Jump to
S1C2
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -133.698472 |
| Energy at 298.15K | -133.693577 |
| HF Energy | -133.072779 |
| Nuclear repulsion energy | 71.351285 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3730 |
3730 |
1.18 |
|
|
|
| 2 |
A' |
3304 |
3304 |
19.15 |
|
|
|
| 3 |
A' |
3213 |
3213 |
62.44 |
|
|
|
| 4 |
A' |
3184 |
3184 |
20.50 |
|
|
|
| 5 |
A' |
1917 |
1917 |
92.98 |
|
|
|
| 6 |
A' |
1609 |
1609 |
8.54 |
|
|
|
| 7 |
A' |
1579 |
1579 |
36.55 |
|
|
|
| 8 |
A' |
1526 |
1526 |
7.07 |
|
|
|
| 9 |
A' |
1381 |
1381 |
29.45 |
|
|
|
| 10 |
A' |
1152 |
1152 |
26.49 |
|
|
|
| 11 |
A' |
983 |
983 |
5.72 |
|
|
|
| 12 |
A' |
520 |
520 |
20.48 |
|
|
|
| 13 |
A" |
3242 |
3242 |
29.58 |
|
|
|
| 14 |
A" |
1614 |
1614 |
7.15 |
|
|
|
| 15 |
A" |
1251 |
1251 |
11.36 |
|
|
|
| 16 |
A" |
1178 |
1178 |
20.31 |
|
|
|
| 17 |
A" |
741 |
741 |
58.41 |
|
|
|
| 18 |
A" |
199 |
199 |
3.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16160.8 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16160.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.020 |
-0.654 |
0.000 |
| C2 |
0.000 |
0.448 |
0.000 |
| N3 |
1.230 |
0.212 |
0.000 |
| H4 |
-0.526 |
-1.619 |
0.000 |
| H5 |
-1.662 |
-0.577 |
0.877 |
| H6 |
-1.662 |
-0.577 |
-0.877 |
| H7 |
-0.403 |
1.461 |
0.000 |
| H8 |
1.765 |
1.062 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5019 | 2.4115 | 1.0843 | 1.0894 | 1.0894 | 2.2035 | 3.2715 |
C2 | 1.5019 | | 1.2525 | 2.1326 | 2.1409 | 2.1409 | 1.0901 | 1.8686 | N3 | 2.4115 | 1.2525 | | 2.5368 | 3.1239 | 3.1239 | 2.0554 | 1.0040 | H4 | 1.0843 | 2.1326 | 2.5368 | | 1.7738 | 1.7738 | 3.0823 | 3.5260 | H5 | 1.0894 | 2.1409 | 3.1239 | 1.7738 | | 1.7536 | 2.5516 | 3.8990 | H6 | 1.0894 | 2.1409 | 3.1239 | 1.7738 | 1.7536 | | 2.5516 | 3.8990 | H7 | 2.2035 | 1.0901 | 2.0554 | 3.0823 | 2.5516 | 2.5516 | | 2.2038 | H8 | 3.2715 | 1.8686 | 1.0040 | 3.5260 | 3.8990 | 3.8990 | 2.2038 | |
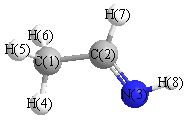 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
121.954 |
|
C1 |
C2 |
H7 |
115.531 |
| C2 |
C1 |
H4 |
110.044 |
|
C2 |
C1 |
H5 |
110.402 |
| C2 |
C1 |
H6 |
110.402 |
|
C2 |
N3 |
H8 |
111.340 |
| N3 |
C2 |
H7 |
122.514 |
|
H4 |
C1 |
H5 |
109.372 |
| H4 |
C1 |
H6 |
109.372 |
|
H5 |
C1 |
H6 |
107.194 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -133.697336 |
| Energy at 298.15K | -133.703070 |
| HF Energy | -133.071718 |
| Nuclear repulsion energy | 71.320554 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3697 |
3697 |
3.22 |
|
|
|
| 2 |
A' |
3280 |
3280 |
68.29 |
|
|
|
| 3 |
A' |
3261 |
3261 |
2.31 |
|
|
|
| 4 |
A' |
3185 |
3185 |
14.37 |
|
|
|
| 5 |
A' |
1919 |
1919 |
99.58 |
|
|
|
| 6 |
A' |
1616 |
1616 |
22.56 |
|
|
|
| 7 |
A' |
1554 |
1554 |
17.98 |
|
|
|
| 8 |
A' |
1528 |
1528 |
5.19 |
|
|
|
| 9 |
A' |
1394 |
1394 |
65.63 |
|
|
|
| 10 |
A' |
1151 |
1151 |
23.71 |
|
|
|
| 11 |
A' |
972 |
972 |
1.21 |
|
|
|
| 12 |
A' |
526 |
526 |
9.97 |
|
|
|
| 13 |
A" |
3250 |
3250 |
22.63 |
|
|
|
| 14 |
A" |
1608 |
1608 |
7.09 |
|
|
|
| 15 |
A" |
1259 |
1259 |
48.72 |
|
|
|
| 16 |
A" |
1183 |
1183 |
17.08 |
|
|
|
| 17 |
A" |
754 |
754 |
18.16 |
|
|
|
| 18 |
A" |
182 |
182 |
0.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16158.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16158.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.005 |
-0.636 |
0.000 |
| C2 |
0.000 |
0.484 |
0.000 |
| N3 |
1.246 |
0.391 |
0.000 |
| H4 |
-0.523 |
-1.610 |
0.000 |
| H5 |
-1.647 |
-0.564 |
0.876 |
| H6 |
-1.647 |
-0.564 |
-0.876 |
| H7 |
-0.406 |
1.489 |
0.000 |
| H8 |
1.530 |
-0.574 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5046 | 2.4747 | 1.0868 | 1.0887 | 1.0887 | 2.2079 | 2.5357 |
C2 | 1.5046 | | 1.2497 | 2.1580 | 2.1400 | 2.1400 | 1.0841 | 1.8600 | N3 | 2.4747 | 1.2497 | | 2.6714 | 3.1709 | 3.1709 | 1.9835 | 1.0064 | H4 | 1.0868 | 2.1580 | 2.6714 | | 1.7679 | 1.7679 | 3.1013 | 2.2994 | H5 | 1.0887 | 2.1400 | 3.1709 | 1.7679 | | 1.7521 | 2.5544 | 3.2959 | H6 | 1.0887 | 2.1400 | 3.1709 | 1.7679 | 1.7521 | | 2.5544 | 3.2959 | H7 | 2.2079 | 1.0841 | 1.9835 | 3.1013 | 2.5544 | 2.5544 | | 2.8291 | H8 | 2.5357 | 1.8600 | 1.0064 | 2.2994 | 3.2959 | 3.2959 | 2.8291 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
127.680 |
|
C1 |
C2 |
H7 |
116.104 |
| C2 |
C1 |
H4 |
111.754 |
|
C2 |
C1 |
H5 |
110.188 |
| C2 |
C1 |
H6 |
110.188 |
|
C2 |
N3 |
H8 |
110.601 |
| N3 |
C2 |
H7 |
116.215 |
|
H4 |
C1 |
H5 |
108.710 |
| H4 |
C1 |
H6 |
108.710 |
|
H5 |
C1 |
H6 |
107.161 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
