Jump to
S1C2
Vibrational Frequencies calculated at CBS-Q
Geometric Data calculated at CBS-Q
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -193.066692 |
| Energy at 298.15K | -193.071999 |
| HF Energy | -192.189727 |
| Nuclear repulsion energy | 149.030128 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3418 |
3418 |
0.75 |
|
|
|
| 2 |
A1 |
3407 |
3407 |
24.57 |
|
|
|
| 3 |
A1 |
3380 |
3380 |
0.69 |
|
|
|
| 4 |
A1 |
1739 |
1739 |
0.13 |
|
|
|
| 5 |
A1 |
1560 |
1560 |
0.20 |
|
|
|
| 6 |
A1 |
1223 |
1223 |
0.49 |
|
|
|
| 7 |
A1 |
1134 |
1134 |
6.44 |
|
|
|
| 8 |
A1 |
1084 |
1084 |
2.38 |
|
|
|
| 9 |
A1 |
892 |
892 |
0.35 |
|
|
|
| 10 |
A2 |
1033 |
1033 |
0.00 |
|
|
|
| 11 |
A2 |
809 |
809 |
0.00 |
|
|
|
| 12 |
A2 |
563 |
563 |
0.00 |
|
|
|
| 13 |
B1 |
904 |
904 |
14.17 |
|
|
|
| 14 |
B1 |
795 |
795 |
17.56 |
|
|
|
| 15 |
B1 |
707 |
707 |
74.15 |
|
|
|
| 16 |
B1 |
520 |
520 |
7.66 |
|
|
|
| 17 |
B2 |
7892 |
7892 |
206595.40 |
|
|
|
| 18 |
B2 |
3389 |
3389 |
105.32 |
|
|
|
| 19 |
B2 |
3380 |
3380 |
20.10 |
|
|
|
| 20 |
B2 |
1504 |
1504 |
43.59 |
|
|
|
| 21 |
B2 |
1403 |
1403 |
0.04 |
|
|
|
| 22 |
B2 |
1248 |
1248 |
14.09 |
|
|
|
| 23 |
B2 |
1022 |
1022 |
1.43 |
|
|
|
| 24 |
B2 |
913 |
913 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21957.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 21957.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
1.238 |
| C2 |
0.000 |
1.111 |
0.386 |
| C3 |
0.000 |
0.666 |
-1.011 |
| C4 |
0.000 |
-0.666 |
-1.011 |
| C5 |
0.000 |
-1.111 |
0.386 |
| H6 |
0.000 |
0.000 |
2.313 |
| H7 |
0.000 |
2.129 |
0.733 |
| H8 |
0.000 |
1.333 |
-1.855 |
| H9 |
0.000 |
-1.333 |
-1.855 |
| H10 |
0.000 |
-2.129 |
0.733 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.3999 | 2.3459 | 2.3459 | 1.3999 | 1.0750 | 2.1881 | 3.3683 | 3.3683 | 2.1881 |
C2 | 1.3999 | | 1.4663 | 2.2605 | 2.2215 | 2.2242 | 1.0759 | 2.2521 | 3.3161 | 3.2584 | C3 | 2.3459 | 1.4663 | | 1.3324 | 2.2605 | 3.3904 | 2.2766 | 1.0757 | 2.1704 | 3.2949 | C4 | 2.3459 | 2.2605 | 1.3324 | | 1.4663 | 3.3904 | 3.2949 | 2.1704 | 1.0757 | 2.2766 | C5 | 1.3999 | 2.2215 | 2.2605 | 1.4663 | | 2.2242 | 3.2584 | 3.3161 | 2.2521 | 1.0759 | H6 | 1.0750 | 2.2242 | 3.3904 | 3.3904 | 2.2242 | | 2.6512 | 4.3762 | 4.3762 | 2.6512 | H7 | 2.1881 | 1.0759 | 2.2766 | 3.2949 | 3.2584 | 2.6512 | | 2.7077 | 4.3230 | 4.2582 | H8 | 3.3683 | 2.2521 | 1.0757 | 2.1704 | 3.3161 | 4.3762 | 2.7077 | | 2.6669 | 4.3230 | H9 | 3.3683 | 3.3161 | 2.1704 | 1.0757 | 2.2521 | 4.3762 | 4.3230 | 2.6669 | | 2.7077 | H10 | 2.1881 | 3.2584 | 3.2949 | 2.2766 | 1.0759 | 2.6512 | 4.2582 | 4.3230 | 2.7077 | |
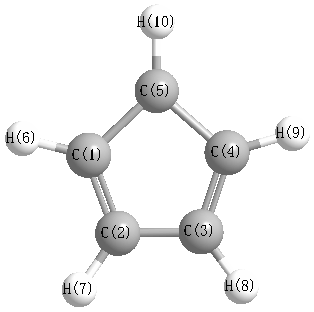 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
109.842 |
|
C1 |
C2 |
H7 |
123.684 |
| C1 |
C5 |
C4 |
109.842 |
|
C1 |
C5 |
H10 |
123.684 |
| C2 |
C1 |
C5 |
105.018 |
|
C2 |
C1 |
H6 |
127.491 |
| C2 |
C3 |
C4 |
107.649 |
|
C2 |
C3 |
H8 |
124.014 |
| C3 |
C2 |
H7 |
126.474 |
|
C3 |
C4 |
C5 |
107.649 |
| C3 |
C4 |
H9 |
128.336 |
|
C4 |
C3 |
H8 |
128.336 |
| C4 |
C5 |
H10 |
126.474 |
|
C5 |
C1 |
H6 |
127.491 |
| C5 |
C4 |
H9 |
124.014 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
