Jump to
S1C2
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -235.320221 |
| Energy at 298.15K | -235.312323 |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
Geometric Data calculated at CBS-Q
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -235.320221 |
| Energy at 298.15K | -235.334509 |
| HF Energy | -234.167706 |
| Nuclear repulsion energy | 249.623156 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3256 |
3256 |
175.73 |
|
|
|
| 2 |
A' |
3244 |
3244 |
32.76 |
|
|
|
| 3 |
A' |
3238 |
3238 |
36.52 |
|
|
|
| 4 |
A' |
3217 |
3217 |
12.31 |
|
|
|
| 5 |
A' |
3198 |
3198 |
9.34 |
|
|
|
| 6 |
A' |
3196 |
3196 |
14.01 |
|
|
|
| 7 |
A' |
3176 |
3176 |
40.67 |
|
|
|
| 8 |
A' |
3174 |
3174 |
30.22 |
|
|
|
| 9 |
A' |
1644 |
1644 |
3.24 |
|
|
|
| 10 |
A' |
1632 |
1632 |
2.17 |
|
|
|
| 11 |
A' |
1630 |
1630 |
4.45 |
|
|
|
| 12 |
A' |
1553 |
1553 |
5.08 |
|
|
|
| 13 |
A' |
1552 |
1552 |
0.91 |
|
|
|
| 14 |
A' |
1508 |
1508 |
2.69 |
|
|
|
| 15 |
A' |
1495 |
1495 |
7.17 |
|
|
|
| 16 |
A' |
1409 |
1409 |
1.53 |
|
|
|
| 17 |
A' |
1358 |
1358 |
0.48 |
|
|
|
| 18 |
A' |
1300 |
1300 |
0.13 |
|
|
|
| 19 |
A' |
1203 |
1203 |
0.39 |
|
|
|
| 20 |
A' |
1120 |
1120 |
0.32 |
|
|
|
| 21 |
A' |
1035 |
1035 |
0.67 |
|
|
|
| 22 |
A' |
1019 |
1019 |
0.41 |
|
|
|
| 23 |
A' |
927 |
927 |
0.69 |
|
|
|
| 24 |
A' |
851 |
851 |
0.81 |
|
|
|
| 25 |
A' |
695 |
695 |
1.15 |
|
|
|
| 26 |
A' |
461 |
461 |
0.03 |
|
|
|
| 27 |
A' |
345 |
345 |
0.07 |
|
|
|
| 28 |
A' |
103 |
103 |
0.01 |
|
|
|
| 29 |
A" |
3246 |
3246 |
45.84 |
|
|
|
| 30 |
A" |
3242 |
3242 |
24.93 |
|
|
|
| 31 |
A" |
3240 |
3240 |
38.27 |
|
|
|
| 32 |
A" |
3196 |
3196 |
88.67 |
|
|
|
| 33 |
A" |
1635 |
1635 |
1.67 |
|
|
|
| 34 |
A" |
1632 |
1632 |
0.20 |
|
|
|
| 35 |
A" |
1615 |
1615 |
1.99 |
|
|
|
| 36 |
A" |
1425 |
1425 |
1.05 |
|
|
|
| 37 |
A" |
1393 |
1393 |
0.01 |
|
|
|
| 38 |
A" |
1358 |
1358 |
0.11 |
|
|
|
| 39 |
A" |
1245 |
1245 |
0.44 |
|
|
|
| 40 |
A" |
1228 |
1228 |
0.00 |
|
|
|
| 41 |
A" |
1077 |
1077 |
2.07 |
|
|
|
| 42 |
A" |
974 |
974 |
0.06 |
|
|
|
| 43 |
A" |
970 |
970 |
0.30 |
|
|
|
| 44 |
A" |
870 |
870 |
0.10 |
|
|
|
| 45 |
A" |
399 |
399 |
0.00 |
|
|
|
| 46 |
A" |
277 |
277 |
0.06 |
|
|
|
| 47 |
A" |
264 |
264 |
0.00 |
|
|
|
| 48 |
A" |
247 |
247 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 39035.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 39035.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.250 |
0.081 |
1.080 |
| C2 |
0.250 |
0.081 |
-1.080 |
| C3 |
-0.657 |
0.722 |
0.000 |
| C4 |
0.841 |
-0.863 |
0.000 |
| C5 |
-0.840 |
2.233 |
0.000 |
| C6 |
0.250 |
-2.270 |
0.000 |
| H7 |
0.990 |
0.786 |
1.455 |
| H8 |
-0.240 |
-0.385 |
1.933 |
| H9 |
0.990 |
0.786 |
-1.455 |
| H10 |
-0.240 |
-0.385 |
-1.933 |
| H11 |
-1.634 |
0.242 |
0.000 |
| H12 |
1.926 |
-0.931 |
0.000 |
| H13 |
0.122 |
2.744 |
0.000 |
| H14 |
-0.839 |
-2.247 |
0.000 |
| H15 |
-1.391 |
2.563 |
-0.880 |
| H16 |
-1.391 |
2.563 |
0.880 |
| H17 |
0.568 |
-2.826 |
-0.880 |
| H18 |
0.568 |
-2.826 |
0.880 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 2.1607 | 1.5494 | 1.5513 | 2.6430 | 2.5871 | 1.0885 | 1.0882 | 2.7333 | 3.0882 | 2.1774 | 2.2364 | 2.8768 | 2.7884 | 3.5630 | 2.9820 | 3.5211 | 2.9315 |
C2 | 2.1607 | | 1.5494 | 1.5513 | 2.6430 | 2.5871 | 2.7333 | 3.0882 | 1.0885 | 1.0882 | 2.1774 | 2.2364 | 2.8768 | 2.7884 | 2.9820 | 3.5630 | 2.9315 | 3.5211 | C3 | 1.5494 | 1.5494 | | 2.1804 | 1.5217 | 3.1263 | 2.1984 | 2.2664 | 2.1984 | 2.2664 | 1.0886 | 3.0669 | 2.1668 | 2.9752 | 2.1682 | 2.1682 | 3.8559 | 3.8559 | C4 | 1.5513 | 1.5513 | 2.1804 | | 3.5223 | 1.5260 | 2.2036 | 2.2653 | 2.2036 | 2.2653 | 2.7097 | 1.0877 | 3.6778 | 2.1770 | 4.1819 | 4.1819 | 2.1691 | 2.1691 | C5 | 2.6430 | 2.6430 | 1.5217 | 3.5223 | | 4.6326 | 2.7494 | 3.3092 | 2.7494 | 3.3092 | 2.1434 | 4.2027 | 1.0894 | 4.4803 | 1.0894 | 1.0894 | 5.3248 | 5.3248 | C6 | 2.5871 | 2.5871 | 3.1263 | 1.5260 | 4.6326 | | 3.4641 | 2.7436 | 3.4641 | 2.7436 | 3.1395 | 2.1451 | 5.0155 | 1.0894 | 5.1789 | 5.1789 | 1.0891 | 1.0891 | H7 | 1.0885 | 2.7333 | 2.1984 | 2.2036 | 2.7494 | 3.4641 | | 1.7644 | 2.9092 | 3.7894 | 3.0491 | 2.4372 | 2.5894 | 3.8292 | 3.7786 | 3.0263 | 4.3217 | 3.6817 | H8 | 1.0882 | 3.0882 | 2.2664 | 2.2653 | 3.3092 | 2.7436 | 1.7644 | | 3.7894 | 3.8659 | 2.4642 | 2.9540 | 3.6960 | 2.7501 | 4.2342 | 3.3354 | 3.8114 | 2.7784 | H9 | 2.7333 | 1.0885 | 2.1984 | 2.2036 | 2.7494 | 3.4641 | 2.9092 | 3.7894 | | 1.7644 | 3.0491 | 2.4372 | 2.5894 | 3.8292 | 3.0263 | 3.7786 | 3.6817 | 4.3217 | H10 | 3.0882 | 1.0882 | 2.2664 | 2.2653 | 3.3092 | 2.7436 | 3.7894 | 3.8659 | 1.7644 | | 2.4642 | 2.9540 | 3.6960 | 2.7501 | 3.3354 | 4.2342 | 2.7784 | 3.8114 | H11 | 2.1774 | 2.1774 | 1.0886 | 2.7097 | 2.1434 | 3.1395 | 3.0491 | 2.4642 | 3.0491 | 2.4642 | | 3.7482 | 3.0568 | 2.6130 | 2.4940 | 2.4940 | 3.8778 | 3.8778 | H12 | 2.2364 | 2.2364 | 3.0669 | 1.0877 | 4.2027 | 2.1451 | 2.4372 | 2.9540 | 2.4372 | 2.9540 | 3.7482 | | 4.0943 | 3.0627 | 4.8975 | 4.8975 | 2.4921 | 2.4921 | H13 | 2.8768 | 2.8768 | 2.1668 | 3.6778 | 1.0894 | 5.0155 | 2.5894 | 3.6960 | 2.5894 | 3.6960 | 3.0568 | 4.0943 | | 5.0833 | 1.7597 | 1.7597 | 5.6572 | 5.6572 | H14 | 2.7884 | 2.7884 | 2.9752 | 2.1770 | 4.4803 | 1.0894 | 3.8292 | 2.7501 | 3.8292 | 2.7501 | 2.6130 | 3.0627 | 5.0833 | | 4.9211 | 4.9211 | 1.7582 | 1.7582 | H15 | 3.5630 | 2.9820 | 2.1682 | 4.1819 | 1.0894 | 5.1789 | 3.7786 | 4.2342 | 3.0263 | 3.3354 | 2.4940 | 4.8975 | 1.7597 | 4.9211 | | 1.7598 | 5.7342 | 5.9983 | H16 | 2.9820 | 3.5630 | 2.1682 | 4.1819 | 1.0894 | 5.1789 | 3.0263 | 3.3354 | 3.7786 | 4.2342 | 2.4940 | 4.8975 | 1.7597 | 4.9211 | 1.7598 | | 5.9983 | 5.7342 | H17 | 3.5211 | 2.9315 | 3.8559 | 2.1691 | 5.3248 | 1.0891 | 4.3217 | 3.8114 | 3.6817 | 2.7784 | 3.8778 | 2.4921 | 5.6572 | 1.7582 | 5.7342 | 5.9983 | | 1.7609 | H18 | 2.9315 | 3.5211 | 3.8559 | 2.1691 | 5.3248 | 1.0891 | 3.6817 | 2.7784 | 4.3217 | 3.8114 | 3.8778 | 2.4921 | 5.6572 | 1.7582 | 5.9983 | 5.7342 | 1.7609 | |
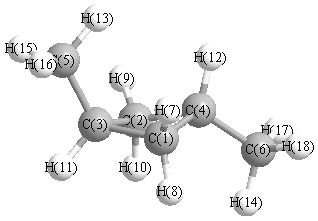 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C2 |
88.418 |
|
C1 |
C3 |
C5 |
118.766 |
| C1 |
C3 |
H11 |
110.037 |
|
C1 |
C4 |
C2 |
88.278 |
| C1 |
C4 |
C6 |
114.427 |
|
C1 |
C4 |
H12 |
114.726 |
| C2 |
C3 |
C5 |
118.766 |
|
C2 |
C3 |
H11 |
110.037 |
| C2 |
C4 |
C6 |
114.427 |
|
C2 |
C4 |
H12 |
114.726 |
| C3 |
C1 |
C4 |
89.370 |
|
C3 |
C1 |
H7 |
111.705 |
| C3 |
C1 |
H8 |
117.399 |
|
C3 |
C2 |
C4 |
89.370 |
| C3 |
C2 |
H9 |
111.705 |
|
C3 |
C2 |
H10 |
117.399 |
| C3 |
C5 |
H13 |
111.082 |
|
C3 |
C5 |
H15 |
111.193 |
| C3 |
C5 |
H16 |
111.193 |
|
C4 |
C1 |
H7 |
111.986 |
| C4 |
C1 |
H8 |
117.154 |
|
C4 |
C2 |
H9 |
111.986 |
| C4 |
C2 |
H10 |
117.154 |
|
C4 |
C6 |
H14 |
111.598 |
| C4 |
C6 |
H17 |
110.981 |
|
C4 |
C6 |
H18 |
110.981 |
| C5 |
C3 |
H11 |
109.274 |
|
C6 |
C4 |
H12 |
109.164 |
| H7 |
C1 |
H8 |
108.308 |
|
H9 |
C2 |
H10 |
108.308 |
| H13 |
C5 |
H15 |
107.731 |
|
H13 |
C5 |
H16 |
107.731 |
| H14 |
C6 |
H17 |
107.616 |
|
H14 |
C6 |
H18 |
107.616 |
| H15 |
C5 |
H16 |
107.742 |
|
H17 |
C6 |
H18 |
107.876 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
