Jump to
S1C2
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -377.127554 |
| Energy at 298.15K | -377.121699 |
| HF Energy | -375.774039 |
| Nuclear repulsion energy | 193.032284 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3328 |
3328 |
63.79 |
23.63 |
0.20 |
0.33 |
| 2 |
A |
3311 |
3311 |
13.74 |
106.44 |
0.48 |
0.65 |
| 3 |
A |
3256 |
3256 |
22.05 |
93.56 |
0.10 |
0.19 |
| 4 |
A |
1647 |
1647 |
4.80 |
8.18 |
0.75 |
0.86 |
| 5 |
A |
1620 |
1620 |
17.95 |
2.29 |
0.40 |
0.58 |
| 6 |
A |
1560 |
1560 |
37.83 |
4.20 |
0.75 |
0.85 |
| 7 |
A |
1481 |
1481 |
37.26 |
2.63 |
0.75 |
0.86 |
| 8 |
A |
1384 |
1384 |
18.65 |
8.79 |
0.73 |
0.84 |
| 9 |
A |
1284 |
1284 |
134.09 |
4.76 |
0.74 |
0.85 |
| 10 |
A |
1250 |
1250 |
134.40 |
4.75 |
0.75 |
0.86 |
| 11 |
A |
1233 |
1233 |
104.40 |
1.62 |
0.21 |
0.35 |
| 12 |
A |
1199 |
1199 |
85.03 |
4.83 |
0.49 |
0.66 |
| 13 |
A |
993 |
993 |
40.59 |
5.42 |
0.33 |
0.49 |
| 14 |
A |
632 |
632 |
6.29 |
1.54 |
0.42 |
0.59 |
| 15 |
A |
516 |
516 |
24.87 |
0.60 |
0.73 |
0.85 |
| 16 |
A |
464 |
464 |
7.05 |
1.50 |
0.49 |
0.65 |
| 17 |
A |
262 |
262 |
10.51 |
0.04 |
0.54 |
0.70 |
| 18 |
A |
128 |
128 |
10.24 |
0.01 |
0.72 |
0.84 |
Unscaled Zero Point Vibrational Energy (zpe) 12773.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12773.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.775 |
-0.587 |
-0.279 |
| C2 |
0.468 |
0.020 |
0.325 |
| F3 |
-1.856 |
0.118 |
0.151 |
| F4 |
1.513 |
-0.750 |
-0.008 |
| F5 |
0.678 |
1.238 |
-0.183 |
| H6 |
-0.718 |
-0.533 |
-1.360 |
| H7 |
-0.875 |
-1.618 |
0.039 |
| H8 |
0.418 |
0.102 |
1.402 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5097 | 1.3600 | 2.3103 | 2.3346 | 1.0839 | 1.0837 | 2.1736 |
C2 | 1.5097 | | 2.3332 | 1.3406 | 1.3356 | 2.1339 | 2.1379 | 1.0816 | F3 | 1.3600 | 2.3332 | | 3.4834 | 2.7914 | 2.0001 | 1.9970 | 2.5964 | F4 | 2.3103 | 1.3406 | 3.4834 | | 2.1634 | 2.6184 | 2.5415 | 1.9785 | F5 | 2.3346 | 1.3356 | 2.7914 | 2.1634 | | 2.5441 | 3.2584 | 1.9675 | H6 | 1.0839 | 2.1339 | 2.0001 | 2.6184 | 2.5441 | | 1.7770 | 3.0538 | H7 | 1.0837 | 2.1379 | 1.9970 | 2.5415 | 3.2584 | 1.7770 | | 2.5475 | H8 | 2.1736 | 1.0816 | 2.5964 | 1.9785 | 1.9675 | 3.0538 | 2.5475 | |
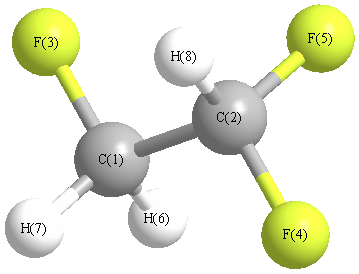 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.155 |
|
C1 |
C2 |
F5 |
110.123 |
| C1 |
C2 |
H8 |
112.989 |
|
C2 |
C1 |
F3 |
108.682 |
| C2 |
C1 |
H6 |
109.629 |
|
C2 |
C1 |
H7 |
109.958 |
| F3 |
C1 |
H6 |
109.330 |
|
F3 |
C1 |
H7 |
109.086 |
| F4 |
C2 |
F5 |
107.871 |
|
F4 |
C2 |
H8 |
109.067 |
| F5 |
C2 |
H8 |
108.509 |
|
H6 |
C1 |
H7 |
110.130 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -377.125291 |
| Energy at 298.15K | -377.119443 |
| HF Energy | -375.771174 |
| Nuclear repulsion energy | 195.954237 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3289 |
3289 |
69.13 |
85.88 |
0.15 |
0.25 |
| 2 |
A' |
3245 |
3245 |
16.50 |
79.80 |
0.20 |
0.33 |
| 3 |
A' |
1643 |
1643 |
6.16 |
6.40 |
0.75 |
0.86 |
| 4 |
A' |
1594 |
1594 |
35.67 |
3.01 |
0.40 |
0.57 |
| 5 |
A' |
1560 |
1560 |
37.37 |
3.19 |
0.72 |
0.83 |
| 6 |
A' |
1303 |
1303 |
157.35 |
5.12 |
0.55 |
0.71 |
| 7 |
A' |
1210 |
1210 |
26.01 |
3.52 |
0.75 |
0.86 |
| 8 |
A' |
950 |
950 |
49.70 |
7.46 |
0.18 |
0.30 |
| 9 |
A' |
823 |
823 |
65.98 |
2.66 |
0.52 |
0.69 |
| 10 |
A' |
566 |
566 |
17.42 |
1.37 |
0.75 |
0.85 |
| 11 |
A' |
251 |
251 |
2.67 |
0.11 |
0.43 |
0.60 |
| 12 |
A" |
3305 |
3305 |
35.61 |
52.98 |
0.75 |
0.86 |
| 13 |
A" |
1559 |
1559 |
47.38 |
1.73 |
0.75 |
0.86 |
| 14 |
A" |
1404 |
1404 |
24.30 |
14.69 |
0.75 |
0.86 |
| 15 |
A" |
1276 |
1276 |
122.08 |
1.28 |
0.75 |
0.86 |
| 16 |
A" |
1064 |
1064 |
68.99 |
3.57 |
0.75 |
0.86 |
| 17 |
A" |
397 |
397 |
0.11 |
0.13 |
0.75 |
0.86 |
| 18 |
A" |
121 |
121 |
2.84 |
0.02 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12780.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12780.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.735 |
-0.830 |
0.000 |
| C2 |
0.351 |
0.629 |
0.000 |
| F3 |
-0.384 |
-1.597 |
0.000 |
| F4 |
-0.384 |
0.912 |
1.080 |
| F5 |
-0.384 |
0.912 |
-1.080 |
| H6 |
1.316 |
-1.055 |
-0.888 |
| H7 |
1.316 |
-1.055 |
0.888 |
| H8 |
1.223 |
1.272 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5085 | 1.3560 | 2.3351 | 2.3351 | 1.0848 | 1.0848 | 2.1581 |
C2 | 1.5085 | | 2.3436 | 1.3366 | 1.3366 | 2.1339 | 2.1339 | 1.0839 | F3 | 1.3560 | 2.3436 | | 2.7312 | 2.7312 | 1.9932 | 1.9932 | 3.2881 | F4 | 2.3351 | 1.3366 | 2.7312 | | 2.1597 | 3.2604 | 2.6069 | 1.9696 | F5 | 2.3351 | 1.3366 | 2.7312 | 2.1597 | | 2.6069 | 3.2604 | 1.9696 | H6 | 1.0848 | 2.1339 | 1.9932 | 3.2604 | 2.6069 | | 1.7753 | 2.4918 | H7 | 1.0848 | 2.1339 | 1.9932 | 2.6069 | 3.2604 | 1.7753 | | 2.4918 | H8 | 2.1581 | 1.0839 | 3.2881 | 1.9696 | 1.9696 | 2.4918 | 2.4918 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.172 |
|
C1 |
C2 |
F5 |
110.172 |
| C1 |
C2 |
H8 |
111.664 |
|
C2 |
C1 |
F3 |
109.684 |
| C2 |
C1 |
H6 |
109.663 |
|
C2 |
C1 |
H7 |
109.663 |
| F3 |
C1 |
H6 |
108.992 |
|
F3 |
C1 |
H7 |
108.992 |
| F4 |
C2 |
F5 |
107.784 |
|
F4 |
C2 |
H8 |
108.471 |
| F5 |
C2 |
H8 |
108.471 |
|
H6 |
C1 |
H7 |
109.830 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
