Jump to
S1C2
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -154.104750 |
| Energy at 298.15K | -154.099030 |
| HF Energy | -153.437900 |
| Nuclear repulsion energy | 75.265457 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
4110 |
4110 |
25.95 |
|
|
|
| 2 |
A' |
3302 |
3302 |
12.43 |
|
|
|
| 3 |
A' |
3167 |
3167 |
78.00 |
|
|
|
| 4 |
A' |
1665 |
1665 |
2.22 |
|
|
|
| 5 |
A' |
1617 |
1617 |
7.81 |
|
|
|
| 6 |
A' |
1561 |
1561 |
8.04 |
|
|
|
| 7 |
A' |
1341 |
1341 |
114.75 |
|
|
|
| 8 |
A' |
1170 |
1170 |
111.49 |
|
|
|
| 9 |
A' |
1068 |
1068 |
2.60 |
|
|
|
| 10 |
A' |
645 |
645 |
10.85 |
|
|
|
| 11 |
A' |
394 |
394 |
35.62 |
|
|
|
| 12 |
A" |
3407 |
3407 |
22.02 |
|
|
|
| 13 |
A" |
3200 |
3200 |
84.56 |
|
|
|
| 14 |
A" |
1411 |
1411 |
1.12 |
|
|
|
| 15 |
A" |
1273 |
1273 |
2.63 |
|
|
|
| 16 |
A" |
859 |
859 |
1.15 |
|
|
|
| 17 |
A" |
312 |
312 |
168.80 |
|
|
|
| 18 |
A" |
82 |
82 |
0.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15291.7 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 15291.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-1.105 |
-0.340 |
0.000 |
| C2 |
0.000 |
0.536 |
0.000 |
| C3 |
1.261 |
-0.270 |
0.000 |
| H4 |
-1.906 |
0.158 |
0.000 |
| H5 |
-0.031 |
1.180 |
0.880 |
| H6 |
-0.031 |
1.180 |
-0.880 |
| H7 |
1.622 |
-0.697 |
-0.922 |
| H8 |
1.622 |
-0.697 |
0.922 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| O1 | | 1.4108 | 2.3676 | 0.9428 | 2.0592 | 2.0592 | 2.9010 | 2.9010 |
C2 | 1.4108 | | 1.4967 | 1.9431 | 1.0909 | 1.0909 | 2.2361 | 2.2361 | C3 | 2.3676 | 1.4967 | | 3.1957 | 2.1317 | 2.1317 | 1.0779 | 1.0779 | H4 | 0.9428 | 1.9431 | 3.1957 | | 2.3099 | 2.3099 | 3.7451 | 3.7451 | H5 | 2.0592 | 1.0909 | 2.1317 | 2.3099 | | 1.7608 | 3.0820 | 2.5006 | H6 | 2.0592 | 1.0909 | 2.1317 | 2.3099 | 1.7608 | | 2.5006 | 3.0820 | H7 | 2.9010 | 2.2361 | 1.0779 | 3.7451 | 3.0820 | 2.5006 | | 1.8433 | H8 | 2.9010 | 2.2361 | 1.0779 | 3.7451 | 2.5006 | 3.0820 | 1.8433 | |
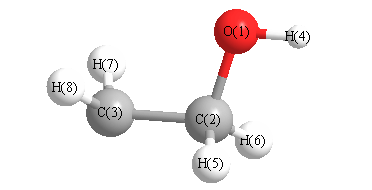 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
109.007 |
|
O1 |
C2 |
H5 |
110.147 |
| O1 |
C2 |
H6 |
110.147 |
|
C2 |
O1 |
H4 |
109.693 |
| C2 |
C3 |
H7 |
119.696 |
|
C2 |
C3 |
H8 |
119.696 |
| C3 |
C2 |
H5 |
109.953 |
|
C3 |
C2 |
H6 |
109.953 |
| H5 |
C2 |
H6 |
107.625 |
|
H7 |
C3 |
H8 |
117.524 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -154.106492 |
| Energy at 298.15K | -154.111571 |
| HF Energy | -153.438828 |
| Nuclear repulsion energy | 75.452832 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4124 |
4124 |
32.68 |
|
|
|
| 2 |
A |
3422 |
3422 |
13.43 |
|
|
|
| 3 |
A |
3310 |
3310 |
10.14 |
|
|
|
| 4 |
A |
3161 |
3161 |
87.98 |
|
|
|
| 5 |
A |
3109 |
3109 |
90.07 |
|
|
|
| 6 |
A |
1654 |
1654 |
3.64 |
|
|
|
| 7 |
A |
1613 |
1613 |
11.48 |
|
|
|
| 8 |
A |
1552 |
1552 |
2.58 |
|
|
|
| 9 |
A |
1392 |
1392 |
93.22 |
|
|
|
| 10 |
A |
1357 |
1357 |
28.60 |
|
|
|
| 11 |
A |
1217 |
1217 |
33.68 |
|
|
|
| 12 |
A |
1152 |
1152 |
44.87 |
|
|
|
| 13 |
A |
1051 |
1051 |
7.81 |
|
|
|
| 14 |
A |
942 |
942 |
15.55 |
|
|
|
| 15 |
A |
481 |
481 |
19.89 |
|
|
|
| 16 |
A |
422 |
422 |
34.06 |
|
|
|
| 17 |
A |
317 |
317 |
151.44 |
|
|
|
| 18 |
A |
178 |
178 |
14.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15226.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 15226.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-1.093 |
-0.381 |
-0.056 |
| C2 |
-0.023 |
0.524 |
0.037 |
| C3 |
1.251 |
-0.248 |
-0.022 |
| H4 |
-1.901 |
0.080 |
0.091 |
| H5 |
-0.097 |
1.090 |
0.972 |
| H6 |
-0.070 |
1.250 |
-0.776 |
| H7 |
2.181 |
0.257 |
-0.212 |
| H8 |
1.263 |
-1.279 |
0.283 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| O1 | | 1.4047 | 2.3481 | 0.9414 | 2.0524 | 2.0558 | 3.3398 | 2.5442 |
C2 | 1.4047 | | 1.4909 | 1.9303 | 1.0949 | 1.0913 | 2.2343 | 2.2280 | C3 | 2.3481 | 1.4909 | | 3.1708 | 2.1436 | 2.1352 | 1.0759 | 1.0747 | H4 | 0.9414 | 1.9303 | 3.1708 | | 2.2472 | 2.3393 | 4.0972 | 3.4486 | H5 | 2.0524 | 1.0949 | 2.1436 | 2.2472 | | 1.7551 | 2.6990 | 2.8168 | H6 | 2.0558 | 1.0913 | 2.1352 | 2.3393 | 1.7551 | | 2.5244 | 3.0486 | H7 | 3.3398 | 2.2343 | 1.0759 | 4.0972 | 2.6990 | 2.5244 | | 1.8568 | H8 | 2.5442 | 2.2280 | 1.0747 | 3.4486 | 2.8168 | 3.0486 | 1.8568 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
108.340 |
|
O1 |
C2 |
H5 |
109.768 |
| O1 |
C2 |
H6 |
110.270 |
|
C2 |
O1 |
H4 |
109.132 |
| C2 |
C3 |
H7 |
120.165 |
|
C2 |
C3 |
H8 |
119.675 |
| C3 |
C2 |
H5 |
111.060 |
|
C3 |
C2 |
H6 |
110.612 |
| H5 |
C2 |
H6 |
106.794 |
|
H7 |
C3 |
H8 |
119.406 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
