Jump to
S1C2
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -218.017889 |
| Energy at 298.15K | -218.011804 |
| HF Energy | -217.107995 |
| Nuclear repulsion energy | 128.608102 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3265 |
3265 |
45.67 |
|
|
|
| 2 |
A' |
3219 |
3219 |
56.61 |
|
|
|
| 3 |
A' |
3204 |
3204 |
20.57 |
|
|
|
| 4 |
A' |
3185 |
3185 |
28.65 |
|
|
|
| 5 |
A' |
1665 |
1665 |
1.28 |
|
|
|
| 6 |
A' |
1647 |
1647 |
5.62 |
|
|
|
| 7 |
A' |
1629 |
1629 |
0.19 |
|
|
|
| 8 |
A' |
1583 |
1583 |
18.81 |
|
|
|
| 9 |
A' |
1553 |
1553 |
3.43 |
|
|
|
| 10 |
A' |
1456 |
1456 |
3.16 |
|
|
|
| 11 |
A' |
1227 |
1227 |
6.06 |
|
|
|
| 12 |
A' |
1169 |
1169 |
123.47 |
|
|
|
| 13 |
A' |
1105 |
1105 |
9.48 |
|
|
|
| 14 |
A' |
963 |
963 |
7.70 |
|
|
|
| 15 |
A' |
479 |
479 |
7.32 |
|
|
|
| 16 |
A' |
289 |
289 |
4.39 |
|
|
|
| 17 |
A" |
3277 |
3277 |
123.50 |
|
|
|
| 18 |
A" |
3250 |
3250 |
13.66 |
|
|
|
| 19 |
A" |
3231 |
3231 |
1.32 |
|
|
|
| 20 |
A" |
1636 |
1636 |
5.90 |
|
|
|
| 21 |
A" |
1435 |
1435 |
0.02 |
|
|
|
| 22 |
A" |
1387 |
1387 |
0.98 |
|
|
|
| 23 |
A" |
1303 |
1303 |
2.80 |
|
|
|
| 24 |
A" |
962 |
962 |
1.88 |
|
|
|
| 25 |
A" |
815 |
815 |
0.76 |
|
|
|
| 26 |
A" |
248 |
248 |
0.00 |
|
|
|
| 27 |
A" |
142 |
142 |
4.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22662.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 22662.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.131 |
-0.790 |
0.000 |
| C2 |
0.000 |
0.721 |
0.000 |
| C3 |
-1.461 |
1.173 |
0.000 |
| F4 |
1.457 |
-1.141 |
0.000 |
| H5 |
-0.326 |
-1.229 |
0.883 |
| H6 |
-0.326 |
-1.229 |
-0.883 |
| H7 |
0.517 |
1.113 |
-0.874 |
| H8 |
0.517 |
1.113 |
0.874 |
| H9 |
-1.530 |
2.258 |
0.000 |
| H10 |
-1.991 |
0.810 |
-0.880 |
| H11 |
-1.991 |
0.810 |
0.880 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5166 | 2.5282 | 1.3707 | 1.0866 | 1.0866 | 2.1291 | 2.1291 | 3.4715 | 2.7994 | 2.7994 |
C2 | 1.5166 | | 1.5299 | 2.3635 | 2.1652 | 2.1652 | 1.0884 | 1.0884 | 2.1688 | 2.1782 | 2.1782 | C3 | 2.5282 | 1.5299 | | 3.7240 | 2.8003 | 2.8003 | 2.1636 | 2.1636 | 1.0870 | 1.0890 | 1.0890 | F4 | 1.3707 | 2.3635 | 3.7240 | | 1.9907 | 1.9907 | 2.5930 | 2.5930 | 4.5243 | 4.0572 | 4.0572 | H5 | 1.0866 | 2.1652 | 2.8003 | 1.9907 | | 1.7650 | 3.0464 | 2.4889 | 3.7936 | 3.1679 | 2.6326 | H6 | 1.0866 | 2.1652 | 2.8003 | 1.9907 | 1.7650 | | 2.4889 | 3.0464 | 3.7936 | 2.6326 | 3.1679 | H7 | 2.1291 | 1.0884 | 2.1636 | 2.5930 | 3.0464 | 2.4889 | | 1.7483 | 2.5029 | 2.5257 | 3.0749 | H8 | 2.1291 | 1.0884 | 2.1636 | 2.5930 | 2.4889 | 3.0464 | 1.7483 | | 2.5029 | 3.0749 | 2.5257 | H9 | 3.4715 | 2.1688 | 1.0870 | 4.5243 | 3.7936 | 3.7936 | 2.5029 | 2.5029 | | 1.7561 | 1.7561 | H10 | 2.7994 | 2.1782 | 1.0890 | 4.0572 | 3.1679 | 2.6326 | 2.5257 | 3.0749 | 1.7561 | | 1.7592 | H11 | 2.7994 | 2.1782 | 1.0890 | 4.0572 | 2.6326 | 3.1679 | 3.0749 | 2.5257 | 1.7561 | 1.7592 | |
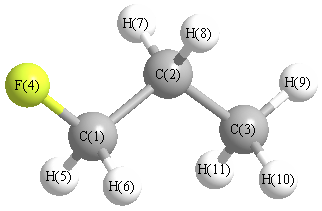 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.171 |
|
C1 |
C2 |
H7 |
108.511 |
| C1 |
C2 |
H8 |
108.511 |
|
C2 |
C1 |
F4 |
109.781 |
| C2 |
C1 |
H5 |
111.490 |
|
C2 |
C1 |
H6 |
111.490 |
| C2 |
C3 |
H9 |
110.803 |
|
C2 |
C3 |
H10 |
111.438 |
| C2 |
C3 |
H11 |
111.438 |
|
C3 |
C2 |
H7 |
110.310 |
| C3 |
C2 |
H8 |
110.310 |
|
F4 |
C1 |
H5 |
107.653 |
| F4 |
C1 |
H6 |
107.653 |
|
H5 |
C1 |
H6 |
108.622 |
| H7 |
C2 |
H8 |
106.858 |
|
H9 |
C3 |
H10 |
107.615 |
| H9 |
C3 |
H11 |
107.615 |
|
H10 |
C3 |
H11 |
107.748 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -218.017893 |
| Energy at 298.15K | -218.011893 |
| HF Energy | -217.108456 |
| Nuclear repulsion energy | 131.033892 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3280 |
3280 |
46.65 |
42.99 |
0.74 |
0.85 |
| 2 |
A |
3275 |
3275 |
79.55 |
61.64 |
0.65 |
0.78 |
| 3 |
A |
3256 |
3256 |
56.12 |
48.47 |
0.69 |
0.82 |
| 4 |
A |
3232 |
3232 |
36.27 |
92.13 |
0.34 |
0.51 |
| 5 |
A |
3220 |
3220 |
22.70 |
162.63 |
0.26 |
0.42 |
| 6 |
A |
3193 |
3193 |
17.02 |
136.65 |
0.09 |
0.16 |
| 7 |
A |
3188 |
3188 |
40.72 |
28.08 |
0.44 |
0.61 |
| 8 |
A |
1663 |
1663 |
0.22 |
5.11 |
0.74 |
0.85 |
| 9 |
A |
1644 |
1644 |
6.86 |
7.19 |
0.75 |
0.86 |
| 10 |
A |
1632 |
1632 |
5.67 |
19.68 |
0.75 |
0.86 |
| 11 |
A |
1616 |
1616 |
1.05 |
18.37 |
0.74 |
0.85 |
| 12 |
A |
1571 |
1571 |
19.98 |
2.13 |
0.53 |
0.69 |
| 13 |
A |
1555 |
1555 |
3.12 |
2.28 |
0.75 |
0.86 |
| 14 |
A |
1506 |
1506 |
1.24 |
1.04 |
0.74 |
0.85 |
| 15 |
A |
1417 |
1417 |
1.24 |
15.59 |
0.72 |
0.83 |
| 16 |
A |
1391 |
1391 |
1.83 |
6.69 |
0.75 |
0.86 |
| 17 |
A |
1285 |
1285 |
2.61 |
1.05 |
0.20 |
0.33 |
| 18 |
A |
1224 |
1224 |
31.11 |
2.13 |
0.69 |
0.82 |
| 19 |
A |
1173 |
1173 |
56.80 |
6.37 |
0.70 |
0.83 |
| 20 |
A |
1061 |
1061 |
39.05 |
5.56 |
0.74 |
0.85 |
| 21 |
A |
994 |
994 |
4.05 |
0.72 |
0.75 |
0.86 |
| 22 |
A |
927 |
927 |
3.93 |
12.40 |
0.18 |
0.30 |
| 23 |
A |
830 |
830 |
0.81 |
0.98 |
0.38 |
0.55 |
| 24 |
A |
514 |
514 |
5.18 |
0.43 |
0.69 |
0.82 |
| 25 |
A |
336 |
336 |
1.37 |
0.19 |
0.12 |
0.22 |
| 26 |
A |
242 |
242 |
2.06 |
0.07 |
0.45 |
0.62 |
| 27 |
A |
151 |
151 |
3.12 |
0.04 |
0.67 |
0.80 |
Unscaled Zero Point Vibrational Energy (zpe) 22688.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 22688.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.809 |
0.502 |
0.295 |
| C2 |
-0.587 |
0.663 |
-0.276 |
| C3 |
-1.526 |
-0.478 |
0.117 |
| F4 |
1.374 |
-0.658 |
-0.173 |
| H5 |
1.457 |
1.322 |
0.001 |
| H6 |
0.791 |
0.440 |
1.379 |
| H7 |
-0.512 |
0.727 |
-1.361 |
| H8 |
-0.984 |
1.617 |
0.070 |
| H9 |
-2.512 |
-0.339 |
-0.321 |
| H10 |
-1.137 |
-1.433 |
-0.224 |
| H11 |
-1.648 |
-0.532 |
1.198 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5167 | 2.5387 | 1.3724 | 1.0859 | 1.0866 | 2.1298 | 2.1239 | 3.4805 | 2.7921 | 2.8143 |
C2 | 1.5167 | | 1.5295 | 2.3670 | 2.1652 | 2.1654 | 1.0888 | 1.0901 | 2.1706 | 2.1672 | 2.1744 | C3 | 2.5387 | 1.5295 | | 2.9209 | 3.4862 | 2.7938 | 2.1597 | 2.1645 | 1.0876 | 1.0862 | 1.0891 | F4 | 1.3724 | 2.3670 | 2.9209 | | 1.9896 | 1.9891 | 2.6244 | 3.2863 | 3.9023 | 2.6283 | 3.3210 | H5 | 1.0859 | 2.1652 | 3.4862 | 1.9896 | | 1.7669 | 2.4668 | 2.4600 | 4.3143 | 3.7902 | 3.8092 | H6 | 1.0866 | 2.1654 | 2.7938 | 1.9891 | 1.7669 | | 3.0476 | 2.5002 | 3.7956 | 3.1294 | 2.6314 | H7 | 2.1298 | 1.0888 | 2.1597 | 2.6244 | 2.4668 | 3.0476 | | 1.7501 | 2.4930 | 2.5190 | 3.0695 | H8 | 2.1239 | 1.0901 | 2.1645 | 3.2863 | 2.4600 | 2.5002 | 1.7501 | | 2.5123 | 3.0677 | 2.5163 | H9 | 3.4805 | 2.1706 | 1.0876 | 3.9023 | 4.3143 | 3.7956 | 2.4930 | 2.5123 | | 1.7600 | 1.7587 | H10 | 2.7921 | 2.1672 | 1.0862 | 2.6283 | 3.7902 | 3.1294 | 2.5190 | 3.0677 | 1.7600 | | 1.7591 | H11 | 2.8143 | 2.1744 | 1.0891 | 3.3210 | 3.8092 | 2.6314 | 3.0695 | 2.5163 | 1.7587 | 1.7591 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.900 |
|
C1 |
C2 |
H7 |
108.537 |
| C1 |
C2 |
H8 |
108.004 |
|
C2 |
C1 |
F4 |
109.926 |
| C2 |
C1 |
H5 |
111.519 |
|
C2 |
C1 |
H6 |
111.494 |
| C2 |
C3 |
H9 |
110.946 |
|
C2 |
C3 |
H10 |
110.759 |
| C2 |
C3 |
H11 |
111.156 |
|
C3 |
C2 |
H7 |
110.005 |
| C3 |
C2 |
H8 |
110.311 |
|
F4 |
C1 |
H5 |
107.489 |
| F4 |
C1 |
H6 |
107.410 |
|
H5 |
C1 |
H6 |
108.835 |
| H7 |
C2 |
H8 |
106.877 |
|
H9 |
C3 |
H10 |
108.122 |
| H9 |
C3 |
H11 |
107.793 |
|
H10 |
C3 |
H11 |
107.932 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
