Jump to
S1C2
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -192.764569 |
| Energy at 298.15K | -192.759536 |
| HF Energy | -191.906744 |
| Nuclear repulsion energy | 129.297213 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3242 |
3242 |
27.54 |
|
|
|
| 2 |
A1 |
3211 |
3211 |
11.22 |
|
|
|
| 3 |
A1 |
1689 |
1689 |
0.00 |
|
|
|
| 4 |
A1 |
1628 |
1628 |
2.27 |
|
|
|
| 5 |
A1 |
1523 |
1523 |
10.99 |
|
|
|
| 6 |
A1 |
1139 |
1139 |
22.50 |
|
|
|
| 7 |
A1 |
1025 |
1025 |
31.80 |
|
|
|
| 8 |
A1 |
902 |
902 |
5.01 |
|
|
|
| 9 |
A2 |
3253 |
3253 |
0.00 |
|
|
|
| 10 |
A2 |
1340 |
1340 |
0.00 |
|
|
|
| 11 |
A2 |
1273 |
1273 |
0.00 |
|
|
|
| 12 |
A2 |
897 |
897 |
0.00 |
|
|
|
| 13 |
B1 |
3295 |
3295 |
77.68 |
|
|
|
| 14 |
B1 |
3250 |
3250 |
74.30 |
|
|
|
| 15 |
B1 |
1314 |
1314 |
0.09 |
|
|
|
| 16 |
B1 |
1266 |
1266 |
6.56 |
|
|
|
| 17 |
B1 |
820 |
820 |
0.01 |
|
|
|
| 18 |
B1 |
99 |
99 |
4.72 |
|
|
|
| 19 |
B2 |
3203 |
3203 |
170.83 |
|
|
|
| 20 |
B2 |
1658 |
1658 |
5.72 |
|
|
|
| 21 |
B2 |
1439 |
1439 |
0.01 |
|
|
|
| 22 |
B2 |
1388 |
1388 |
6.35 |
|
|
|
| 23 |
B2 |
1166 |
1166 |
135.13 |
|
|
|
| 24 |
B2 |
1004 |
1004 |
1.65 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20512.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 20512.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
1.049 |
| C2 |
0.000 |
0.000 |
-1.074 |
| C3 |
0.000 |
1.028 |
0.072 |
| C4 |
0.000 |
-1.028 |
0.072 |
| H5 |
0.885 |
0.000 |
-1.701 |
| H6 |
-0.885 |
0.000 |
-1.701 |
| H7 |
0.886 |
1.654 |
0.147 |
| H8 |
-0.886 |
1.654 |
0.147 |
| H9 |
-0.886 |
-1.654 |
0.147 |
| H10 |
0.886 |
-1.654 |
0.147 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1228 | 1.4183 | 1.4183 | 2.8893 | 2.8893 | 2.0818 | 2.0818 | 2.0818 | 2.0818 |
C2 | 2.1228 | | 1.5394 | 1.5394 | 1.0849 | 1.0849 | 2.2388 | 2.2388 | 2.2388 | 2.2388 | C3 | 1.4183 | 1.5394 | | 2.0562 | 2.2327 | 2.2327 | 1.0874 | 1.0874 | 2.8257 | 2.8257 | C4 | 1.4183 | 1.5394 | 2.0562 | | 2.2327 | 2.2327 | 2.8257 | 2.8257 | 1.0874 | 1.0874 | H5 | 2.8893 | 1.0849 | 2.2327 | 2.2327 | | 1.7699 | 2.4808 | 3.0480 | 3.0480 | 2.4808 | H6 | 2.8893 | 1.0849 | 2.2327 | 2.2327 | 1.7699 | | 3.0480 | 2.4808 | 2.4808 | 3.0480 | H7 | 2.0818 | 2.2388 | 1.0874 | 2.8257 | 2.4808 | 3.0480 | | 1.7717 | 3.7528 | 3.3083 | H8 | 2.0818 | 2.2388 | 1.0874 | 2.8257 | 3.0480 | 2.4808 | 1.7717 | | 3.3083 | 3.7528 | H9 | 2.0818 | 2.2388 | 2.8257 | 1.0874 | 3.0480 | 2.4808 | 3.7528 | 3.3083 | | 1.7717 | H10 | 2.0818 | 2.2388 | 2.8257 | 1.0874 | 2.4808 | 3.0480 | 3.3083 | 3.7528 | 1.7717 | |
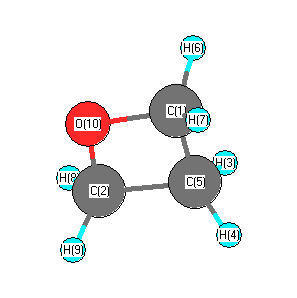 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
91.641 |
|
O1 |
C3 |
H7 |
111.684 |
| O1 |
C3 |
H8 |
111.684 |
|
O1 |
C4 |
C2 |
91.641 |
| O1 |
C4 |
H9 |
111.684 |
|
O1 |
C4 |
H10 |
111.684 |
| C2 |
C3 |
H7 |
115.861 |
|
C2 |
C3 |
H8 |
115.861 |
| C2 |
C4 |
H9 |
115.861 |
|
C2 |
C4 |
H10 |
115.861 |
| C3 |
O1 |
C4 |
92.914 |
|
C3 |
C2 |
C4 |
83.805 |
| C3 |
C2 |
H5 |
115.505 |
|
C3 |
C2 |
H6 |
115.505 |
| C4 |
C2 |
H5 |
115.505 |
|
C4 |
C2 |
H6 |
115.505 |
| H5 |
C2 |
H6 |
109.306 |
|
H7 |
C3 |
H8 |
109.113 |
| H9 |
C4 |
H10 |
109.113 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -192.764536 |
| Energy at 298.15K | -192.772179 |
| HF Energy | -191.906746 |
| Nuclear repulsion energy | 129.323158 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3297 |
3297 |
76.70 |
|
|
|
| 2 |
A' |
3252 |
3252 |
75.50 |
|
|
|
| 3 |
A' |
3244 |
3244 |
27.66 |
|
|
|
| 4 |
A' |
3213 |
3213 |
11.05 |
|
|
|
| 5 |
A' |
1689 |
1689 |
0.00 |
|
|
|
| 6 |
A' |
1628 |
1628 |
2.31 |
|
|
|
| 7 |
A' |
1523 |
1523 |
11.09 |
|
|
|
| 8 |
A' |
1314 |
1314 |
0.09 |
|
|
|
| 9 |
A' |
1266 |
1266 |
6.57 |
|
|
|
| 10 |
A' |
1140 |
1140 |
22.76 |
|
|
|
| 11 |
A' |
1025 |
1025 |
31.55 |
|
|
|
| 12 |
A' |
903 |
903 |
4.96 |
|
|
|
| 13 |
A' |
819 |
819 |
0.01 |
|
|
|
| 14 |
A' |
94 |
94 |
4.71 |
|
|
|
| 15 |
A" |
3255 |
3255 |
0.00 |
|
|
|
| 16 |
A" |
3205 |
3205 |
171.11 |
|
|
|
| 17 |
A" |
1658 |
1658 |
5.74 |
|
|
|
| 18 |
A" |
1438 |
1438 |
0.01 |
|
|
|
| 19 |
A" |
1387 |
1387 |
6.43 |
|
|
|
| 20 |
A" |
1340 |
1340 |
0.00 |
|
|
|
| 21 |
A" |
1272 |
1272 |
0.00 |
|
|
|
| 22 |
A" |
1168 |
1168 |
135.15 |
|
|
|
| 23 |
A" |
1004 |
1004 |
1.64 |
|
|
|
| 24 |
A" |
896 |
896 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20514.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 20514.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.217 |
-1.027 |
0.000 |
| C2 |
-0.059 |
1.066 |
0.000 |
| C3 |
-0.059 |
-0.073 |
1.024 |
| C4 |
-0.059 |
-0.073 |
-1.024 |
| H5 |
0.866 |
1.631 |
0.000 |
| H6 |
-0.904 |
1.744 |
0.000 |
| H7 |
0.707 |
-0.072 |
1.790 |
| H8 |
-1.028 |
-0.268 |
1.473 |
| H9 |
0.707 |
-0.072 |
-1.790 |
| H10 |
-1.028 |
-0.268 |
-1.473 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1107 | 1.4269 | 1.4269 | 2.7363 | 2.9887 | 2.0879 | 2.0726 | 2.0879 | 2.0726 |
C2 | 2.1107 | | 1.5313 | 1.5313 | 1.0839 | 1.0836 | 2.2553 | 2.2108 | 2.2553 | 2.2108 | C3 | 1.4269 | 1.5313 | | 2.0488 | 2.1925 | 2.2501 | 1.0834 | 1.0858 | 2.9172 | 2.6863 | C4 | 1.4269 | 1.5313 | 2.0488 | | 2.1925 | 2.2501 | 2.9172 | 2.6863 | 1.0834 | 1.0858 | H5 | 2.7363 | 1.0839 | 2.1925 | 2.1925 | | 1.7736 | 2.4760 | 3.0599 | 2.4760 | 3.0599 | H6 | 2.9887 | 1.0836 | 2.2501 | 2.2501 | 1.7736 | | 3.0163 | 2.4964 | 3.0163 | 2.4964 | H7 | 2.0879 | 2.2553 | 1.0834 | 2.9172 | 2.4760 | 3.0163 | | 1.7748 | 3.5810 | 3.7016 | H8 | 2.0726 | 2.2108 | 1.0858 | 2.6863 | 3.0599 | 2.4964 | 1.7748 | | 3.7016 | 2.9467 | H9 | 2.0879 | 2.2553 | 2.9172 | 1.0834 | 2.4760 | 3.0163 | 3.5810 | 3.7016 | | 1.7748 | H10 | 2.0726 | 2.2108 | 2.6863 | 1.0858 | 3.0599 | 2.4964 | 3.7016 | 2.9467 | 1.7748 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
90.970 |
|
O1 |
C3 |
H7 |
111.829 |
| O1 |
C3 |
H8 |
110.414 |
|
O1 |
C4 |
C2 |
90.970 |
| O1 |
C4 |
H9 |
111.829 |
|
O1 |
C4 |
H10 |
110.414 |
| C2 |
C3 |
H7 |
118.194 |
|
C2 |
C3 |
H8 |
114.215 |
| C2 |
C4 |
H9 |
118.194 |
|
C2 |
C4 |
H10 |
114.215 |
| C3 |
O1 |
C4 |
91.765 |
|
C3 |
C2 |
C4 |
83.974 |
| C3 |
C2 |
H5 |
112.813 |
|
C3 |
C2 |
H6 |
117.718 |
| C4 |
C2 |
H5 |
112.813 |
|
C4 |
C2 |
H6 |
117.718 |
| H5 |
C2 |
H6 |
109.822 |
|
H7 |
C3 |
H8 |
109.808 |
| H9 |
C4 |
H10 |
109.808 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
