Jump to
S1C2
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -267.955707 |
| Energy at 298.15K | -267.966372 |
| Nuclear repulsion energy | 195.700953 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4126 |
4126 |
45.28 |
|
|
|
| 2 |
A |
4096 |
4096 |
110.77 |
|
|
|
| 3 |
A |
3257 |
3257 |
68.38 |
|
|
|
| 4 |
A |
3249 |
3249 |
61.02 |
|
|
|
| 5 |
A |
3209 |
3209 |
87.35 |
|
|
|
| 6 |
A |
3194 |
3194 |
23.29 |
|
|
|
| 7 |
A |
3167 |
3167 |
66.35 |
|
|
|
| 8 |
A |
3144 |
3144 |
70.43 |
|
|
|
| 9 |
A |
1668 |
1668 |
1.88 |
|
|
|
| 10 |
A |
1654 |
1654 |
0.07 |
|
|
|
| 11 |
A |
1611 |
1611 |
11.42 |
|
|
|
| 12 |
A |
1599 |
1599 |
49.08 |
|
|
|
| 13 |
A |
1587 |
1587 |
64.69 |
|
|
|
| 14 |
A |
1525 |
1525 |
7.09 |
|
|
|
| 15 |
A |
1506 |
1506 |
0.50 |
|
|
|
| 16 |
A |
1428 |
1428 |
42.09 |
|
|
|
| 17 |
A |
1396 |
1396 |
8.38 |
|
|
|
| 18 |
A |
1333 |
1333 |
9.91 |
|
|
|
| 19 |
A |
1306 |
1306 |
129.70 |
|
|
|
| 20 |
A |
1238 |
1238 |
28.62 |
|
|
|
| 21 |
A |
1212 |
1212 |
65.76 |
|
|
|
| 22 |
A |
1194 |
1194 |
54.59 |
|
|
|
| 23 |
A |
1040 |
1040 |
2.46 |
|
|
|
| 24 |
A |
995 |
995 |
7.19 |
|
|
|
| 25 |
A |
980 |
980 |
9.79 |
|
|
|
| 26 |
A |
874 |
874 |
7.47 |
|
|
|
| 27 |
A |
548 |
548 |
83.08 |
|
|
|
| 28 |
A |
534 |
534 |
138.30 |
|
|
|
| 29 |
A |
416 |
416 |
8.07 |
|
|
|
| 30 |
A |
356 |
356 |
67.53 |
|
|
|
| 31 |
A |
312 |
312 |
59.07 |
|
|
|
| 32 |
A |
209 |
209 |
3.39 |
|
|
|
| 33 |
A |
119 |
119 |
1.61 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27040.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 27040.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.018 |
1.032 |
-0.340 |
| C2 |
1.301 |
0.457 |
0.155 |
| C3 |
-1.248 |
0.399 |
0.316 |
| H4 |
-0.091 |
0.903 |
-1.418 |
| H5 |
-0.012 |
2.104 |
-0.138 |
| O6 |
1.296 |
-0.937 |
-0.060 |
| O7 |
-1.519 |
-0.888 |
-0.152 |
| H8 |
2.098 |
-1.324 |
0.248 |
| H9 |
-0.725 |
-1.397 |
-0.116 |
| H10 |
2.135 |
0.917 |
-0.375 |
| H11 |
1.423 |
0.669 |
1.218 |
| H12 |
-1.127 |
0.402 |
1.403 |
| H13 |
-2.127 |
0.995 |
0.091 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
H4 |
H5 |
O6 |
O7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5220 | 1.5306 | 1.0882 | 1.0903 | 2.3843 | 2.4441 | 3.2210 | 2.5402 | 2.1565 | 2.1537 | 2.1593 | 2.1529 |
C2 | 1.5220 | | 2.5546 | 2.1467 | 2.1268 | 1.4102 | 3.1387 | 1.9527 | 2.7593 | 1.0901 | 1.0914 | 2.7303 | 3.4709 | C3 | 1.5306 | 2.5546 | | 2.1440 | 2.1538 | 2.8982 | 1.3959 | 3.7640 | 1.9205 | 3.4913 | 2.8326 | 1.0940 | 1.0857 | H4 | 1.0882 | 2.1467 | 2.1440 | | 1.7571 | 2.6740 | 2.6162 | 3.5384 | 2.7174 | 2.4579 | 3.0490 | 3.0461 | 2.5359 | H5 | 1.0903 | 2.1268 | 2.1538 | 1.7571 | | 3.3113 | 3.3496 | 4.0434 | 3.5731 | 2.4643 | 2.4413 | 2.5520 | 2.3994 | O6 | 2.3843 | 1.4102 | 2.8982 | 2.6740 | 3.3113 | | 2.8167 | 0.9419 | 2.0733 | 2.0595 | 2.0566 | 3.1312 | 3.9339 | O7 | 2.4441 | 3.1387 | 1.3959 | 2.6162 | 3.3496 | 2.8167 | | 3.6646 | 0.9442 | 4.0812 | 3.5995 | 2.0581 | 1.9934 | H8 | 3.2210 | 1.9527 | 3.7640 | 3.5384 | 4.0434 | 0.9419 | 3.6646 | | 2.8469 | 2.3263 | 2.3166 | 3.8355 | 4.8221 | H9 | 2.5402 | 2.7593 | 1.9205 | 2.7174 | 3.5731 | 2.0733 | 0.9442 | 2.8469 | | 3.6880 | 3.2656 | 2.3891 | 2.7808 | H10 | 2.1565 | 1.0901 | 3.4913 | 2.4579 | 2.4643 | 2.0595 | 4.0812 | 2.3263 | 3.6880 | | 1.7627 | 3.7502 | 4.2883 | H11 | 2.1537 | 1.0914 | 2.8326 | 3.0490 | 2.4413 | 2.0566 | 3.5995 | 2.3166 | 3.2656 | 1.7627 | | 2.5706 | 3.7398 | H12 | 2.1593 | 2.7303 | 1.0940 | 3.0461 | 2.5520 | 3.1312 | 2.0581 | 3.8355 | 2.3891 | 3.7502 | 2.5706 | | 1.7535 | H13 | 2.1529 | 3.4709 | 1.0857 | 2.5359 | 2.3994 | 3.9339 | 1.9934 | 4.8221 | 2.7808 | 4.2883 | 3.7398 | 1.7535 | |
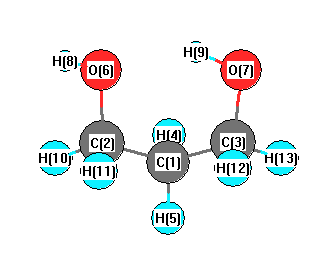 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
108.747 |
|
C1 |
C2 |
H10 |
110.194 |
| C1 |
C2 |
H11 |
109.896 |
|
C1 |
C3 |
O7 |
113.185 |
| C1 |
C3 |
H12 |
109.597 |
|
C1 |
C3 |
H13 |
109.581 |
| C2 |
C1 |
C3 |
113.626 |
|
C2 |
C1 |
H4 |
109.539 |
| C2 |
C1 |
H5 |
107.862 |
|
C2 |
O6 |
H8 |
110.661 |
| C3 |
C1 |
H4 |
108.732 |
|
C3 |
C1 |
H5 |
109.383 |
| C3 |
O7 |
H9 |
108.771 |
|
H4 |
C1 |
H5 |
107.525 |
| O6 |
C2 |
H10 |
110.260 |
|
O6 |
C2 |
H11 |
109.940 |
| O7 |
C3 |
H12 |
110.918 |
|
O7 |
C3 |
H13 |
106.211 |
| H10 |
C2 |
H11 |
107.798 |
|
H12 |
C3 |
H13 |
107.118 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -269.134608 |
| Energy at 298.15K | -269.127543 |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
Geometric Data calculated at CBS-Q
Point Group is C1
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
