Jump to
S1C2
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -151.708638 |
| Energy at 298.15K | -151.708183 |
| HF Energy | -151.083422 |
| Nuclear repulsion energy | 53.087089 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3566 |
3566 |
66.16 |
|
|
|
| 2 |
A' |
2169 |
2169 |
355.03 |
|
|
|
| 3 |
A' |
1370 |
1370 |
6.96 |
|
|
|
| 4 |
A' |
624 |
624 |
15.68 |
|
|
|
| 5 |
A' |
319 |
319 |
332.19 |
|
|
|
| 6 |
A" |
550 |
550 |
21.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4298.5 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 4298.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.044 |
0.000 |
| C2 |
0.866 |
-0.898 |
0.000 |
| O3 |
-0.888 |
0.778 |
0.000 |
| H4 |
1.910 |
-1.092 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.2792 | 1.1520 | 2.2225 |
C2 | 1.2792 | | 2.4256 | 1.0626 | O3 | 1.1520 | 2.4256 | | 3.3655 | H4 | 2.2225 | 1.0626 | 3.3655 | |
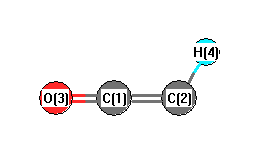 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
143.104 |
|
C2 |
C1 |
O3 |
172.159 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -151.708632 |
| Energy at 298.15K | -151.704020 |
| HF Energy | -151.083422 |
| Nuclear repulsion energy | 53.084261 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3565 |
3565 |
65.89 |
73.76 |
0.27 |
0.43 |
| 2 |
A' |
2168 |
2168 |
354.20 |
76.64 |
0.18 |
0.31 |
| 3 |
A' |
1369 |
1369 |
6.99 |
48.40 |
0.31 |
0.47 |
| 4 |
A' |
624 |
624 |
15.70 |
18.65 |
0.58 |
0.73 |
| 5 |
A' |
322 |
322 |
332.45 |
7.39 |
0.52 |
0.68 |
| 6 |
A" |
550 |
550 |
20.79 |
33.93 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 4299.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 4299.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.044 |
0.000 |
| C2 |
0.870 |
-0.894 |
0.000 |
| O3 |
-0.892 |
0.773 |
0.000 |
| H4 |
1.917 |
-1.081 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.2792 | 1.1522 | 2.2220 |
C2 | 1.2792 | | 2.4257 | 1.0627 | O3 | 1.1522 | 2.4257 | | 3.3652 | H4 | 2.2220 | 1.0627 | 3.3652 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
143.019 |
|
C2 |
C1 |
O3 |
172.117 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
