return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H10O (Methyl propyl ether)

using model chemistry: CBS-Q

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A'
Energy calculated at CBS-Q
 hartrees
Energy at 0K-233.205855
Energy at 298.15K-233.197992
HF Energy-232.137429
Nuclear repulsion energy189.568414
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3288 3288 50.34 97.05 0.51 0.67
2 A' 3258 3258 52.78 84.48 0.59 0.74
3 A' 3205 3205 25.18 101.15 0.15 0.26
4 A' 3183 3183 43.76 112.18 0.01 0.02
5 A' 3158 3158 80.47 134.12 0.05 0.09
6 A' 3136 3136 30.79 14.08 0.21 0.34
7 A' 1672 1672 1.19 9.27 0.75 0.86
8 A' 1646 1646 8.72 4.87 0.74 0.85
9 A' 1645 1645 3.23 5.20 0.74 0.85
10 A' 1629 1629 0.38 29.43 0.74 0.85
11 A' 1625 1625 0.63 5.29 0.67 0.80
12 A' 1578 1578 33.78 2.80 0.71 0.83
13 A' 1553 1553 3.31 1.40 0.58 0.74
14 A' 1460 1460 6.40 1.06 0.31 0.48
15 A' 1362 1362 83.30 1.87 0.35 0.52
16 A' 1288 1288 164.94 8.94 0.41 0.59
17 A' 1220 1220 9.55 3.59 0.21 0.35
18 A' 1127 1127 7.62 11.00 0.58 0.73
19 A' 1070 1070 25.60 3.89 0.69 0.81
20 A' 973 973 3.42 8.41 0.29 0.45
21 A' 467 467 0.71 3.35 0.19 0.32
22 A' 436 436 3.38 1.47 0.39 0.56
23 A' 207 207 1.61 0.14 0.17 0.30
24 A" 3258 3258 106.85 2.32 0.75 0.86
25 A" 3232 3232 0.21 110.53 0.75 0.86
26 A" 3206 3206 89.04 60.60 0.75 0.86
27 A" 3167 3167 54.13 58.33 0.75 0.86
28 A" 1636 1636 4.92 18.28 0.75 0.86
29 A" 1633 1633 4.56 17.58 0.75 0.86
30 A" 1438 1438 0.50 20.44 0.75 0.86
31 A" 1389 1389 2.84 0.22 0.75 0.86
32 A" 1310 1310 9.05 1.98 0.75 0.86
33 A" 1282 1282 0.23 4.95 0.75 0.86
34 A" 971 971 1.86 0.11 0.75 0.86
35 A" 817 817 0.55 0.05 0.75 0.86
36 A" 255 255 1.77 0.02 0.75 0.86
37 A" 243 243 3.81 0.07 0.75 0.86
38 A" 126 126 2.38 0.01 0.75 0.86
39 A" 109 109 1.66 0.14 0.75 0.86

Unscaled Zero Point Vibrational Energy (zpe) 32127.4 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 32127.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CBS-Q
ABC
0.65865 0.07097 0.06732

See section I.F.4 to change rotational constant units
Geometric Data calculated at CBS-Q

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.321 0.639 0.000
O2 -1.259 -0.258 0.000
C3 0.000 0.342 0.000
C4 1.062 -0.746 0.000
C5 2.482 -0.178 0.000
H6 -3.239 0.061 0.000
H7 -2.306 1.277 0.885
H8 -2.306 1.277 -0.885
H9 0.111 0.982 0.880
H10 0.111 0.982 -0.880
H11 0.909 -1.377 -0.873
H12 0.909 -1.377 0.873
H13 2.664 0.439 0.879
H14 2.664 0.439 -0.879
H15 3.220 -0.977 0.000

Atom - Atom Distances (Å)
  C1 O2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.39012.34033.65594.87261.08451.09181.09182.60942.60943.90683.90685.06665.06665.7723
O21.39011.39522.37233.74272.00532.05872.05872.04732.04732.59182.59184.08114.08114.5370
C32.34031.39521.52022.53613.25142.64172.64171.09381.09382.13162.13162.80742.80743.4797
C43.65592.37231.52021.52964.37634.02794.02792.15972.15971.08831.08832.17792.17792.1704
C54.87263.74272.53611.52965.72655.08265.08262.78242.78242.16232.16231.08921.08921.0876
H61.08452.00533.25144.37635.72651.77031.77033.58433.58434.47664.47665.98065.98066.5423
H71.09182.05872.64174.02795.08261.77031.77082.43543.00794.52544.16975.04095.34096.0339
H81.09182.05872.64174.02795.08261.77031.77083.00792.43544.16974.52545.34095.04096.0339
H92.60942.04731.09382.15972.78243.58432.43543.00791.75963.04572.49062.61043.14793.7786
H102.60942.04731.09382.15972.78243.58433.00792.43541.75962.49063.04573.14792.61043.7786
H113.90682.59182.13161.08832.16234.47664.52544.16973.04572.49061.74643.07392.52532.5027
H123.90682.59182.13161.08832.16234.47664.16974.52542.49063.04571.74642.52533.07392.5027
H135.06664.08112.80742.17791.08925.98065.04095.34092.61043.14793.07392.52531.75881.7565
H145.06664.08112.80742.17791.08925.98065.34095.04093.14792.61042.52533.07391.75881.7565
H155.77234.53703.47972.17041.08766.54236.03396.03393.77863.77862.50272.50271.75651.7565

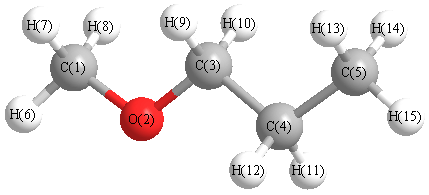
picture of Methyl propyl ether state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O2 C3 114.331 O2 C1 H6 107.615
O2 C1 H7 111.527 O2 C1 H8 111.527
O2 C3 C4 108.843 O2 C3 H9 110.092
O2 C3 H10 110.092 C3 C4 C5 112.517
C3 C4 H11 108.473 C3 C4 H12 108.473
C4 C3 H9 110.358 C4 C3 H10 110.358
C4 C5 H13 111.425 C4 C5 H14 111.425
C4 C5 H15 110.924 C5 C4 H11 110.236
C5 C4 H12 110.236 H6 C1 H7 108.863
H6 C1 H8 108.863 H7 C1 H8 108.381
H9 C3 H10 107.090 H11 C4 H12 106.712
H13 C5 H14 107.689 H13 C5 H15 107.595
H14 C5 H15 107.595
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability