return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 (2,3-dimethylbut-1-ene)

using model chemistry: CBS-Q

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A'
Energy calculated at CBS-Q
 hartrees
Energy at 0K-235.329408
Energy at 298.15K-235.320470
HF Energy-234.174319
Nuclear repulsion energy243.738956
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3382 3382 26.98      
2 A' 3308 3308 15.28      
3 A' 3274 3274 32.85      
4 A' 3261 3261 64.69      
5 A' 3255 3255 98.47      
6 A' 3200 3200 3.84      
7 A' 3191 3191 46.12      
8 A' 3186 3186 11.64      
9 A' 1874 1874 16.89      
10 A' 1649 1649 3.86      
11 A' 1644 1644 6.67      
12 A' 1633 1633 8.54      
13 A' 1589 1589 2.26      
14 A' 1559 1559 1.57      
15 A' 1550 1550 4.03      
16 A' 1467 1467 7.00      
17 A' 1418 1418 0.46      
18 A' 1292 1292 3.15      
19 A' 1208 1208 8.52      
20 A' 1096 1096 0.80      
21 A' 1043 1043 0.24      
22 A' 959 959 0.74      
23 A' 773 773 0.66      
24 A' 556 556 0.78      
25 A' 478 478 1.23      
26 A' 354 354 0.16      
27 A' 297 297 0.42      
28 A' 276 276 0.10      
29 A" 3258 3258 14.03      
30 A" 3250 3250 25.07      
31 A" 3240 3240 29.37      
32 A" 3183 3183 31.26      
33 A" 1637 1637 0.35      
34 A" 1625 1625 0.80      
35 A" 1618 1618 6.39      
36 A" 1538 1538 3.54      
37 A" 1469 1469 1.91      
38 A" 1219 1219 2.73      
39 A" 1160 1160 0.70      
40 A" 1038 1038 36.62      
41 A" 1035 1035 9.19      
42 A" 1002 1002 0.15      
43 A" 805 805 0.61      
44 A" 599 599 6.49      
45 A" 282 282 0.04      
46 A" 231 231 0.27      
47 A" 182 182 0.07      
48 A" 41 41 0.10      

Unscaled Zero Point Vibrational Energy (zpe) 38590.7 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 38590.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CBS-Q
ABC
0.14706 0.08313 0.08211

See section I.F.4 to change rotational constant units
Geometric Data calculated at CBS-Q

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.951 -1.807 0.000
C2 -0.065 -0.823 0.000
C3 -0.514 0.633 0.000
C4 1.421 -1.113 0.000
C5 -0.065 1.382 1.265
C6 -0.065 1.382 -1.265
H7 -0.652 -2.844 0.000
H8 -2.014 -1.617 0.000
H9 -1.603 0.624 0.000
H10 1.607 -2.183 0.000
H11 1.911 -0.690 -0.875
H12 1.911 -0.690 0.875
H13 -0.499 2.380 1.289
H14 -0.381 0.858 2.165
H15 1.017 1.497 1.305
H16 -0.499 2.380 -1.289
H17 -0.381 0.858 -2.165
H18 1.017 1.497 -1.305

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.32442.47862.47123.54343.54341.07941.07942.51632.58533.19423.19424.40443.48024.06134.40443.48024.0613
C21.32441.52361.51362.54212.54212.10502.10452.11152.15522.16472.16473.47992.75872.87333.47992.75872.8733
C32.47861.52362.60651.53771.53773.47982.70371.08873.52572.89782.89782.17132.18072.18982.17132.18072.1898
C42.47121.51362.60653.16763.16762.70063.47123.48711.08621.08881.08884.18943.43742.94644.18943.43742.9464
C53.54342.54211.53773.16762.53064.45053.79392.13154.13603.57472.88921.08891.08821.08842.77643.48452.7910
C63.54342.54211.53773.16762.53064.45053.79392.13154.13602.88923.57472.77643.48452.79101.08891.08821.0884
H71.07942.10503.47982.70064.45054.45051.83293.59562.35353.46043.46045.38334.29694.83085.38334.29694.8308
H81.07942.10452.70373.47123.79393.79391.83292.27783.66444.12624.12624.46483.67114.53724.46483.67114.5372
H92.51632.11151.08873.48712.13152.13153.59562.27784.26393.85203.85202.44282.49723.05472.44282.49723.0547
H102.58532.15523.52571.08624.13604.13602.35353.66444.26391.75751.75755.18864.22893.94955.18864.22893.9495
H113.19422.16472.89781.08883.57472.88923.46044.12623.85201.75751.75104.46284.10963.21533.92473.05082.4016
H123.19422.16472.89781.08882.88923.57473.46044.12623.85201.75751.75103.92473.05082.40164.46284.10963.2153
H134.40443.47992.17134.18941.08892.77645.38334.46482.44285.18864.46283.92471.76091.75432.57763.77643.1313
H143.48022.75872.18073.43741.08823.48454.29693.67112.49724.22894.10963.05081.76091.76123.77644.32993.7951
H154.06132.87332.18982.94641.08842.79104.83084.53723.05473.94953.21532.40161.75431.76123.13133.79512.6101
H164.40443.47992.17134.18942.77641.08895.38334.46482.44285.18863.92474.46282.57763.77643.13131.76091.7543
H173.48022.75872.18073.43743.48451.08824.29693.67112.49724.22893.05084.10963.77644.32993.79511.76091.7612
H184.06132.87332.18982.94642.79101.08844.83084.53723.05473.94952.40163.21533.13133.79512.61011.75431.7612

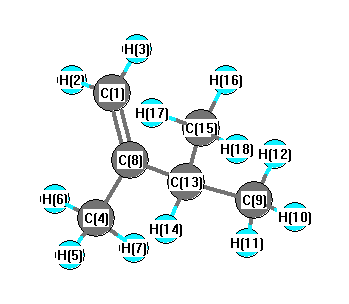
picture of 2,3-dimethylbut-1-ene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.826 C1 C2 C4 120.946
C2 C1 H7 121.917 C2 C1 H8 121.876
C2 C3 C5 112.281 C2 C3 C6 112.281
C2 C3 H9 106.667 C2 C4 H10 110.919
C2 C4 H11 111.528 C2 C4 H12 111.528
C3 C2 C4 118.229 C3 C5 H13 110.353
C3 C5 H14 111.135 C3 C5 H15 111.854
C3 C6 H16 110.353 C3 C6 H17 111.135
C3 C6 H18 111.854 C5 C3 C6 110.749
C5 C3 H9 107.259 C6 C3 H9 107.259
H7 C1 H8 116.208 H10 C4 H11 107.813
H10 C4 H12 107.813 H11 C4 H12 107.042
H13 C5 H14 107.958 H13 C5 H15 107.354
H14 C5 H15 108.021 H16 C6 H17 107.958
H16 C6 H18 107.354 H17 C6 H18 108.021
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability