Jump to
S1C2
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -551.019972 |
| Energy at 298.15K | -551.013345 |
| HF Energy | -548.939662 |
| Nuclear repulsion energy | 358.743655 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
1902 |
1902 |
0.00 |
|
|
|
| 2 |
Ag |
1570 |
1570 |
0.00 |
|
|
|
| 3 |
Ag |
714 |
714 |
0.00 |
|
|
|
| 4 |
Ag |
628 |
628 |
0.00 |
|
|
|
| 5 |
Ag |
666i |
666i |
0.00 |
|
|
|
| 6 |
Au |
1619 |
1619 |
1099.76 |
|
|
|
| 7 |
Au |
1062 |
1062 |
29.44 |
|
|
|
| 8 |
Au |
271 |
271 |
7.56 |
|
|
|
| 9 |
Bg |
827 |
827 |
0.00 |
|
|
|
| 10 |
Bg |
742 |
742 |
0.00 |
|
|
|
| 11 |
Bg |
239 |
239 |
0.00 |
|
|
|
| 12 |
Bg |
5813i |
5813i |
0.00 |
|
|
|
| 13 |
Bu |
1837 |
1837 |
8.04 |
|
|
|
| 14 |
Bu |
1049 |
1049 |
403.61 |
|
|
|
| 15 |
Bu |
726 |
726 |
159.01 |
|
|
|
| 16 |
Bu |
318 |
318 |
1.27 |
|
|
|
| 17 |
Bu |
154 |
154 |
0.00 |
|
|
|
| 18 |
Bu |
238i |
238i |
22.98 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3468.7 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 3468.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.956 |
0.000 |
| C2 |
0.000 |
0.000 |
1.042 |
| C3 |
0.000 |
-0.956 |
0.000 |
| C4 |
0.000 |
0.000 |
-1.042 |
| F5 |
0.000 |
2.261 |
0.000 |
| F6 |
0.000 |
0.000 |
2.393 |
| F7 |
0.000 |
-2.261 |
0.000 |
| F8 |
0.000 |
0.000 |
-2.393 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
| C1 | | 1.4137 | 1.9115 | 1.4137 | 1.3055 | 2.5766 | 3.2170 | 2.5766 |
C2 | 1.4137 | | 1.4137 | 2.0834 | 2.4897 | 1.3511 | 2.4897 | 3.4345 | C3 | 1.9115 | 1.4137 | | 1.4137 | 3.2170 | 2.5766 | 1.3055 | 2.5766 | C4 | 1.4137 | 2.0834 | 1.4137 | | 2.4897 | 3.4345 | 2.4897 | 1.3511 | F5 | 1.3055 | 2.4897 | 3.2170 | 2.4897 | | 3.2922 | 4.5225 | 3.2922 | F6 | 2.5766 | 1.3511 | 2.5766 | 3.4345 | 3.2922 | | 3.2922 | 4.7856 | F7 | 3.2170 | 2.4897 | 1.3055 | 2.4897 | 4.5225 | 3.2922 | | 3.2922 | F8 | 2.5766 | 3.4345 | 2.5766 | 1.3511 | 3.2922 | 4.7856 | 3.2922 | |
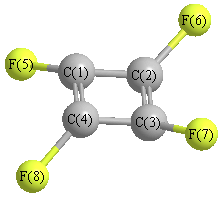 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
85.072 |
|
C1 |
C2 |
F6 |
137.464 |
| C1 |
C4 |
C3 |
85.072 |
|
C1 |
C4 |
F8 |
137.464 |
| C2 |
C1 |
C4 |
94.928 |
|
C2 |
C1 |
F5 |
132.536 |
| C2 |
C3 |
C4 |
94.928 |
|
C2 |
C3 |
F7 |
132.536 |
| C3 |
C2 |
F6 |
137.464 |
|
C3 |
C4 |
F8 |
137.464 |
| C4 |
C1 |
F5 |
132.536 |
|
C4 |
C3 |
F7 |
132.536 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -551.016257 |
| Energy at 298.15K | -551.017006 |
| HF Energy | -548.999616 |
| Nuclear repulsion energy | 356.628733 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
2094 |
2094 |
0.00 |
|
|
|
| 2 |
Ag |
1378 |
1378 |
0.00 |
|
|
|
| 3 |
Ag |
691 |
691 |
0.00 |
|
|
|
| 4 |
Ag |
242 |
242 |
0.00 |
|
|
|
| 5 |
Ag |
185 |
185 |
0.00 |
|
|
|
| 6 |
Au |
1470 |
1470 |
527.56 |
|
|
|
| 7 |
Au |
1021 |
1021 |
97.53 |
|
|
|
| 8 |
Au |
739 |
739 |
0.00 |
|
|
|
| 9 |
Au |
243 |
243 |
0.31 |
|
|
|
| 10 |
Au |
182 |
182 |
0.00 |
|
|
|
| 11 |
Bg |
1516 |
1516 |
0.00 |
|
|
|
| 12 |
Bg |
824 |
824 |
0.00 |
|
|
|
| 13 |
Bg |
597 |
597 |
0.00 |
|
|
|
| 14 |
Bg |
513 |
513 |
0.00 |
|
|
|
| 15 |
Bu |
2099 |
2099 |
94.76 |
|
|
|
| 16 |
Bu |
1062 |
1062 |
232.72 |
|
|
|
| 17 |
Bu |
329 |
329 |
11.80 |
|
|
|
| 18 |
Bu |
242 |
242 |
2.22 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7713.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 7713.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.117 |
0.758 |
0.652 |
| C2 |
-0.117 |
-0.758 |
0.652 |
| C3 |
-0.117 |
-0.758 |
-0.652 |
| C4 |
0.117 |
0.758 |
-0.652 |
| F5 |
-0.117 |
1.654 |
1.569 |
| F6 |
0.117 |
-1.654 |
1.569 |
| F7 |
0.117 |
-1.654 |
-1.569 |
| F8 |
-0.117 |
1.654 |
-1.569 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
| C1 | | 1.5344 | 2.0136 | 1.3038 | 1.3028 | 2.5804 | 3.2787 | 2.4060 |
C2 | 1.5344 | | 1.3038 | 2.0136 | 2.5804 | 1.3028 | 2.4060 | 3.2787 | C3 | 2.0136 | 1.3038 | | 1.5344 | 3.2787 | 2.4060 | 1.3028 | 2.5804 | C4 | 1.3038 | 2.0136 | 1.5344 | | 2.4060 | 3.2787 | 2.5804 | 1.3028 | F5 | 1.3028 | 2.5804 | 3.2787 | 2.4060 | | 3.3154 | 4.5651 | 3.1382 | F6 | 2.5804 | 1.3028 | 2.4060 | 3.2787 | 3.3154 | | 3.1382 | 4.5651 | F7 | 3.2787 | 2.4060 | 1.3028 | 2.5804 | 4.5651 | 3.1382 | | 3.3154 | F8 | 2.4060 | 3.2787 | 2.5804 | 1.3028 | 3.1382 | 4.5651 | 3.3154 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
90.000 |
|
C1 |
C2 |
F6 |
130.686 |
| C1 |
C4 |
C3 |
90.000 |
|
C1 |
C4 |
F8 |
134.748 |
| C2 |
C1 |
C4 |
90.000 |
|
C2 |
C1 |
F5 |
130.686 |
| C2 |
C3 |
C4 |
90.000 |
|
C2 |
C3 |
F7 |
134.748 |
| C3 |
C2 |
F6 |
134.748 |
|
C3 |
C4 |
F8 |
130.686 |
| C4 |
C1 |
F5 |
134.748 |
|
C4 |
C3 |
F7 |
130.686 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
