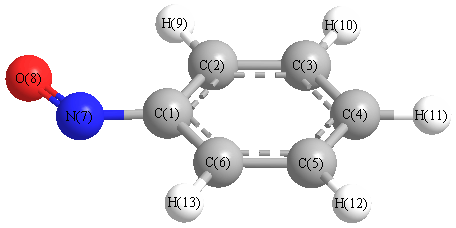
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3402 |
3402 |
3.00 |
84.95 |
0.18 |
0.30 |
| 2 |
A' |
3386 |
3386 |
11.53 |
177.06 |
0.12 |
0.22 |
| 3 |
A' |
3374 |
3374 |
20.17 |
58.26 |
0.28 |
0.44 |
| 4 |
A' |
3365 |
3365 |
11.22 |
106.47 |
0.73 |
0.84 |
| 5 |
A' |
3350 |
3350 |
0.71 |
49.54 |
0.75 |
0.86 |
| 6 |
A' |
1936 |
1936 |
199.53 |
83.17 |
0.29 |
0.45 |
| 7 |
A' |
1799 |
1799 |
3.47 |
41.84 |
0.47 |
0.64 |
| 8 |
A' |
1784 |
1784 |
51.26 |
76.47 |
0.51 |
0.68 |
| 9 |
A' |
1646 |
1646 |
0.80 |
4.75 |
0.30 |
0.47 |
| 10 |
A' |
1613 |
1613 |
17.44 |
2.17 |
0.53 |
0.69 |
| 11 |
A' |
1451 |
1451 |
6.53 |
1.00 |
0.43 |
0.60 |
| 12 |
A' |
1350 |
1350 |
13.50 |
0.23 |
0.72 |
0.84 |
| 13 |
A' |
1301 |
1301 |
31.46 |
30.59 |
0.32 |
0.49 |
| 14 |
A' |
1257 |
1257 |
61.00 |
36.29 |
0.17 |
0.30 |
| 15 |
A' |
1230 |
1230 |
21.11 |
13.63 |
0.73 |
0.85 |
| 16 |
A' |
1172 |
1172 |
0.67 |
0.65 |
0.57 |
0.72 |
| 17 |
A' |
1116 |
1116 |
1.43 |
8.84 |
0.11 |
0.19 |
| 18 |
A' |
1086 |
1086 |
0.92 |
26.66 |
0.10 |
0.18 |
| 19 |
A' |
903 |
903 |
22.01 |
7.08 |
0.38 |
0.55 |
| 20 |
A' |
731 |
731 |
15.33 |
3.64 |
0.17 |
0.29 |
| 21 |
A' |
669 |
669 |
0.05 |
6.51 |
0.75 |
0.86 |
| 22 |
A' |
484 |
484 |
0.15 |
2.57 |
0.33 |
0.49 |
| 23 |
A' |
283 |
283 |
2.82 |
2.49 |
0.48 |
0.65 |
| 24 |
A" |
1122 |
1122 |
0.11 |
0.57 |
0.75 |
0.86 |
| 25 |
A" |
1102 |
1102 |
0.03 |
0.02 |
0.75 |
0.86 |
| 26 |
A" |
1064 |
1064 |
5.01 |
1.96 |
0.75 |
0.86 |
| 27 |
A" |
954 |
954 |
0.00 |
3.00 |
0.75 |
0.86 |
| 28 |
A" |
852 |
852 |
61.54 |
0.05 |
0.75 |
0.86 |
| 29 |
A" |
752 |
752 |
22.65 |
0.54 |
0.75 |
0.86 |
| 30 |
A" |
520 |
520 |
2.00 |
0.29 |
0.75 |
0.86 |
| 31 |
A" |
459 |
459 |
0.03 |
0.04 |
0.75 |
0.86 |
| 32 |
A" |
275 |
275 |
0.13 |
3.89 |
0.75 |
0.86 |
| 33 |
A" |
115 |
115 |
0.42 |
0.30 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 22950.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 22950.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.