Jump to
S1C2
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -616.033578 |
| Energy at 298.15K | -616.026330 |
| HF Energy | -615.008129 |
| Nuclear repulsion energy | 205.374791 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3355 |
3355 |
8.66 |
|
|
|
| 2 |
A' |
3302 |
3302 |
24.42 |
|
|
|
| 3 |
A' |
3265 |
3265 |
24.29 |
|
|
|
| 4 |
A' |
3249 |
3249 |
25.70 |
|
|
|
| 5 |
A' |
3182 |
3182 |
38.68 |
|
|
|
| 6 |
A' |
1918 |
1918 |
1.79 |
|
|
|
| 7 |
A' |
1629 |
1629 |
9.19 |
|
|
|
| 8 |
A' |
1618 |
1618 |
1.68 |
|
|
|
| 9 |
A' |
1554 |
1554 |
4.03 |
|
|
|
| 10 |
A' |
1465 |
1465 |
43.60 |
|
|
|
| 11 |
A' |
1447 |
1447 |
10.70 |
|
|
|
| 12 |
A' |
1420 |
1420 |
1.69 |
|
|
|
| 13 |
A' |
1204 |
1204 |
0.62 |
|
|
|
| 14 |
A' |
1115 |
1115 |
11.60 |
|
|
|
| 15 |
A' |
980 |
980 |
9.19 |
|
|
|
| 16 |
A' |
762 |
762 |
30.78 |
|
|
|
| 17 |
A' |
625 |
625 |
8.23 |
|
|
|
| 18 |
A' |
357 |
357 |
1.76 |
|
|
|
| 19 |
A' |
185 |
185 |
0.83 |
|
|
|
| 20 |
A" |
3302 |
3302 |
11.30 |
|
|
|
| 21 |
A" |
3236 |
3236 |
33.69 |
|
|
|
| 22 |
A" |
1616 |
1616 |
5.26 |
|
|
|
| 23 |
A" |
1319 |
1319 |
2.82 |
|
|
|
| 24 |
A" |
1179 |
1179 |
2.76 |
|
|
|
| 25 |
A" |
1100 |
1100 |
33.87 |
|
|
|
| 26 |
A" |
1045 |
1045 |
0.05 |
|
|
|
| 27 |
A" |
804 |
804 |
0.31 |
|
|
|
| 28 |
A" |
256 |
256 |
4.63 |
|
|
|
| 29 |
A" |
215 |
215 |
0.86 |
|
|
|
| 30 |
A" |
112 |
112 |
1.76 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23406.3 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 23406.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.313 |
0.272 |
0.000 |
| C2 |
0.000 |
1.007 |
0.000 |
| C3 |
1.220 |
0.502 |
0.000 |
| C4 |
2.479 |
1.326 |
0.000 |
| Cl5 |
-1.187 |
-1.518 |
0.000 |
| H6 |
-1.895 |
0.531 |
0.877 |
| H7 |
-1.895 |
0.531 |
-0.877 |
| H8 |
-0.133 |
2.081 |
0.000 |
| H9 |
1.351 |
-0.568 |
0.000 |
| H10 |
2.266 |
2.391 |
0.000 |
| H11 |
3.087 |
1.100 |
0.876 |
| H12 |
3.087 |
1.100 |
-0.876 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5044 | 2.5430 | 3.9361 | 1.7946 | 1.0840 | 1.0840 | 2.1599 | 2.7937 | 4.1591 | 4.5625 | 4.5625 |
C2 | 1.5044 | | 1.3197 | 2.4999 | 2.7899 | 2.1418 | 2.1418 | 1.0830 | 2.0752 | 2.6554 | 3.2105 | 3.2105 | C3 | 2.5430 | 1.3197 | | 1.5050 | 3.1425 | 3.2362 | 3.2362 | 2.0791 | 1.0789 | 2.1593 | 2.1478 | 2.1478 | C4 | 3.9361 | 2.4999 | 1.5050 | | 4.6402 | 4.5320 | 4.5320 | 2.7197 | 2.2046 | 1.0871 | 1.0895 | 1.0895 | Cl5 | 1.7946 | 2.7899 | 3.1425 | 4.6402 | | 2.3388 | 2.3388 | 3.7506 | 2.7104 | 5.2161 | 5.0889 | 5.0889 | H6 | 1.0840 | 2.1418 | 3.2362 | 4.5320 | 2.3388 | | 1.7534 | 2.5054 | 3.5383 | 4.6415 | 5.0152 | 5.3126 | H7 | 1.0840 | 2.1418 | 3.2362 | 4.5320 | 2.3388 | 1.7534 | | 2.5054 | 3.5383 | 4.6415 | 5.3126 | 5.0152 | H8 | 2.1599 | 1.0830 | 2.0791 | 2.7197 | 3.7506 | 2.5054 | 2.5054 | | 3.0372 | 2.4188 | 3.4785 | 3.4785 | H9 | 2.7937 | 2.0752 | 1.0789 | 2.2046 | 2.7104 | 3.5383 | 3.5383 | 3.0372 | | 3.0979 | 2.5625 | 2.5625 | H10 | 4.1591 | 2.6554 | 2.1593 | 1.0871 | 5.2161 | 4.6415 | 4.6415 | 2.4188 | 3.0979 | | 1.7630 | 1.7630 | H11 | 4.5625 | 3.2105 | 2.1478 | 1.0895 | 5.0889 | 5.0152 | 5.3126 | 3.4785 | 2.5625 | 1.7630 | | 1.7512 | H12 | 4.5625 | 3.2105 | 2.1478 | 1.0895 | 5.0889 | 5.3126 | 5.0152 | 3.4785 | 2.5625 | 1.7630 | 1.7512 | |
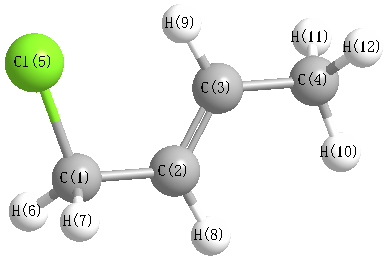 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
128.319 |
|
C1 |
C2 |
H8 |
112.162 |
| C2 |
C1 |
Cl5 |
115.205 |
|
C2 |
C1 |
H6 |
110.623 |
| C2 |
C1 |
H7 |
110.623 |
|
C2 |
C3 |
C4 |
124.382 |
| C2 |
C3 |
H9 |
119.471 |
|
C3 |
C2 |
H8 |
119.518 |
| C3 |
C4 |
H10 |
111.818 |
|
C3 |
C4 |
H11 |
110.740 |
| C3 |
C4 |
H12 |
110.740 |
|
C4 |
C3 |
H9 |
116.147 |
| Cl5 |
C1 |
H6 |
106.031 |
|
Cl5 |
C1 |
H7 |
106.031 |
| H6 |
C1 |
H7 |
107.952 |
|
H10 |
C4 |
H11 |
108.197 |
| H10 |
C4 |
H12 |
108.197 |
|
H11 |
C4 |
H12 |
106.973 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -616.034660 |
| Energy at 298.15K | -616.042319 |
| HF Energy | -615.010776 |
| Nuclear repulsion energy | 201.405449 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3343 |
3343 |
22.44 |
|
|
|
| 2 |
A |
3328 |
3328 |
2.01 |
|
|
|
| 3 |
A |
3311 |
3311 |
26.87 |
|
|
|
| 4 |
A |
3270 |
3270 |
15.10 |
|
|
|
| 5 |
A |
3268 |
3268 |
25.64 |
|
|
|
| 6 |
A |
3238 |
3238 |
31.53 |
|
|
|
| 7 |
A |
3183 |
3183 |
36.74 |
|
|
|
| 8 |
A |
1906 |
1906 |
10.81 |
|
|
|
| 9 |
A |
1628 |
1628 |
10.82 |
|
|
|
| 10 |
A |
1621 |
1621 |
1.59 |
|
|
|
| 11 |
A |
1616 |
1616 |
6.01 |
|
|
|
| 12 |
A |
1553 |
1553 |
3.55 |
|
|
|
| 13 |
A |
1467 |
1467 |
18.17 |
|
|
|
| 14 |
A |
1447 |
1447 |
1.78 |
|
|
|
| 15 |
A |
1406 |
1406 |
58.17 |
|
|
|
| 16 |
A |
1309 |
1309 |
1.23 |
|
|
|
| 17 |
A |
1216 |
1216 |
0.93 |
|
|
|
| 18 |
A |
1177 |
1177 |
2.55 |
|
|
|
| 19 |
A |
1156 |
1156 |
3.57 |
|
|
|
| 20 |
A |
1104 |
1104 |
34.09 |
|
|
|
| 21 |
A |
1031 |
1031 |
5.42 |
|
|
|
| 22 |
A |
966 |
966 |
2.37 |
|
|
|
| 23 |
A |
872 |
872 |
16.90 |
|
|
|
| 24 |
A |
748 |
748 |
82.03 |
|
|
|
| 25 |
A |
524 |
524 |
1.89 |
|
|
|
| 26 |
A |
381 |
381 |
4.14 |
|
|
|
| 27 |
A |
296 |
296 |
2.20 |
|
|
|
| 28 |
A |
220 |
220 |
0.75 |
|
|
|
| 29 |
A |
169 |
169 |
2.26 |
|
|
|
| 30 |
A |
93 |
93 |
0.49 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23422.3 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 23422.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.777 |
0.817 |
0.127 |
| C2 |
0.524 |
0.149 |
0.444 |
| C3 |
1.601 |
0.250 |
-0.316 |
| C4 |
2.933 |
-0.380 |
-0.017 |
| Cl5 |
-2.105 |
-0.390 |
-0.109 |
| H6 |
-1.113 |
1.454 |
0.934 |
| H7 |
-0.724 |
1.395 |
-0.786 |
| H8 |
0.550 |
-0.428 |
1.357 |
| H9 |
1.545 |
0.825 |
-1.232 |
| H10 |
2.908 |
-0.944 |
0.912 |
| H11 |
3.227 |
-1.057 |
-0.818 |
| H12 |
3.711 |
0.377 |
0.064 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4963 | 2.4839 | 3.9005 | 1.8103 | 1.0827 | 1.0814 | 2.1965 | 2.6895 | 4.1586 | 4.5203 | 4.5100 |
C2 | 1.4963 | | 1.3219 | 2.5086 | 2.7407 | 2.1506 | 2.1499 | 1.0804 | 2.0751 | 2.6638 | 3.2176 | 3.2177 | C3 | 2.4839 | 1.3219 | | 1.5039 | 3.7666 | 3.2219 | 2.6333 | 2.0888 | 1.0823 | 2.1550 | 2.1460 | 2.1483 | C4 | 3.9005 | 2.5086 | 1.5039 | | 5.0387 | 4.5433 | 4.1368 | 2.7503 | 2.2038 | 1.0867 | 1.0894 | 1.0895 | Cl5 | 1.8103 | 2.7407 | 3.7666 | 5.0387 | | 2.3399 | 2.3567 | 3.0338 | 4.0074 | 5.1460 | 5.4202 | 5.8695 | H6 | 1.0827 | 2.1506 | 3.2219 | 4.5433 | 2.3399 | | 1.7646 | 2.5473 | 3.4860 | 4.6821 | 5.3119 | 5.0195 | H7 | 1.0814 | 2.1499 | 2.6333 | 4.1368 | 2.3567 | 1.7646 | | 3.0883 | 2.3809 | 4.6414 | 4.6500 | 4.6290 | H8 | 2.1965 | 1.0804 | 2.0888 | 2.7503 | 3.0338 | 2.5473 | 3.0883 | | 3.0430 | 2.4540 | 3.5059 | 3.5087 | H9 | 2.6895 | 2.0751 | 1.0823 | 2.2038 | 4.0074 | 3.4860 | 2.3809 | 3.0430 | | 3.0959 | 2.5583 | 2.5639 | H10 | 4.1586 | 2.6638 | 2.1550 | 1.0867 | 5.1460 | 4.6821 | 4.6414 | 2.4540 | 3.0959 | | 1.7632 | 1.7635 | H11 | 4.5203 | 3.2176 | 2.1460 | 1.0894 | 5.4202 | 5.3119 | 4.6500 | 3.5059 | 2.5583 | 1.7632 | | 1.7522 | H12 | 4.5100 | 3.2177 | 2.1483 | 1.0895 | 5.8695 | 5.0195 | 4.6290 | 3.5087 | 2.5639 | 1.7635 | 1.7522 | |
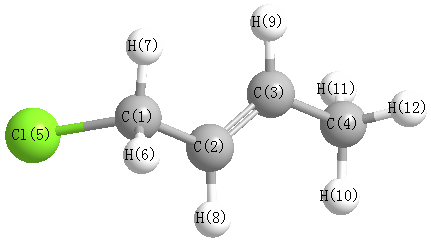 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.499 |
|
C1 |
C2 |
H8 |
116.018 |
| C2 |
C1 |
Cl5 |
111.609 |
|
C2 |
C1 |
H6 |
111.999 |
| C2 |
C1 |
H7 |
112.019 |
|
C2 |
C3 |
C4 |
125.068 |
| C2 |
C3 |
H9 |
119.000 |
|
C3 |
C2 |
H8 |
120.479 |
| C3 |
C4 |
H10 |
111.570 |
|
C3 |
C4 |
H11 |
110.682 |
| C3 |
C4 |
H12 |
110.855 |
|
C4 |
C3 |
H9 |
115.931 |
| Cl5 |
C1 |
H6 |
105.164 |
|
Cl5 |
C1 |
H7 |
106.426 |
| H6 |
C1 |
H7 |
109.253 |
|
H10 |
C4 |
H11 |
108.246 |
| H10 |
C4 |
H12 |
108.263 |
|
H11 |
C4 |
H12 |
107.064 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
