return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 (hex-1-ene)

using model chemistry: CBS-Q

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CBS-Q
 hartrees
Energy at 0K-235.323275
Energy at 298.15K-235.314375
HF Energy-234.172907
Nuclear repulsion energy228.212614
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3388 3388 27.88      
2 A 3314 3314 6.86      
3 A 3297 3297 34.99      
4 A 3251 3251 70.62      
5 A 3251 3251 105.58      
6 A 3231 3231 39.31      
7 A 3210 3210 8.05      
8 A 3196 3196 8.17      
9 A 3183 3183 27.54      
10 A 3179 3179 101.31      
11 A 3172 3172 4.12      
12 A 3168 3168 9.40      
13 A 1874 1874 10.02      
14 A 1647 1647 3.22      
15 A 1636 1636 0.64      
16 A 1635 1635 5.26      
17 A 1625 1625 0.48      
18 A 1621 1621 2.25      
19 A 1583 1583 1.29      
20 A 1555 1555 1.53      
21 A 1543 1543 0.90      
22 A 1484 1484 0.32      
23 A 1450 1450 0.41      
24 A 1433 1433 0.03      
25 A 1429 1429 0.52      
26 A 1384 1384 6.86      
27 A 1357 1357 0.05      
28 A 1297 1297 0.20      
29 A 1218 1218 2.15      
30 A 1143 1143 9.90      
31 A 1131 1131 0.21      
32 A 1119 1119 3.08      
33 A 1091 1091 0.45      
34 A 1056 1056 45.04      
35 A 1011 1011 2.06      
36 A 995 995 0.53      
37 A 961 961 1.06      
38 A 855 855 0.73      
39 A 783 783 1.79      
40 A 702 702 9.15      
41 A 487 487 1.29      
42 A 383 383 0.35      
43 A 378 378 0.15      
44 A 263 263 0.00      
45 A 187 187 0.04      
46 A 138 138 0.04      
47 A 100 100 0.02      
48 A 80 80 0.02      

Unscaled Zero Point Vibrational Energy (zpe) 38735.1 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 38735.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CBS-Q
ABC
0.43591 0.04173 0.04097

See section I.F.4 to change rotational constant units
Geometric Data calculated at CBS-Q

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 3.072 -0.170 -0.437
C2 2.040 -0.211 0.381
C3 0.773 0.585 0.225
C4 -0.472 -0.302 0.089
C5 -1.768 0.502 -0.029
C6 -3.007 -0.383 -0.167
H7 3.080 0.467 -1.308
H8 3.951 -0.768 -0.267
H9 2.085 -0.867 1.240
H10 0.650 1.235 1.092
H11 0.855 1.237 -0.643
H12 -0.358 -0.945 -0.784
H13 -0.536 -0.967 0.951
H14 -1.877 1.143 0.846
H15 -1.703 1.169 -0.889
H16 -3.911 0.216 -0.250
H17 -2.944 -1.012 -1.053
H18 -3.121 -1.038 0.695

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.31712.50843.58494.90336.08841.07871.07722.06663.19052.63363.53323.94725.27814.97916.99596.10616.3551
C21.31711.50462.53073.89615.07982.09562.09351.08162.12852.13302.76522.74514.17054.18625.99985.24855.2368
C32.50841.50461.53422.55503.92122.77203.48922.20451.09141.08852.15272.15662.77832.77634.72224.24284.2448
C43.58492.53071.53421.52962.54923.89304.46172.85992.15102.16011.08981.09082.15272.15243.49442.81472.8157
C54.90333.89612.55501.52961.52835.01395.86344.28102.76442.79302.15622.15301.09011.09002.17302.17342.1736
C66.08845.07983.92122.54921.52836.25066.96935.30444.19224.21522.77712.77402.15192.15151.08761.08841.0885
H71.07872.09562.77203.89305.01396.25061.83503.04273.50102.44643.75284.49815.44654.85197.07486.20836.6877
H81.07722.09353.48924.46175.86346.96931.83502.40084.09433.70794.34344.65406.23376.00877.92356.94447.1425
H92.06661.08162.20452.85994.28105.30443.04272.40082.54983.07963.17252.63874.45994.79766.27215.52885.2371
H103.19052.12851.09142.15102.76444.19223.50104.09432.54981.74733.04732.50562.54033.07594.86214.75094.4210
H112.63362.13301.08852.16012.79304.21522.44643.70793.07961.74732.50053.05573.11282.57064.89034.43484.7727
H123.53322.76522.15271.08982.15622.77713.75284.34343.17253.04732.50051.74373.05282.50713.77572.60143.1352
H133.94722.74512.15661.09082.15302.77404.49814.65402.63872.50563.05571.74372.50253.05073.77233.13312.5986
H145.27814.17052.77832.15271.09012.15195.44656.23374.45992.54033.11283.05282.50251.74302.48933.06422.5155
H154.97914.18622.77632.15241.09002.15154.85196.00874.79763.07592.57062.50713.05071.74302.48852.51523.0640
H166.99595.99984.72223.49442.17301.08767.07487.92356.27214.86214.89033.77573.77232.48932.48851.75721.7572
H176.10615.24854.24282.81472.17341.08846.20836.94445.52884.75094.43482.60143.13313.06422.51521.75721.7566
H186.35515.23684.24482.81572.17361.08856.68777.14255.23714.42104.77273.13522.59862.51553.06401.75721.7566

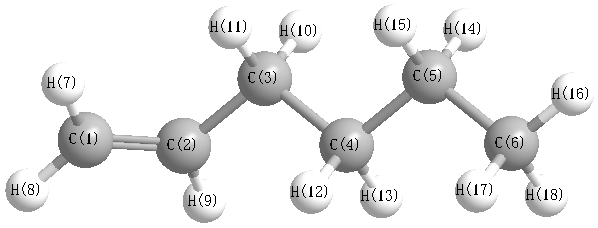
picture of hex-1-ene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 125.354 C1 C2 H9 118.658
C2 C1 H7 121.706 C2 C1 H8 121.619
C2 C3 C4 112.769 C2 C3 H10 109.109
C2 C3 H11 109.641 C3 C2 H9 115.987
C3 C4 C5 113.011 C3 C4 H12 109.075
C3 C4 H13 109.319 C4 C3 H10 108.845
C4 C3 H11 109.724 C4 C5 C6 112.955
C4 C5 H14 109.372 C4 C5 H15 109.353
C5 C4 H12 109.668 C5 C4 H13 109.352
C5 C6 H16 111.225 C5 C6 H17 111.206
C5 C6 H18 111.216 C6 C5 H14 109.400
C6 C5 H15 109.371 H7 C1 H8 116.675
H10 C3 H11 106.555 H12 C4 H13 106.186
H14 C5 H15 106.163 H16 C6 H17 107.714
H16 C6 H18 107.705 H17 C6 H18 107.597
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability