return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12O2 (Methyl pivalate)

using model chemistry: CBS-Q

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at CBS-Q
 hartrees
Energy at 0K-385.659590
Energy at 298.15K-385.648695
HF Energy-383.937417
Nuclear repulsion energy413.298324
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3334 3334 26.75      
2 A 3315 3315 30.80      
3 A 3290 3290 52.78      
4 A 3286 3286 32.45      
5 A 3283 3283 1.78      
6 A 3271 3271 61.05      
7 A 3256 3256 27.19      
8 A 3254 3254 14.16      
9 A 3230 3230 39.64      
10 A 3203 3203 27.81      
11 A 3192 3192 10.94      
12 A 3190 3190 41.89      
13 A 1998 1998 307.34      
14 A 1661 1661 27.60      
15 A 1643 1643 8.88      
16 A 1640 1640 4.94      
17 A 1639 1639 4.07      
18 A 1631 1631 4.61      
19 A 1628 1628 0.00      
20 A 1624 1624 0.88      
21 A 1620 1620 4.92      
22 A 1618 1618 0.01      
23 A 1574 1574 10.56      
24 A 1547 1547 4.23      
25 A 1539 1539 3.02      
26 A 1462 1462 200.75      
27 A 1371 1371 1.54      
28 A 1349 1349 1.08      
29 A 1349 1349 163.93      
30 A 1313 1313 216.28      
31 A 1295 1295 3.39      
32 A 1160 1160 16.47      
33 A 1147 1147 0.80      
34 A 1108 1108 11.20      
35 A 1052 1052 0.01      
36 A 1024 1024 2.06      
37 A 1019 1019 1.84      
38 A 957 957 11.05      
39 A 862 862 12.48      
40 A 861 861 2.18      
41 A 634 634 6.39      
42 A 533 533 3.71      
43 A 406 406 1.71      
44 A 393 393 14.15      
45 A 365 365 0.30      
46 A 344 344 6.83      
47 A 315 315 2.75      
48 A 287 287 0.01      
49 A 280 280 2.84      
50 A 230 230 1.00      
51 A 214 214 2.15      
52 A 181 181 0.08      
53 A 132 132 2.62      
54 A 45 45 1.05      

Unscaled Zero Point Vibrational Energy (zpe) 41073.3 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 41073.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CBS-Q
ABC
0.10498 0.05039 0.04428

See section I.F.4 to change rotational constant units
Geometric Data calculated at CBS-Q

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.907 1.119 -0.004
C2 -0.966 -0.092 -0.000
C3 0.478 0.402 -0.001
C4 -1.202 -0.949 -1.258
C5 -1.202 -0.941 1.263
O6 1.350 -0.598 -0.001
O7 0.813 1.540 -0.001
C8 2.723 -0.256 0.000
H9 -1.750 1.739 -0.882
H10 -1.750 1.744 0.871
H11 -2.941 0.782 -0.003
H12 -0.557 -1.823 -1.271
H13 -1.018 -0.376 -2.166
H14 -2.235 -1.287 -1.284
H15 -0.557 -1.815 1.282
H16 -1.019 -0.363 2.167
H17 -2.235 -1.280 1.290
H18 3.262 -1.195 0.003
H19 2.972 0.322 0.882
H20 2.974 0.318 -0.884

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 O6 O7 C8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20
C11.53352.49012.51992.51993.68202.75174.83021.08611.08601.08823.47602.77482.74533.47612.77462.74575.66285.02235.0237
C21.53351.52641.54051.54062.37112.41383.69332.17792.17792.15982.18602.18462.16482.18602.18472.16504.36944.05724.0590
C32.49011.52642.49582.49501.32731.18542.34012.74332.74283.44032.76362.74423.44422.76252.74303.44373.20942.64702.6491
C42.51991.54052.49582.52162.86633.43994.17952.76903.47662.75681.08541.08871.08802.75973.48002.77024.64434.85974.3797
C52.51991.54062.49502.52162.86843.43674.17993.47672.76892.75702.75963.48002.77041.08541.08871.08804.64484.37764.8615
O63.68202.37111.32732.86632.86842.20411.41513.98093.98204.50802.59863.21613.87012.60053.21963.87152.00242.06322.0631
O72.75172.41381.18543.43993.43672.20412.62222.71722.71473.82973.84663.42174.35053.84343.41634.34813.67072.63132.6349
C84.83023.69332.34014.17954.17991.41512.62224.97684.97715.75913.85134.32455.22503.85134.32555.22521.08191.08371.0837
H91.08612.17792.74332.76903.47673.98092.71724.97681.75281.76283.77592.57993.09074.32803.77473.75065.87395.23604.9328
H101.08602.17792.74283.47662.76893.98202.71474.97711.75281.76294.32803.77493.75013.77572.57953.09105.87434.93175.2373
H111.08822.15983.44032.75682.75704.50803.82975.75911.76281.76293.75183.11732.53363.75223.11722.53436.51035.99705.9985
H123.47602.18602.76361.08542.75962.59863.84663.85133.77594.32803.75181.76201.76162.55303.76383.11014.07454.65774.1478
H132.77482.18462.74421.08873.48003.21613.42174.32452.57993.77493.11731.76201.75783.76384.33273.77404.86685.06914.2497
H142.74532.16483.44421.08802.77043.87014.35055.22503.09073.75012.53361.76161.75783.11083.77392.57415.64635.86505.4658
H153.47612.18602.76252.75971.08542.60053.84343.85134.32803.77573.75222.55303.76383.11081.76211.76164.07474.14514.6592
H162.77462.18472.74303.48001.08873.21963.41634.32553.77472.57953.11723.76384.33273.77391.76211.75794.86804.24825.0709
H172.74572.16503.44372.77021.08803.87154.34815.22523.75063.09102.53433.11013.77402.57411.76161.75795.64655.46375.8668
H185.66284.36943.20944.64434.64482.00243.67071.08195.87395.87436.51034.07454.86685.64634.07474.86805.64651.77721.7772
H195.02234.05722.64704.85974.37762.06322.63131.08375.23604.93175.99704.65775.06915.86504.14514.24825.46371.77721.7659
H205.02374.05902.64914.37974.86152.06312.63491.08374.93285.23735.99854.14784.24975.46584.65925.07095.86681.77721.7659

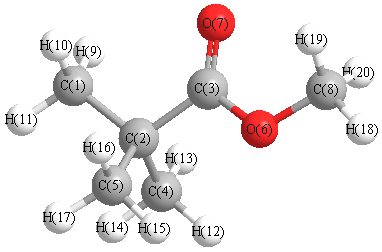
picture of Methyl pivalate state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 108.930 C1 C2 C4 110.119
C1 C2 C5 110.115 C2 C1 H9 111.336
C2 C1 H10 111.339 C2 C1 H11 109.769
C2 C3 O6 112.189 C2 C3 O7 125.303
C2 C4 H12 111.529 C2 C4 H13 111.218
C2 C4 H14 109.691 C2 C5 H15 111.522
C2 C5 H16 111.214 C2 C5 H17 109.696
C3 C2 C4 108.930 C3 C2 C5 108.871
C3 O6 C8 117.113 C4 C2 C5 109.845
O6 C3 O7 122.507 O6 C8 H18 105.862
O6 C8 H19 110.615 O6 C8 H20 110.607
H9 C1 H10 107.600 H9 C1 H11 108.341
H10 C1 H11 108.348 H12 C4 H13 108.275
H12 C4 H14 108.291 H13 C4 H14 107.713
H15 C5 H16 108.281 H15 C5 H17 108.288
H16 C5 H17 107.716 H18 C8 H19 110.305
H18 C8 H20 110.298 H19 C8 H20 109.126
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability