Jump to
S1C2
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -277.950605 |
| Energy at 298.15K | -277.945220 |
| HF Energy | -276.914558 |
| Nuclear repulsion energy | 126.881333 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3246 |
3246 |
0.00 |
194.80 |
0.10 |
0.18 |
| 2 |
Ag |
1664 |
1664 |
0.00 |
18.46 |
0.75 |
0.85 |
| 3 |
Ag |
1608 |
1608 |
0.00 |
2.81 |
0.45 |
0.62 |
| 4 |
Ag |
1183 |
1183 |
0.00 |
11.49 |
0.75 |
0.86 |
| 5 |
Ag |
1160 |
1160 |
0.00 |
8.04 |
0.22 |
0.36 |
| 6 |
Ag |
495 |
495 |
0.00 |
2.67 |
0.48 |
0.65 |
| 7 |
Au |
3318 |
3318 |
87.61 |
0.00 |
0.00 |
0.00 |
| 8 |
Au |
1355 |
1355 |
7.19 |
0.00 |
0.00 |
0.00 |
| 9 |
Au |
871 |
871 |
0.51 |
0.00 |
0.00 |
0.00 |
| 10 |
Au |
143 |
143 |
16.68 |
0.00 |
0.00 |
0.00 |
| 11 |
Bg |
3292 |
3292 |
0.00 |
125.56 |
0.75 |
0.86 |
| 12 |
Bg |
1415 |
1415 |
0.00 |
18.19 |
0.75 |
0.86 |
| 13 |
Bg |
1288 |
1288 |
0.00 |
4.14 |
0.75 |
0.86 |
| 14 |
Bu |
3248 |
3248 |
72.86 |
0.00 |
0.00 |
0.00 |
| 15 |
Bu |
1671 |
1671 |
2.99 |
0.00 |
0.00 |
0.00 |
| 16 |
Bu |
1489 |
1489 |
23.81 |
0.00 |
0.00 |
0.00 |
| 17 |
Bu |
1184 |
1184 |
266.09 |
0.00 |
0.00 |
0.00 |
| 18 |
Bu |
304 |
304 |
23.83 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 14466.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 14466.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
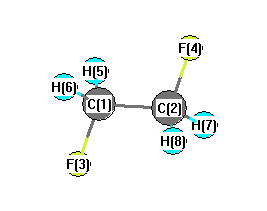
Geometric Data calculated at CBS-Q
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.422 |
0.629 |
0.000 |
| C2 |
-0.422 |
-0.629 |
0.000 |
| F3 |
-0.422 |
1.704 |
0.000 |
| F4 |
0.422 |
-1.704 |
0.000 |
| H5 |
1.047 |
0.676 |
0.885 |
| H6 |
1.047 |
0.676 |
-0.885 |
| H7 |
-1.047 |
-0.676 |
0.885 |
| H8 |
-1.047 |
-0.676 |
-0.885 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5147 | 1.3666 | 2.3336 | 1.0844 | 1.0844 | 2.1548 | 2.1548 |
C2 | 1.5147 | | 2.3336 | 1.3666 | 2.1548 | 2.1548 | 1.0844 | 1.0844 | F3 | 1.3666 | 2.3336 | | 3.5117 | 1.9992 | 1.9992 | 2.6155 | 2.6155 | F4 | 2.3336 | 1.3666 | 3.5117 | | 2.6155 | 2.6155 | 1.9992 | 1.9992 | H5 | 1.0844 | 2.1548 | 1.9992 | 2.6155 | | 1.7695 | 2.4925 | 3.0567 | H6 | 1.0844 | 2.1548 | 1.9992 | 2.6155 | 1.7695 | | 3.0567 | 2.4925 | H7 | 2.1548 | 1.0844 | 2.6155 | 1.9992 | 2.4925 | 3.0567 | | 1.7695 | H8 | 2.1548 | 1.0844 | 2.6155 | 1.9992 | 3.0567 | 2.4925 | 1.7695 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.069 |
|
C1 |
C2 |
H7 |
110.915 |
| C1 |
C2 |
H8 |
110.915 |
|
C2 |
C1 |
F3 |
108.069 |
| C2 |
C1 |
H5 |
110.915 |
|
C2 |
C1 |
H6 |
110.915 |
| F3 |
C1 |
H5 |
108.765 |
|
F3 |
C1 |
H6 |
108.765 |
| F4 |
C2 |
H7 |
108.765 |
|
F4 |
C2 |
H8 |
108.765 |
| H5 |
C1 |
H6 |
109.350 |
|
H7 |
C2 |
H8 |
109.350 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -277.951786 |
| Energy at 298.15K | -277.946493 |
| HF Energy | -276.914080 |
| Nuclear repulsion energy | 129.092948 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3287 |
3287 |
28.06 |
98.08 |
0.73 |
0.85 |
| 2 |
A |
3240 |
3240 |
41.56 |
160.31 |
0.03 |
0.06 |
| 3 |
A |
1645 |
1645 |
0.12 |
2.60 |
0.75 |
0.86 |
| 4 |
A |
1597 |
1597 |
23.54 |
2.74 |
0.38 |
0.55 |
| 5 |
A |
1423 |
1423 |
2.93 |
18.98 |
0.73 |
0.84 |
| 6 |
A |
1239 |
1239 |
26.48 |
1.36 |
0.14 |
0.25 |
| 7 |
A |
1221 |
1221 |
88.67 |
6.01 |
0.72 |
0.84 |
| 8 |
A |
946 |
946 |
32.50 |
7.02 |
0.23 |
0.37 |
| 9 |
A |
345 |
345 |
0.31 |
0.20 |
0.32 |
0.49 |
| 10 |
A |
153 |
153 |
4.70 |
0.03 |
0.75 |
0.86 |
| 11 |
B |
3301 |
3301 |
67.11 |
13.44 |
0.75 |
0.86 |
| 12 |
B |
3225 |
3225 |
21.78 |
41.65 |
0.75 |
0.86 |
| 13 |
B |
1642 |
1642 |
4.67 |
13.61 |
0.75 |
0.86 |
| 14 |
B |
1549 |
1549 |
18.20 |
1.03 |
0.75 |
0.86 |
| 15 |
B |
1382 |
1382 |
7.94 |
5.53 |
0.75 |
0.86 |
| 16 |
B |
1199 |
1199 |
53.72 |
3.81 |
0.75 |
0.86 |
| 17 |
B |
992 |
992 |
53.78 |
3.73 |
0.75 |
0.86 |
| 18 |
B |
537 |
537 |
20.74 |
0.51 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 14461.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 14461.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.262 |
0.706 |
0.509 |
| C2 |
-0.262 |
-0.706 |
0.509 |
| F3 |
-0.262 |
1.399 |
-0.544 |
| F4 |
0.262 |
-1.399 |
-0.544 |
| H5 |
-0.026 |
1.211 |
1.426 |
| H6 |
1.343 |
0.711 |
0.415 |
| H7 |
0.026 |
-1.211 |
1.426 |
| H8 |
-1.343 |
-0.711 |
0.415 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5064 | 1.3644 | 2.3538 | 1.0864 | 1.0848 | 2.1387 | 2.1430 |
C2 | 1.5064 | | 2.3538 | 1.3644 | 2.1387 | 2.1430 | 1.0864 | 1.0848 | F3 | 1.3644 | 2.3538 | | 2.8472 | 1.9929 | 1.9918 | 3.2830 | 2.5579 | F4 | 2.3538 | 1.3644 | 2.8472 | | 3.2830 | 2.5579 | 1.9929 | 1.9918 | H5 | 1.0864 | 2.1387 | 1.9929 | 3.2830 | | 1.7736 | 2.4227 | 2.5397 | H6 | 1.0848 | 2.1430 | 1.9918 | 2.5579 | 1.7736 | | 2.5397 | 3.0388 | H7 | 2.1387 | 1.0864 | 3.2830 | 1.9929 | 2.4227 | 2.5397 | | 1.7736 | H8 | 2.1430 | 1.0848 | 2.5579 | 1.9918 | 2.5397 | 3.0388 | 1.7736 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.053 |
|
C1 |
C2 |
H7 |
110.091 |
| C1 |
C2 |
H8 |
110.528 |
|
C2 |
C1 |
F3 |
110.053 |
| C2 |
C1 |
H5 |
110.091 |
|
C2 |
C1 |
H6 |
110.528 |
| F3 |
C1 |
H5 |
108.281 |
|
F3 |
C1 |
H6 |
108.287 |
| F4 |
C2 |
H7 |
108.281 |
|
F4 |
C2 |
H8 |
108.287 |
| H5 |
C1 |
H6 |
109.548 |
|
H7 |
C2 |
H8 |
109.548 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
