Jump to
S1C2
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -553.408772 |
| Energy at 298.15K | -553.401441 |
| Nuclear repulsion energy | 199.253414 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
Geometric Data calculated at CBS-Q
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -553.410198 |
| Energy at 298.15K | -553.416293 |
| HF Energy | -552.417023 |
| Nuclear repulsion energy | 202.200087 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3414 |
3414 |
13.63 |
70.49 |
0.62 |
0.77 |
| 2 |
A |
3411 |
3411 |
8.42 |
73.14 |
0.69 |
0.81 |
| 3 |
A |
3358 |
3358 |
7.43 |
161.61 |
0.21 |
0.35 |
| 4 |
A |
3355 |
3355 |
9.70 |
89.59 |
0.39 |
0.56 |
| 5 |
A |
3326 |
3326 |
3.12 |
65.61 |
0.32 |
0.48 |
| 6 |
A |
3321 |
3321 |
5.12 |
76.09 |
0.16 |
0.28 |
| 7 |
A |
1828 |
1828 |
23.82 |
24.84 |
0.55 |
0.71 |
| 8 |
A |
1820 |
1820 |
19.76 |
64.26 |
0.09 |
0.17 |
| 9 |
A |
1551 |
1551 |
5.06 |
20.69 |
0.34 |
0.50 |
| 10 |
A |
1544 |
1544 |
5.04 |
35.43 |
0.45 |
0.62 |
| 11 |
A |
1420 |
1420 |
3.52 |
17.69 |
0.24 |
0.38 |
| 12 |
A |
1410 |
1410 |
15.49 |
16.05 |
0.47 |
0.64 |
| 13 |
A |
1152 |
1152 |
5.32 |
5.99 |
0.75 |
0.86 |
| 14 |
A |
1126 |
1126 |
21.46 |
2.08 |
0.74 |
0.85 |
| 15 |
A |
1110 |
1110 |
22.09 |
2.62 |
0.74 |
0.85 |
| 16 |
A |
1098 |
1098 |
20.79 |
1.57 |
0.59 |
0.74 |
| 17 |
A |
1076 |
1076 |
32.66 |
2.11 |
0.74 |
0.85 |
| 18 |
A |
1033 |
1033 |
43.71 |
2.28 |
0.75 |
0.86 |
| 19 |
A |
802 |
802 |
9.40 |
1.79 |
0.54 |
0.70 |
| 20 |
A |
743 |
743 |
1.99 |
5.85 |
0.18 |
0.30 |
| 21 |
A |
690 |
690 |
19.51 |
5.05 |
0.74 |
0.85 |
| 22 |
A |
661 |
661 |
10.59 |
10.07 |
0.69 |
0.82 |
| 23 |
A |
485 |
485 |
0.53 |
6.84 |
0.74 |
0.85 |
| 24 |
A |
397 |
397 |
0.52 |
4.31 |
0.24 |
0.39 |
| 25 |
A |
230 |
230 |
1.70 |
5.11 |
0.68 |
0.81 |
| 26 |
A |
135 |
135 |
4.28 |
6.47 |
0.75 |
0.86 |
| 27 |
A |
89 |
89 |
1.85 |
10.79 |
0.72 |
0.84 |
Unscaled Zero Point Vibrational Energy (zpe) 20291.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 20291.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.012 |
-0.984 |
0.000 |
| C2 |
1.111 |
0.342 |
0.408 |
| C3 |
-1.564 |
-0.180 |
-0.037 |
| C4 |
2.234 |
0.570 |
-0.248 |
| C5 |
-1.817 |
1.113 |
-0.116 |
| H6 |
0.842 |
0.914 |
1.282 |
| H7 |
-2.370 |
-0.896 |
0.013 |
| H8 |
2.924 |
1.328 |
0.088 |
| H9 |
2.510 |
0.010 |
-1.126 |
| H10 |
-1.043 |
1.858 |
-0.188 |
| H11 |
-2.836 |
1.466 |
-0.115 |
Atom - Atom Distances (Å)
| |
S1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| S1 | | 1.7703 | 1.7696 | 2.7234 | 2.7853 | 2.4361 | 2.3833 | 3.7193 | 2.9149 | 3.0377 | 3.7581 |
C2 | 1.7703 | | 2.7623 | 1.3200 | 3.0733 | 1.0787 | 3.7159 | 2.0876 | 2.1019 | 2.7008 | 4.1370 | C3 | 1.7696 | 2.7623 | | 3.8776 | 1.3206 | 2.9542 | 1.0786 | 4.7363 | 4.2214 | 2.1098 | 2.0814 | C4 | 2.7234 | 1.3200 | 3.8776 | | 4.0895 | 2.0966 | 4.8388 | 1.0784 | 1.0776 | 3.5213 | 5.1498 | C5 | 2.7853 | 3.0733 | 1.3206 | 4.0895 | | 3.0105 | 2.0879 | 4.7498 | 4.5783 | 1.0769 | 1.0778 | H6 | 2.4361 | 1.0787 | 2.9542 | 2.0966 | 3.0105 | | 3.8990 | 2.4346 | 3.0653 | 2.5695 | 3.9723 | H7 | 2.3833 | 3.7159 | 1.0786 | 4.8388 | 2.0879 | 3.8990 | | 5.7422 | 5.0920 | 3.0640 | 2.4106 | H8 | 3.7193 | 2.0876 | 4.7363 | 1.0784 | 4.7498 | 2.4346 | 5.7422 | | 1.8398 | 4.0111 | 5.7643 | H9 | 2.9149 | 2.1019 | 4.2214 | 1.0776 | 4.5783 | 3.0653 | 5.0920 | 1.8398 | | 4.1132 | 5.6317 | H10 | 3.0377 | 2.7008 | 2.1098 | 3.5213 | 1.0769 | 2.5695 | 3.0640 | 4.0111 | 4.1132 | | 1.8368 | H11 | 3.7581 | 4.1370 | 2.0814 | 5.1498 | 1.0778 | 3.9723 | 2.4106 | 5.7643 | 5.6317 | 1.8368 | |
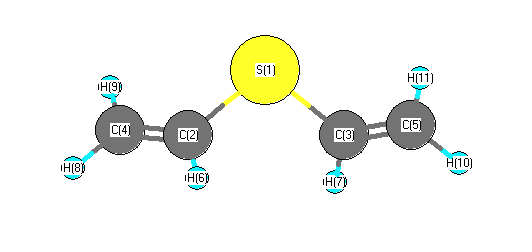 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
C2 |
C4 |
122.931 |
|
S1 |
C2 |
H6 |
115.380 |
| S1 |
C3 |
C5 |
128.078 |
|
S1 |
C3 |
H7 |
111.277 |
| C2 |
S1 |
C3 |
102.582 |
|
C2 |
C4 |
H8 |
120.680 |
| C2 |
C4 |
H9 |
122.163 |
|
C3 |
C5 |
H10 |
122.964 |
| C3 |
C5 |
H11 |
120.080 |
|
C4 |
C2 |
H6 |
121.536 |
| C5 |
C3 |
H7 |
120.645 |
|
H8 |
C4 |
H9 |
117.152 |
| H10 |
C5 |
H11 |
116.956 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
