Jump to
S1C2
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -750.629453 |
| Energy at 298.15K | -750.620476 |
| HF Energy | -747.971852 |
| Nuclear repulsion energy | 569.227652 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
2067 |
2067 |
0.00 |
|
|
|
| 2 |
Ag |
1571 |
1571 |
0.00 |
|
|
|
| 3 |
Ag |
1469 |
1469 |
0.00 |
|
|
|
| 4 |
Ag |
1294 |
1294 |
0.00 |
|
|
|
| 5 |
Ag |
782 |
782 |
0.00 |
|
|
|
| 6 |
Ag |
641 |
641 |
0.00 |
|
|
|
| 7 |
Ag |
408 |
408 |
0.00 |
|
|
|
| 8 |
Ag |
382 |
382 |
0.00 |
|
|
|
| 9 |
Ag |
239 |
239 |
0.00 |
|
|
|
| 10 |
Au |
683 |
683 |
12.03 |
|
|
|
| 11 |
Au |
390 |
390 |
10.75 |
|
|
|
| 12 |
Au |
125 |
125 |
0.24 |
|
|
|
| 13 |
Au |
11i |
11i |
0.15 |
|
|
|
| 14 |
Bg |
753 |
753 |
0.00 |
|
|
|
| 15 |
Bg |
545 |
545 |
0.00 |
|
|
|
| 16 |
Bg |
204 |
204 |
0.00 |
|
|
|
| 17 |
Bu |
1988 |
1988 |
530.08 |
|
|
|
| 18 |
Bu |
1503 |
1503 |
471.66 |
|
|
|
| 19 |
Bu |
1327 |
1327 |
463.85 |
|
|
|
| 20 |
Bu |
1051 |
1051 |
370.85 |
|
|
|
| 21 |
Bu |
684 |
684 |
6.81 |
|
|
|
| 22 |
Bu |
545 |
545 |
4.21 |
|
|
|
| 23 |
Bu |
311 |
311 |
8.32 |
|
|
|
| 24 |
Bu |
147 |
147 |
1.22 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9548.8 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 9548.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.156 |
1.839 |
0.000 |
| C2 |
0.466 |
0.562 |
0.000 |
| C3 |
-0.466 |
-0.562 |
0.000 |
| C4 |
-0.156 |
-1.839 |
0.000 |
| F5 |
1.043 |
2.780 |
0.000 |
| F6 |
-1.043 |
2.318 |
0.000 |
| F7 |
1.750 |
0.234 |
0.000 |
| F8 |
-1.750 |
-0.234 |
0.000 |
| F9 |
1.043 |
-2.318 |
0.000 |
| F10 |
-1.043 |
-2.780 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
F9 |
F10 |
| C1 | | 1.3138 | 2.4809 | 3.6914 | 1.2933 | 1.2916 | 2.2619 | 2.8162 | 4.2505 | 4.7723 |
C2 | 1.3138 | | 1.4613 | 2.4809 | 2.2914 | 2.3153 | 1.3248 | 2.3551 | 2.9374 | 3.6676 | C3 | 2.4809 | 1.4613 | | 1.3138 | 3.6676 | 2.9374 | 2.3551 | 1.3248 | 2.3153 | 2.2914 | C4 | 3.6914 | 2.4809 | 1.3138 | | 4.7723 | 4.2505 | 2.8162 | 2.2619 | 1.2916 | 1.2933 | F5 | 1.2933 | 2.2914 | 3.6676 | 4.7723 | | 2.1374 | 2.6422 | 4.1094 | 5.0978 | 5.9388 | F6 | 1.2916 | 2.3153 | 2.9374 | 4.2505 | 2.1374 | | 3.4849 | 2.6478 | 5.0836 | 5.0978 | F7 | 2.2619 | 1.3248 | 2.3551 | 2.8162 | 2.6422 | 3.4849 | | 3.5309 | 2.6478 | 4.1094 | F8 | 2.8162 | 2.3551 | 1.3248 | 2.2619 | 4.1094 | 2.6478 | 3.5309 | | 3.4849 | 2.6422 | F9 | 4.2505 | 2.9374 | 2.3153 | 1.2916 | 5.0978 | 5.0836 | 2.6478 | 3.4849 | | 2.1374 | F10 | 4.7723 | 3.6676 | 2.2914 | 1.2933 | 5.9388 | 5.0978 | 4.1094 | 2.6422 | 2.1374 | |
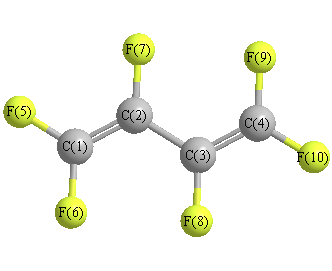 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
126.673 |
|
C1 |
C2 |
F7 |
118.010 |
| C2 |
C1 |
F5 |
123.025 |
|
C2 |
C1 |
F6 |
125.413 |
| C2 |
C3 |
C4 |
126.673 |
|
C2 |
C3 |
F8 |
115.317 |
| C3 |
C2 |
F7 |
115.317 |
|
C3 |
C4 |
F9 |
125.413 |
| C3 |
C4 |
F10 |
123.025 |
|
C4 |
C3 |
F8 |
118.010 |
| F5 |
C1 |
F6 |
111.562 |
|
F9 |
C4 |
F10 |
111.562 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -750.631405 |
| Energy at 298.15K | -750.633621 |
| HF Energy | -747.973094 |
| Nuclear repulsion energy | 572.488168 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
2049 |
2049 |
93.93 |
|
|
|
| 2 |
A |
1541 |
1541 |
0.24 |
|
|
|
| 3 |
A |
1484 |
1484 |
243.42 |
|
|
|
| 4 |
A |
1254 |
1254 |
366.95 |
|
|
|
| 5 |
A |
787 |
787 |
0.04 |
|
|
|
| 6 |
A |
762 |
762 |
2.00 |
|
|
|
| 7 |
A |
602 |
602 |
0.45 |
|
|
|
| 8 |
A |
524 |
524 |
0.94 |
|
|
|
| 9 |
A |
410 |
410 |
1.08 |
|
|
|
| 10 |
A |
279 |
279 |
0.22 |
|
|
|
| 11 |
A |
207 |
207 |
0.50 |
|
|
|
| 12 |
A |
106 |
106 |
0.52 |
|
|
|
| 13 |
A |
47 |
47 |
0.00 |
|
|
|
| 14 |
B |
2024 |
2024 |
315.71 |
|
|
|
| 15 |
B |
1499 |
1499 |
320.88 |
|
|
|
| 16 |
B |
1323 |
1323 |
166.21 |
|
|
|
| 17 |
B |
1073 |
1073 |
242.46 |
|
|
|
| 18 |
B |
726 |
726 |
8.65 |
|
|
|
| 19 |
B |
687 |
687 |
8.40 |
|
|
|
| 20 |
B |
623 |
623 |
5.11 |
|
|
|
| 21 |
B |
465 |
465 |
7.25 |
|
|
|
| 22 |
B |
323 |
323 |
5.73 |
|
|
|
| 23 |
B |
228 |
228 |
5.66 |
|
|
|
| 24 |
B |
121 |
121 |
0.52 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9571.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 9571.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.175 |
1.571 |
-0.360 |
| C2 |
0.162 |
0.712 |
0.570 |
| C3 |
-0.162 |
-0.712 |
0.570 |
| C4 |
0.175 |
-1.571 |
-0.360 |
| F5 |
0.162 |
2.817 |
-0.355 |
| F6 |
-0.847 |
1.264 |
-1.420 |
| F7 |
0.824 |
1.146 |
1.635 |
| F8 |
-0.824 |
-1.146 |
1.635 |
| F9 |
0.847 |
-1.264 |
-1.420 |
| F10 |
-0.162 |
-2.817 |
-0.355 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
F9 |
F10 |
| C1 | | 1.3104 | 2.4650 | 3.1617 | 1.2909 | 1.2925 | 2.2711 | 3.4327 | 3.1943 | 4.3885 |
C2 | 1.3104 | | 1.4598 | 2.4650 | 2.2997 | 2.2987 | 1.3269 | 2.3574 | 2.8865 | 3.6625 | C3 | 2.4650 | 1.4598 | | 1.3104 | 3.6625 | 2.8865 | 2.3574 | 1.3269 | 2.2987 | 2.2997 | C4 | 3.1617 | 2.4650 | 1.3104 | | 4.3885 | 3.1943 | 3.4327 | 2.2711 | 1.2925 | 1.2909 | F5 | 1.2909 | 2.2997 | 3.6625 | 4.3885 | | 2.1372 | 2.6813 | 4.5429 | 4.2729 | 5.6439 | F6 | 1.2925 | 2.2987 | 2.8865 | 3.1943 | 2.1372 | | 3.4842 | 3.8910 | 3.0420 | 4.2729 | F7 | 2.2711 | 1.3269 | 2.3574 | 3.4327 | 2.6813 | 3.4842 | | 2.8233 | 3.8910 | 4.5429 | F8 | 3.4327 | 2.3574 | 1.3269 | 2.2711 | 4.5429 | 3.8910 | 2.8233 | | 3.4842 | 2.6813 | F9 | 3.1943 | 2.8865 | 2.2987 | 1.2925 | 4.2729 | 3.0420 | 3.8910 | 3.4842 | | 2.1372 | F10 | 4.3885 | 3.6625 | 2.2997 | 1.2909 | 5.6439 | 4.2729 | 4.5429 | 2.6813 | 2.1372 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
125.619 |
|
C1 |
C2 |
F7 |
118.889 |
| C2 |
C1 |
F5 |
124.273 |
|
C2 |
C1 |
F6 |
124.045 |
| C2 |
C3 |
C4 |
125.619 |
|
C2 |
C3 |
F8 |
115.465 |
| C3 |
C2 |
F7 |
115.465 |
|
C3 |
C4 |
F9 |
124.045 |
| C3 |
C4 |
F10 |
124.273 |
|
C4 |
C3 |
F8 |
118.889 |
| F5 |
C1 |
F6 |
111.640 |
|
F9 |
C4 |
F10 |
111.640 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
