Jump to
S1C2
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -477.377992 |
| Energy at 298.15K | -477.372317 |
| HF Energy | -476.735582 |
| Nuclear repulsion energy | 107.457127 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3268 |
3268 |
41.42 |
|
|
|
| 2 |
A' |
3232 |
3232 |
24.92 |
|
|
|
| 3 |
A' |
3196 |
3196 |
27.91 |
|
|
|
| 4 |
A' |
2908 |
2908 |
25.93 |
|
|
|
| 5 |
A' |
1639 |
1639 |
1.09 |
|
|
|
| 6 |
A' |
1630 |
1630 |
4.14 |
|
|
|
| 7 |
A' |
1552 |
1552 |
2.40 |
|
|
|
| 8 |
A' |
1439 |
1439 |
49.98 |
|
|
|
| 9 |
A' |
1211 |
1211 |
3.41 |
|
|
|
| 10 |
A' |
1060 |
1060 |
2.97 |
|
|
|
| 11 |
A' |
950 |
950 |
3.96 |
|
|
|
| 12 |
A' |
727 |
727 |
3.41 |
|
|
|
| 13 |
A' |
327 |
327 |
2.70 |
|
|
|
| 14 |
A" |
3293 |
3293 |
41.04 |
|
|
|
| 15 |
A" |
3267 |
3267 |
4.92 |
|
|
|
| 16 |
A" |
1630 |
1630 |
6.19 |
|
|
|
| 17 |
A" |
1384 |
1384 |
0.30 |
|
|
|
| 18 |
A" |
1146 |
1146 |
0.33 |
|
|
|
| 19 |
A" |
849 |
849 |
2.58 |
|
|
|
| 20 |
A" |
273 |
273 |
2.30 |
|
|
|
| 21 |
A" |
203 |
203 |
21.52 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17590.3 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 17590.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.521 |
0.695 |
0.000 |
| C2 |
0.000 |
0.826 |
0.000 |
| S3 |
-0.759 |
-0.838 |
0.000 |
| H4 |
1.977 |
1.683 |
0.000 |
| H5 |
1.869 |
0.162 |
0.881 |
| H6 |
1.869 |
0.162 |
-0.881 |
| H7 |
-0.332 |
1.366 |
0.880 |
| H8 |
-0.332 |
1.366 |
-0.880 |
| H9 |
-2.031 |
-0.460 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5269 | 2.7482 | 1.0878 | 1.0871 | 1.0871 | 2.1585 | 2.1585 | 3.7360 |
C2 | 1.5269 | | 1.8289 | 2.1551 | 2.1706 | 2.1706 | 1.0849 | 1.0849 | 2.4043 | S3 | 2.7482 | 1.8289 | | 3.7208 | 2.9474 | 2.9474 | 2.4118 | 2.4118 | 1.3270 | H4 | 1.0878 | 2.1551 | 3.7208 | | 1.7609 | 1.7609 | 2.4913 | 2.4913 | 4.5456 | H5 | 1.0871 | 2.1706 | 2.9474 | 1.7609 | | 1.7621 | 2.5092 | 3.0655 | 4.0473 | H6 | 1.0871 | 2.1706 | 2.9474 | 1.7609 | 1.7621 | | 3.0655 | 2.5092 | 4.0473 | H7 | 2.1585 | 1.0849 | 2.4118 | 2.4913 | 2.5092 | 3.0655 | | 1.7601 | 2.6457 | H8 | 2.1585 | 1.0849 | 2.4118 | 2.4913 | 3.0655 | 2.5092 | 1.7601 | | 2.6457 | H9 | 3.7360 | 2.4043 | 1.3270 | 4.5456 | 4.0473 | 4.0473 | 2.6457 | 2.6457 | |
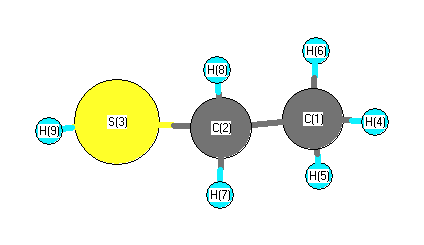 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
109.632 |
|
C1 |
C2 |
H7 |
110.323 |
| C1 |
C2 |
H8 |
110.323 |
|
C2 |
C1 |
H4 |
109.877 |
| C2 |
C1 |
H5 |
111.156 |
|
C2 |
C1 |
H6 |
111.156 |
| C2 |
S3 |
H9 |
97.989 |
|
S3 |
C2 |
H7 |
109.049 |
| S3 |
C2 |
H8 |
109.049 |
|
H4 |
C1 |
H5 |
108.131 |
| H4 |
C1 |
H6 |
108.131 |
|
H5 |
C1 |
H6 |
108.284 |
| H7 |
C2 |
H8 |
108.431 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -477.378438 |
| Energy at 298.15K | -477.385167 |
| HF Energy | -476.736035 |
| Nuclear repulsion energy | 107.193447 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3292 |
3292 |
40.79 |
|
|
|
| 2 |
A |
3269 |
3269 |
8.85 |
|
|
|
| 3 |
A |
3254 |
3254 |
44.77 |
|
|
|
| 4 |
A |
3231 |
3231 |
14.05 |
|
|
|
| 5 |
A |
3189 |
3189 |
32.73 |
|
|
|
| 6 |
A |
2902 |
2902 |
22.40 |
|
|
|
| 7 |
A |
1633 |
1633 |
1.84 |
|
|
|
| 8 |
A |
1628 |
1628 |
6.97 |
|
|
|
| 9 |
A |
1616 |
1616 |
1.48 |
|
|
|
| 10 |
A |
1550 |
1550 |
1.95 |
|
|
|
| 11 |
A |
1443 |
1443 |
26.71 |
|
|
|
| 12 |
A |
1397 |
1397 |
3.61 |
|
|
|
| 13 |
A |
1230 |
1230 |
15.35 |
|
|
|
| 14 |
A |
1156 |
1156 |
0.33 |
|
|
|
| 15 |
A |
1053 |
1053 |
5.42 |
|
|
|
| 16 |
A |
951 |
951 |
9.71 |
|
|
|
| 17 |
A |
792 |
792 |
1.12 |
|
|
|
| 18 |
A |
716 |
716 |
6.68 |
|
|
|
| 19 |
A |
351 |
351 |
1.81 |
|
|
|
| 20 |
A |
276 |
276 |
2.89 |
|
|
|
| 21 |
A |
229 |
229 |
20.29 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17579.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 17579.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.641 |
-0.352 |
-0.051 |
| C2 |
0.498 |
0.645 |
0.088 |
| S3 |
-1.164 |
-0.100 |
-0.080 |
| H4 |
2.597 |
0.164 |
0.023 |
| H5 |
1.608 |
-1.103 |
0.735 |
| H6 |
1.601 |
-0.864 |
-1.007 |
| H7 |
0.546 |
1.170 |
1.035 |
| H8 |
0.542 |
1.393 |
-0.697 |
| H9 |
-1.110 |
-0.913 |
0.965 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5239 | 2.8163 | 1.0884 | 1.0873 | 1.0856 | 2.1671 | 2.1614 | 2.9858 |
C2 | 1.5239 | | 1.8283 | 2.1544 | 2.1697 | 2.1665 | 1.0841 | 1.0852 | 2.4040 | S3 | 2.8163 | 1.8283 | | 3.7707 | 3.0579 | 3.0139 | 2.4044 | 2.3492 | 1.3247 | H4 | 1.0884 | 2.1544 | 3.7707 | | 1.7578 | 1.7638 | 2.4979 | 2.5000 | 3.9729 | H5 | 1.0873 | 2.1697 | 3.0579 | 1.7578 | | 1.7585 | 2.5270 | 3.0687 | 2.7342 | H6 | 1.0856 | 2.1665 | 3.0139 | 1.7638 | 1.7585 | | 3.0694 | 2.5120 | 3.3524 | H7 | 2.1671 | 1.0841 | 2.4044 | 2.4979 | 2.5270 | 3.0694 | | 1.7466 | 2.6621 | H8 | 2.1614 | 1.0852 | 2.3492 | 2.5000 | 3.0687 | 2.5120 | 1.7466 | | 3.2874 | H9 | 2.9858 | 2.4040 | 1.3247 | 3.9729 | 2.7342 | 3.3524 | 2.6621 | 3.2874 | |
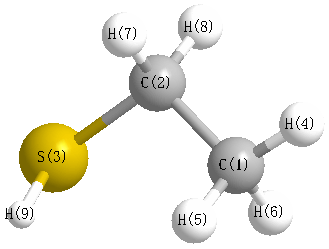 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
114.001 |
|
C1 |
C2 |
H7 |
111.277 |
| C1 |
C2 |
H8 |
110.749 |
|
C2 |
C1 |
H4 |
110.000 |
| C2 |
C1 |
H5 |
111.290 |
|
C2 |
C1 |
H6 |
111.142 |
| C2 |
S3 |
H9 |
98.093 |
|
S3 |
C2 |
H7 |
108.582 |
| S3 |
C2 |
H8 |
104.569 |
|
H4 |
C1 |
H5 |
107.789 |
| H4 |
C1 |
H6 |
108.454 |
|
H5 |
C1 |
H6 |
108.051 |
| H7 |
C2 |
H8 |
107.250 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
