return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H6O (Propylene oxide)

using model chemistry: CBS-Q

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at CBS-Q
 hartrees
Energy at 0K-192.770598
Energy at 298.15K-192.765229
Nuclear repulsion energy125.883859
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3363 3363 41.22      
2 A 3297 3297 48.26      
3 A 3276 3276 41.42      
4 A 3270 3270 3.10      
5 A 3260 3260 33.01      
6 A 3193 3193 23.70      
7 A 1686 1686 9.23      
8 A 1632 1632 4.43      
9 A 1619 1619 2.92      
10 A 1587 1587 17.12      
11 A 1534 1534 4.70      
12 A 1411 1411 7.23      
13 A 1310 1310 7.37      
14 A 1287 1287 1.29      
15 A 1269 1269 5.31      
16 A 1240 1240 9.29      
17 A 1141 1141 9.32      
18 A 1074 1074 21.67      
19 A 986 986 2.87      
20 A 947 947 49.54      
21 A 854 854 6.55      
22 A 438 438 5.83      
23 A 395 395 4.18      
24 A 220 220 0.67      

Unscaled Zero Point Vibrational Energy (zpe) 20143.6 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 20143.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CBS-Q
See section I.F.4 to change rotational constant units
Geometric Data calculated at CBS-Q Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 0.810 -0.767 -0.255
C2 -1.505 0.107 -0.143
H3 -1.417 0.352 -1.198
H4 -2.072 -0.818 -0.055
H5 -2.066 0.896 0.354
C6 -0.143 -0.050 0.483
H7 -0.143 -0.297 1.536
C8 1.044 0.601 -0.049
H9 0.967 1.218 -0.933
H10 1.876 0.832 0.600

Atom - Atom Distances (Å)
  O1 C2 H3 H4 H5 C6 H7 C8 H9 H10
O12.47702.66432.88943.37731.40272.08271.40302.10272.1030
C22.47701.08661.08781.08841.50692.19972.59832.82323.5368
H32.66431.08661.76141.76782.14663.08532.72712.55003.7821
H42.88941.08781.76141.76212.14442.55443.42373.76144.3284
H53.37731.08841.76781.76212.14652.55343.14943.31013.9497
C61.40271.50692.14662.14442.14651.08201.45472.20092.2063
H72.08272.19973.08532.55442.55341.08202.17453.10222.4951
C81.40302.59832.72713.42373.14941.45472.17451.08041.0801
H92.10272.82322.55003.76143.31012.20093.10221.08041.8235
H102.10303.53683.78214.32843.94972.20632.49511.08011.8235

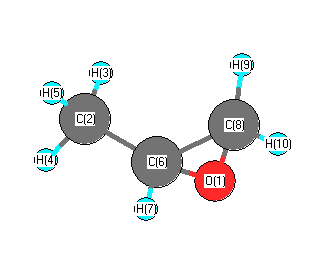
picture of Propylene oxide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C6 C2 116.668 O1 C6 H7 113.272
O1 C6 C8 58.777 O1 C8 C6 58.760
O1 C8 H9 115.096 O1 C8 H10 115.142
C2 C6 H7 115.371 C2 C6 C8 122.628
H3 C2 H4 108.203 H3 C2 H5 108.745
H3 C2 C6 110.680 H4 C2 H5 108.140
H4 C2 C6 110.432 H5 C2 C6 110.561
C6 O1 C8 62.464 C6 C8 H9 119.766
C6 C8 H10 120.287 H7 C6 C8 117.251
H9 C8 H10 115.133
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability