Jump to
S1C2
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -1071.891069 |
| Energy at 298.15K | -1071.884662 |
| HF Energy | -1070.703461 |
| Nuclear repulsion energy | 264.590906 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3372 |
3372 |
1.63 |
|
|
|
| 2 |
A' |
3211 |
3211 |
40.33 |
|
|
|
| 3 |
A' |
2043 |
2043 |
236.67 |
|
|
|
| 4 |
A' |
1528 |
1528 |
9.47 |
|
|
|
| 5 |
A' |
1348 |
1348 |
15.14 |
|
|
|
| 6 |
A' |
1160 |
1160 |
25.07 |
|
|
|
| 7 |
A' |
860 |
860 |
37.49 |
|
|
|
| 8 |
A' |
473 |
473 |
6.66 |
|
|
|
| 9 |
A' |
356 |
356 |
30.18 |
|
|
|
| 10 |
A' |
278 |
278 |
2.29 |
|
|
|
| 11 |
A" |
1376 |
1376 |
40.34 |
|
|
|
| 12 |
A" |
1134 |
1134 |
17.66 |
|
|
|
| 13 |
A" |
821 |
821 |
139.33 |
|
|
|
| 14 |
A" |
307 |
307 |
3.14 |
|
|
|
| 15 |
A" |
87 |
87 |
11.97 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9177.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 9177.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.379 |
-0.012 |
0.000 |
| C2 |
-0.170 |
1.409 |
0.000 |
| H3 |
1.457 |
0.006 |
0.000 |
| Cl4 |
-0.170 |
-0.843 |
1.468 |
| Cl5 |
-0.170 |
-0.843 |
-1.468 |
| O6 |
0.541 |
2.347 |
0.000 |
| H7 |
-1.261 |
1.484 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5239 | 1.0776 | 1.7738 | 1.7738 | 2.3649 | 2.2200 |
C2 | 1.5239 | | 2.1484 | 2.6882 | 2.6882 | 1.1769 | 1.0932 | H3 | 1.0776 | 2.1484 | | 2.3497 | 2.3497 | 2.5140 | 3.0934 | Cl4 | 1.7738 | 2.6882 | 2.3497 | | 2.9360 | 3.5827 | 2.9596 | Cl5 | 1.7738 | 2.6882 | 2.3497 | 2.9360 | | 3.5827 | 2.9596 | O6 | 2.3649 | 1.1769 | 2.5140 | 3.5827 | 3.5827 | | 1.9976 | H7 | 2.2200 | 1.0932 | 3.0934 | 2.9596 | 2.9596 | 1.9976 | |
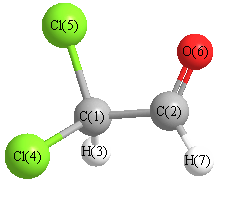 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
121.710 |
|
C1 |
C2 |
H7 |
115.060 |
| C2 |
C1 |
H3 |
110.168 |
|
C2 |
C1 |
Cl4 |
108.973 |
| C2 |
C1 |
Cl5 |
108.973 |
|
H3 |
C1 |
Cl4 |
108.507 |
| H3 |
C1 |
Cl5 |
108.507 |
|
Cl4 |
C1 |
Cl5 |
111.706 |
| O6 |
C2 |
H7 |
123.230 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CBS-Q
| | hartrees |
|---|
| Energy at 0K | -1071.888070 |
| Energy at 298.15K | -1071.881710 |
| HF Energy | -1070.700864 |
| Nuclear repulsion energy | 267.219161 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3316 |
3316 |
5.81 |
77.01 |
0.27 |
0.42 |
| 2 |
A |
3192 |
3192 |
49.34 |
111.80 |
0.33 |
0.49 |
| 3 |
A |
2054 |
2054 |
201.67 |
15.30 |
0.51 |
0.67 |
| 4 |
A |
1527 |
1527 |
25.97 |
3.48 |
0.63 |
0.78 |
| 5 |
A |
1414 |
1414 |
10.38 |
4.25 |
0.74 |
0.85 |
| 6 |
A |
1362 |
1362 |
23.79 |
9.47 |
0.73 |
0.84 |
| 7 |
A |
1144 |
1144 |
9.10 |
3.19 |
0.53 |
0.69 |
| 8 |
A |
1030 |
1030 |
10.99 |
6.92 |
0.62 |
0.77 |
| 9 |
A |
907 |
907 |
99.84 |
11.69 |
0.74 |
0.85 |
| 10 |
A |
700 |
700 |
51.59 |
13.05 |
0.13 |
0.23 |
| 11 |
A |
664 |
664 |
35.20 |
7.58 |
0.43 |
0.60 |
| 12 |
A |
378 |
378 |
1.12 |
3.40 |
0.23 |
0.38 |
| 13 |
A |
299 |
299 |
4.26 |
4.58 |
0.65 |
0.79 |
| 14 |
A |
240 |
240 |
3.67 |
1.33 |
0.71 |
0.83 |
| 15 |
A |
91 |
91 |
15.48 |
2.14 |
0.74 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 9158.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 9158.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CBS-Q
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.093 |
-0.039 |
0.505 |
| C2 |
-0.754 |
-1.269 |
0.169 |
| H3 |
0.208 |
0.029 |
1.578 |
| Cl4 |
1.727 |
-0.293 |
-0.160 |
| Cl5 |
-0.630 |
1.461 |
-0.057 |
| O6 |
-1.830 |
-1.229 |
-0.297 |
| H7 |
-0.255 |
-2.208 |
0.429 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5304 | 1.0811 | 1.7815 | 1.7579 | 2.3990 | 2.1982 |
C2 | 1.5304 | | 2.1428 | 2.6855 | 2.7423 | 1.1732 | 1.0948 | H3 | 1.0811 | 2.1428 | | 2.3303 | 2.3291 | 3.0405 | 2.5568 | Cl4 | 1.7815 | 2.6855 | 2.3303 | | 2.9402 | 3.6798 | 2.8181 | Cl5 | 1.7579 | 2.7423 | 2.3291 | 2.9402 | | 2.9550 | 3.7206 | O6 | 2.3990 | 1.1732 | 3.0405 | 3.6798 | 2.9550 | | 1.9914 | H7 | 2.1982 | 1.0948 | 2.5568 | 2.8181 | 3.7206 | 1.9914 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
124.552 |
|
C1 |
C2 |
H7 |
112.664 |
| C2 |
C1 |
H3 |
109.059 |
|
C2 |
C1 |
Cl4 |
108.118 |
| C2 |
C1 |
Cl5 |
112.829 |
|
H3 |
C1 |
Cl4 |
106.398 |
| H3 |
C1 |
Cl5 |
107.862 |
|
Cl4 |
C1 |
Cl5 |
112.340 |
| O6 |
C2 |
H7 |
122.775 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
