return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C9H8 (Indene)

using model chemistry: CBS-Q

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CBS-Q
 hartrees
Energy at 0K-347.107666
Energy at 298.15K-347.100266
HF Energy-345.457304
Nuclear repulsion energy396.209785
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3398 3398 24.91      
2 A' 3372 3372 29.97      
3 A' 3371 3371 12.57      
4 A' 3357 3357 42.86      
5 A' 3344 3344 4.72      
6 A' 3336 3336 3.73      
7 A' 3189 3189 28.73      
8 A' 1819 1819 3.42      
9 A' 1799 1799 0.50      
10 A' 1765 1765 0.28      
11 A' 1629 1629 16.60      
12 A' 1617 1617 8.31      
13 A' 1589 1589 1.94      
14 A' 1498 1498 3.53      
15 A' 1448 1448 0.13      
16 A' 1372 1372 1.20      
17 A' 1348 1348 1.65      
18 A' 1308 1308 0.73      
19 A' 1272 1272 1.78      
20 A' 1223 1223 0.49      
21 A' 1218 1218 0.96      
22 A' 1172 1172 0.74      
23 A' 1115 1115 2.98      
24 A' 1016 1016 8.65      
25 A' 928 928 2.58      
26 A' 894 894 2.16      
27 A' 788 788 3.03      
28 A' 643 643 1.45      
29 A' 575 575 0.17      
30 A' 410 410 1.19      
31 A" 3219 3219 22.86      
32 A" 1264 1264 1.89      
33 A" 1092 1092 0.09      
34 A" 1077 1077 0.07      
35 A" 1057 1057 1.76      
36 A" 1037 1037 7.90      
37 A" 969 969 0.07      
38 A" 862 862 64.54      
39 A" 805 805 11.70      
40 A" 775 775 13.54      
41 A" 607 607 4.36      
42 A" 467 467 7.13      
43 A" 429 429 2.75      
44 A" 229 229 2.82      
45 A" 211 211 0.74      

Unscaled Zero Point Vibrational Energy (zpe) 32956.4 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 32956.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CBS-Q
ABC
0.12511 0.05240 0.03719

See section I.F.4 to change rotational constant units
Geometric Data calculated at CBS-Q

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.271 0.000 0.000
C2 -1.857 -1.322 0.000
C3 -0.507 -1.641 0.000
C4 -1.341 1.032 0.000
C5 0.000 0.719 0.000
C6 0.418 -0.613 0.000
C7 1.888 -0.632 0.000
C8 2.350 0.612 0.000
C9 1.218 1.610 0.000
H10 -3.323 0.229 0.000
H11 -2.591 -2.109 0.000
H12 -0.189 -2.670 0.000
H13 -1.670 2.058 0.000
H14 2.485 -1.526 0.000
H15 3.386 0.897 0.000
H16 1.253 2.261 0.873
H17 1.253 2.261 -0.873

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 C9 H10 H11 H12 H13 H14 H15 H16 H17
C11.38562.40991.38962.38222.75794.20744.66193.84311.07642.13333.38612.14404.99495.72814.27734.2773
C21.38561.38722.41012.75902.38223.80824.63014.24882.13441.07662.14433.38524.34605.69304.82404.8240
C32.40991.38722.80052.41361.38242.59903.63823.68043.38072.13641.07703.87792.99344.64704.36884.3688
C41.38962.41012.80051.37702.40823.63283.71512.62362.13893.38103.87751.07754.60234.72903.00013.0001
C52.38222.75902.41361.37701.39592.32172.35281.50963.35903.83553.39442.14103.34883.39092.17072.1707
C62.75792.38221.38242.40821.39591.47092.28812.36303.83433.36012.14493.39052.25983.33053.11813.1181
C74.20743.80822.59903.63282.32171.47091.32632.33985.28214.71682.91084.46101.07522.14003.08813.0881
C84.66194.63013.63823.71512.35282.28811.32631.50955.68645.64094.14984.27302.14221.07442.16542.1654
C93.84314.24883.68042.62361.50962.36302.33981.50954.74695.32394.50592.92323.38252.28221.09001.0900
H101.07642.13443.38072.13893.35903.83435.28215.68644.74692.44994.26912.46546.06726.74255.08265.0826
H112.13331.07662.13643.38103.83553.36014.71685.64095.32392.44992.46644.26785.10906.69055.88555.8855
H123.38612.14431.07703.87753.39442.14492.91084.14984.50594.26912.46644.95502.90835.05065.21185.2118
H132.14403.38523.87791.07752.14103.39054.46104.27302.92322.46544.26784.95505.48755.18823.05773.0577
H144.99494.34602.99344.60233.34882.25981.07522.14223.38256.06725.10902.90835.48752.58554.07764.0776
H155.72815.69304.64704.72903.39093.33052.14001.07442.28226.74256.69055.05065.18822.58552.67882.6788
H164.27734.82404.36883.00012.17073.11813.08812.16541.09005.08265.88555.21183.05774.07762.67881.7470
H174.27734.82404.36883.00012.17073.11813.08812.16541.09005.08265.88555.21183.05774.07762.67881.7470

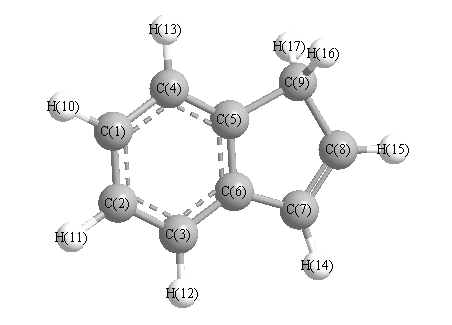
picture of Indene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.711 C1 C2 H11 119.572
C1 C4 C5 118.873 C1 C4 H13 120.164
C2 C1 C4 120.556 C2 C1 H10 119.682
C2 C3 C6 118.661 C2 C3 H12 120.430
C3 C2 H11 119.718 C3 C6 C5 120.630
C3 C6 C7 131.234 C4 C1 H10 119.762
C4 C5 C6 120.569 C4 C5 C9 130.655
C5 C4 H13 120.964 C5 C6 C7 108.136
C5 C9 C8 102.398 C5 C9 H16 112.225
C5 C9 H17 112.225 C6 C3 H12 120.909
C6 C5 C9 108.776 C6 C7 C8 109.658
C6 C7 H14 124.399 C7 C8 C9 111.033
C7 C8 H15 125.774 C8 C7 H14 125.943
C8 C9 H16 111.803 C8 C9 H17 111.803
C9 C8 H15 123.193 H16 C9 H17 106.519
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability