return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10O (2-methylbutyraldehyde)

using model chemistry: CBS-Q

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at CBS-Q
 hartrees
Energy at 0K-271.260363
Energy at 298.15K-271.251777
HF Energy-270.018958
Nuclear repulsion energy240.521205
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CBS-Q
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3286 3286 28.90      
2 A 3277 3277 36.41      
3 A 3267 3267 59.61      
4 A 3255 3255 52.57      
5 A 3214 3214 28.90      
6 A 3206 3206 15.36      
7 A 3190 3190 22.38      
8 A 3179 3179 52.29      
9 A 3154 3154 29.63      
10 A 3107 3107 98.20      
11 A 2020 2020 235.85      
12 A 1646 1646 2.16      
13 A 1642 1642 6.86      
14 A 1634 1634 3.64      
15 A 1631 1631 6.29      
16 A 1622 1622 0.62      
17 A 1569 1569 9.74      
18 A 1556 1556 2.04      
19 A 1544 1544 5.38      
20 A 1516 1516 2.01      
21 A 1492 1492 2.38      
22 A 1434 1434 0.70      
23 A 1396 1396 0.44      
24 A 1298 1298 0.77      
25 A 1263 1263 3.98      
26 A 1218 1218 8.24      
27 A 1121 1121 0.89      
28 A 1091 1091 8.70      
29 A 1059 1059 3.49      
30 A 1016 1016 1.49      
31 A 972 972 12.53      
32 A 849 849 5.26      
33 A 828 828 3.99      
34 A 711 711 14.25      
35 A 431 431 0.81      
36 A 414 414 0.93      
37 A 319 319 6.28      
38 A 281 281 0.81      
39 A 253 253 3.47      
40 A 223 223 0.40      
41 A 102 102 3.93      
42 A 82 82 5.77      

Unscaled Zero Point Vibrational Energy (zpe) 33182.5 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 33182.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CBS-Q
ABC
0.23315 0.07006 0.05861

See section I.F.4 to change rotational constant units
Geometric Data calculated at CBS-Q

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.378 -0.697 0.213
C2 -0.090 0.080 0.399
C3 1.054 -0.682 -0.303
C4 2.452 -0.145 0.013
C5 -0.244 1.532 -0.052
O6 -2.377 -0.264 -0.254
H7 -1.337 -1.745 0.545
H8 0.111 0.053 1.475
H9 1.011 -1.732 -0.011
H10 0.888 -0.657 -1.380
H11 3.213 -0.765 -0.456
H12 2.590 0.871 -0.349
H13 2.640 -0.145 1.086
H14 -0.412 1.589 -1.125
H15 -1.094 2.000 0.436
H16 0.643 2.113 0.186

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 O6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
C11.51572.48663.87562.51521.18441.10052.09102.61402.77084.64074.30354.14892.81912.72043.4614
C21.51571.54332.58141.52902.40332.21491.09432.15942.16003.51532.89272.82402.16902.16682.1719
C32.48661.54331.53102.57863.45682.75062.14221.09061.09042.16632.18432.17542.82483.51402.8665
C43.87562.58141.53103.17594.83854.14762.76712.14382.15671.08801.08691.08923.53634.16562.8982
C52.51521.52902.57863.17592.79633.50622.15543.49742.79924.17032.92493.52511.08751.08591.0865
O61.18442.40333.45684.83852.79631.97843.04643.70083.47615.61655.09595.19442.83782.69223.8687
H71.10052.21492.75064.14763.50621.97842.48892.41333.13714.76094.80224.32053.84173.75434.3513
H82.09101.09432.14222.76712.15543.04642.48892.49093.04273.74453.18422.56623.06442.51392.4873
H92.61402.15941.09062.14383.49743.70082.41332.49091.74522.44563.06242.52443.78044.30763.8672
H102.77082.16001.09042.15672.79923.47613.13713.04271.74522.50462.50853.06822.60683.77893.1910
H114.64073.51532.16631.08804.17035.61654.76093.74452.44562.50461.75371.75824.37375.19513.9113
H124.30352.89272.18431.08692.92495.09594.80223.18423.06242.50851.75371.75923.18263.93182.3703
H134.14892.82402.17541.08923.52515.19444.32052.56622.52443.06821.75821.75924.14864.35483.1459
H142.81912.16902.82483.53631.08752.83783.84173.06443.78042.60684.37373.18264.14861.75231.7628
H152.72042.16683.51404.16561.08592.69223.75432.51394.30763.77895.19513.93184.35481.75231.7586
H163.46142.17192.86652.89821.08653.86874.35132.48733.86723.19103.91132.37033.14591.76281.7586

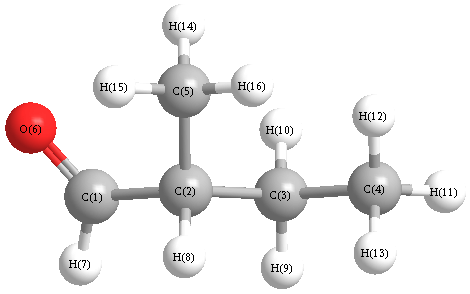
picture of 2-methylbutyraldehyde state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 108.757 C1 C2 C5 111.402
C1 C2 H8 105.337 C2 C1 O6 125.323
C2 C1 H7 114.760 C2 C3 C4 114.214
C2 C3 H9 108.929 C2 C3 H10 108.985
C2 C5 H14 110.860 C2 C5 H15 110.780
C2 C5 H16 111.146 C3 C2 C5 114.132
C3 C2 H8 107.395 C3 C4 H11 110.470
C3 C4 H12 111.982 C3 C4 H13 111.126
C4 C3 H9 108.547 C4 C3 H10 109.575
C5 C2 H8 109.375 O6 C1 H7 119.916
H9 C3 H10 106.291 H11 C4 H12 107.476
H11 C4 H13 107.717 H12 C4 H13 107.889
H14 C5 H15 107.465 H14 C5 H16 108.363
H15 C5 H16 108.094
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability