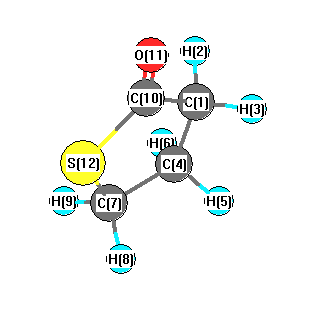
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3153 |
3017 |
10.30 |
|
|
|
| 2 |
A |
3142 |
3006 |
7.56 |
|
|
|
| 3 |
A |
3125 |
2990 |
16.95 |
|
|
|
| 4 |
A |
3087 |
2953 |
18.91 |
|
|
|
| 5 |
A |
3076 |
2943 |
22.97 |
|
|
|
| 6 |
A |
3057 |
2925 |
6.65 |
|
|
|
| 7 |
A |
1846 |
1766 |
356.82 |
|
|
|
| 8 |
A |
1523 |
1457 |
0.37 |
|
|
|
| 9 |
A |
1508 |
1442 |
6.55 |
|
|
|
| 10 |
A |
1481 |
1416 |
7.73 |
|
|
|
| 11 |
A |
1375 |
1315 |
0.44 |
|
|
|
| 12 |
A |
1327 |
1270 |
6.03 |
|
|
|
| 13 |
A |
1321 |
1264 |
6.35 |
|
|
|
| 14 |
A |
1260 |
1205 |
2.44 |
|
|
|
| 15 |
A |
1202 |
1150 |
1.55 |
|
|
|
| 16 |
A |
1129 |
1080 |
7.84 |
|
|
|
| 17 |
A |
1081 |
1035 |
58.34 |
|
|
|
| 18 |
A |
1044 |
998 |
4.82 |
|
|
|
| 19 |
A |
1041 |
996 |
49.63 |
|
|
|
| 20 |
A |
932 |
892 |
9.54 |
|
|
|
| 21 |
A |
880 |
842 |
8.04 |
|
|
|
| 22 |
A |
846 |
810 |
15.02 |
|
|
|
| 23 |
A |
706 |
675 |
2.22 |
|
|
|
| 24 |
A |
632 |
604 |
37.44 |
|
|
|
| 25 |
A |
602 |
576 |
11.38 |
|
|
|
| 26 |
A |
483 |
462 |
0.97 |
|
|
|
| 27 |
A |
477 |
456 |
1.96 |
|
|
|
| 28 |
A |
401 |
384 |
3.17 |
|
|
|
| 29 |
A |
228 |
218 |
1.50 |
|
|
|
| 30 |
A |
131 |
125 |
0.62 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21047.2 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 20135.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.385 |
|
|
-0.180 |
| 2 |
H |
0.214 |
|
|
0.095 |
| 3 |
H |
0.209 |
|
|
0.104 |
| 4 |
C |
-0.347 |
|
|
-0.101 |
| 5 |
H |
0.187 |
|
|
0.070 |
| 6 |
H |
0.196 |
|
|
0.080 |
| 7 |
C |
-0.496 |
|
|
-0.193 |
| 8 |
H |
0.209 |
|
|
0.124 |
| 9 |
H |
0.213 |
|
|
0.143 |
| 10 |
C |
0.268 |
|
|
0.442 |
| 11 |
O |
-0.423 |
|
|
-0.399 |
| 12 |
S |
0.154 |
|
|
-0.185 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.694 |
1.860 |
0.390 |
4.154 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
3.694 |
1.891 |
0.423 |
4.171 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.275 |
-0.345 |
-0.035 |
| y |
-0.345 |
-42.041 |
0.147 |
| z |
-0.035 |
0.147 |
-41.730 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.390 |
-0.345 |
-0.035 |
| y |
-0.345 |
1.961 |
0.147 |
| z |
-0.035 |
0.147 |
2.428 |
|
| Polar |
|---|
| 3z2-r2 | 4.856 |
|---|
| x2-y2 | -4.234 |
|---|
| xy | -0.345 |
|---|
| xz | -0.035 |
|---|
| yz | 0.147 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
178.617 |
| (<r2>)1/2 |
13.365 |