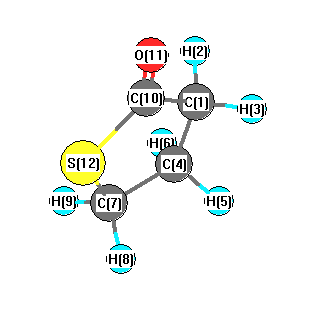
Vibrational Frequencies calculated at PBE1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3171 |
3014 |
8.36 |
|
|
|
| 2 |
A |
3161 |
3004 |
5.96 |
|
|
|
| 3 |
A |
3145 |
2989 |
14.88 |
|
|
|
| 4 |
A |
3103 |
2949 |
17.34 |
|
|
|
| 5 |
A |
3093 |
2939 |
20.78 |
|
|
|
| 6 |
A |
3074 |
2921 |
5.49 |
|
|
|
| 7 |
A |
1867 |
1774 |
360.18 |
|
|
|
| 8 |
A |
1525 |
1449 |
0.53 |
|
|
|
| 9 |
A |
1509 |
1434 |
7.36 |
|
|
|
| 10 |
A |
1481 |
1408 |
8.56 |
|
|
|
| 11 |
A |
1379 |
1311 |
0.35 |
|
|
|
| 12 |
A |
1332 |
1266 |
6.40 |
|
|
|
| 13 |
A |
1326 |
1260 |
5.81 |
|
|
|
| 14 |
A |
1265 |
1202 |
2.60 |
|
|
|
| 15 |
A |
1209 |
1148 |
1.26 |
|
|
|
| 16 |
A |
1133 |
1077 |
11.33 |
|
|
|
| 17 |
A |
1093 |
1039 |
64.57 |
|
|
|
| 18 |
A |
1050 |
998 |
40.86 |
|
|
|
| 19 |
A |
1047 |
995 |
6.57 |
|
|
|
| 20 |
A |
941 |
895 |
9.04 |
|
|
|
| 21 |
A |
884 |
840 |
9.24 |
|
|
|
| 22 |
A |
851 |
809 |
12.82 |
|
|
|
| 23 |
A |
718 |
683 |
2.08 |
|
|
|
| 24 |
A |
641 |
609 |
38.62 |
|
|
|
| 25 |
A |
606 |
576 |
10.20 |
|
|
|
| 26 |
A |
486 |
462 |
0.78 |
|
|
|
| 27 |
A |
480 |
456 |
2.05 |
|
|
|
| 28 |
A |
406 |
386 |
3.08 |
|
|
|
| 29 |
A |
231 |
220 |
1.51 |
|
|
|
| 30 |
A |
132 |
125 |
0.62 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21168.3 cm
-1
Scaled (by 0.9503) Zero Point Vibrational Energy (zpe) 20116.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.396 |
|
|
|
| 2 |
H |
0.219 |
|
|
|
| 3 |
H |
0.214 |
|
|
|
| 4 |
C |
-0.360 |
|
|
|
| 5 |
H |
0.193 |
|
|
|
| 6 |
H |
0.201 |
|
|
|
| 7 |
C |
-0.511 |
|
|
|
| 8 |
H |
0.214 |
|
|
|
| 9 |
H |
0.219 |
|
|
|
| 10 |
C |
0.265 |
|
|
|
| 11 |
O |
-0.422 |
|
|
|
| 12 |
S |
0.163 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.699 |
1.861 |
0.394 |
4.159 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.353 |
-0.358 |
-0.028 |
| y |
-0.358 |
-42.150 |
0.148 |
| z |
-0.028 |
0.148 |
-41.824 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.366 |
-0.358 |
-0.028 |
| y |
-0.358 |
1.938 |
0.148 |
| z |
-0.028 |
0.148 |
2.428 |
|
| Polar |
|---|
| 3z2-r2 | 4.856 |
|---|
| x2-y2 | -4.203 |
|---|
| xy | -0.358 |
|---|
| xz | -0.028 |
|---|
| yz | 0.148 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.002 |
-0.428 |
0.137 |
| y |
-0.428 |
9.299 |
0.148 |
| z |
0.137 |
0.148 |
6.274 |
<r2> (average value of r
2) Å
2
| <r2> |
177.843 |
| (<r2>)1/2 |
13.336 |