return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6OS (Dihydro-2-(3H)-thiophenone)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-629.050132
Energy at 298.15K-629.057770
HF Energy-628.776994
Nuclear repulsion energy290.769772
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3176 3015 10.92      
2 A 3162 3002 8.51      
3 A 3147 2987 17.00      
4 A 3112 2953 18.43      
5 A 3099 2941 21.77      
6 A 3080 2923 6.80      
7 A 1810 1718 317.18      
8 A 1548 1469 0.10      
9 A 1534 1456 5.39      
10 A 1508 1431 6.58      
11 A 1393 1322 0.72      
12 A 1345 1277 6.39      
13 A 1341 1273 7.24      
14 A 1275 1210 2.26      
15 A 1214 1152 2.03      
16 A 1142 1084 6.96      
17 A 1088 1033 59.07      
18 A 1058 1004 5.71      
19 A 1047 994 43.04      
20 A 934 887 9.88      
21 A 892 847 7.07      
22 A 856 813 15.90      
23 A 707 671 2.45      
24 A 627 595 37.07      
25 A 605 575 14.21      
26 A 486 461 1.23      
27 A 478 454 1.86      
28 A 401 381 3.33      
29 A 234 222 1.45      
30 A 128 121 0.60      

Unscaled Zero Point Vibrational Energy (zpe) 21212.9 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 20135.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.14679 0.10394 0.06427

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.117 1.432 0.027
H2 -0.454 2.130 -0.743
H3 -0.253 1.906 1.003
C4 1.349 1.014 -0.227
H5 2.089 1.720 0.159
H6 1.488 0.934 -1.308
C7 1.522 -0.398 0.351
H8 1.581 -0.389 1.440
H9 2.411 -0.889 -0.049
C10 -0.964 0.196 0.000
O11 -2.185 0.192 0.114
S12 -0.007 -1.276 -0.145

Atom - Atom Distances (Å)
  C1 H2 H3 C4 H5 H6 C7 H8 H9 C10 O11 S12
C11.09231.09361.54572.22832.14642.47822.86373.43281.49912.41262.7160
H21.09231.77172.18252.72922.34993.39013.90614.21952.13422.73603.4872
H31.09361.77172.20802.49583.05262.98082.97074.00202.10622.73133.3919
C41.54572.18252.20801.09261.09311.53582.19192.18672.46463.64432.6633
H52.22832.72922.49581.09261.77022.20072.51922.63703.41594.53843.6689
H62.14642.34993.05261.09311.77022.12793.05202.40012.87604.00772.9109
C72.47823.39012.98081.53582.20072.12791.09121.09132.58013.76081.8316
H82.86373.90612.97072.19192.51923.05201.09121.77662.98254.03472.4128
H93.43284.21954.00202.18672.63702.40011.09131.77663.54554.72372.4505
C101.49912.13422.10622.46463.41592.87602.58012.98253.54551.22571.7618
O112.41262.73602.73133.64434.53844.00773.76084.03474.72371.22572.6389
S122.71603.48723.39192.66333.66892.91091.83162.41282.45051.76182.6389

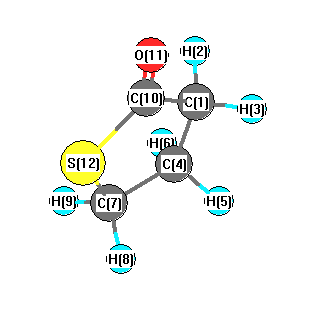
picture of Dihydro-2-(3H)-thiophenone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C4 H5 114.160 C1 C4 H6 107.619
C1 C4 C7 107.069 C1 C10 O11 124.310
C1 C10 S12 112.552 H2 C1 H3 108.286
H2 C1 C4 110.468 H2 C1 C10 109.893
H3 C1 C4 112.436 H3 C1 C10 107.629
C4 C1 C10 108.077 C4 C7 H8 111.991
C4 C7 H9 111.564 C4 C7 S12 104.195
H5 C4 H6 108.169 H5 C4 C7 112.613
H6 C4 C7 106.867 C7 S12 C10 91.760
H8 C7 H9 108.983 H8 C7 S12 108.619
H9 C7 S12 111.408 O11 C10 S12 123.092
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.358      
2 H 0.202      
3 H 0.195      
4 C -0.314      
5 H 0.174      
6 H 0.181      
7 C -0.462      
8 H 0.193      
9 H 0.199      
10 C 0.315      
11 O -0.467      
12 S 0.143      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 10.053 -0.457 0.145
y -0.457 9.269 0.152
z 0.145 0.152 6.231


<r2> (average value of r2) Å2
<r2> 179.457
(<r2>)1/2 13.396