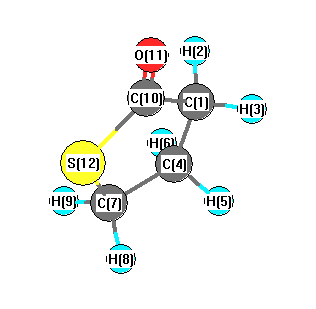
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3075 |
3024 |
10.32 |
|
|
|
| 2 |
A |
3065 |
3014 |
7.70 |
|
|
|
| 3 |
A |
3048 |
2998 |
16.07 |
|
|
|
| 4 |
A |
3008 |
2958 |
18.21 |
|
|
|
| 5 |
A |
3001 |
2951 |
22.78 |
|
|
|
| 6 |
A |
2978 |
2929 |
6.74 |
|
|
|
| 7 |
A |
1784 |
1754 |
319.25 |
|
|
|
| 8 |
A |
1475 |
1451 |
0.32 |
|
|
|
| 9 |
A |
1462 |
1437 |
6.10 |
|
|
|
| 10 |
A |
1435 |
1411 |
6.73 |
|
|
|
| 11 |
A |
1330 |
1308 |
0.56 |
|
|
|
| 12 |
A |
1280 |
1259 |
7.52 |
|
|
|
| 13 |
A |
1273 |
1252 |
5.12 |
|
|
|
| 14 |
A |
1221 |
1201 |
2.36 |
|
|
|
| 15 |
A |
1159 |
1140 |
1.82 |
|
|
|
| 16 |
A |
1091 |
1073 |
5.62 |
|
|
|
| 17 |
A |
1043 |
1026 |
33.97 |
|
|
|
| 18 |
A |
1009 |
993 |
9.21 |
|
|
|
| 19 |
A |
1002 |
986 |
59.36 |
|
|
|
| 20 |
A |
908 |
893 |
9.23 |
|
|
|
| 21 |
A |
852 |
838 |
8.44 |
|
|
|
| 22 |
A |
819 |
805 |
16.60 |
|
|
|
| 23 |
A |
678 |
667 |
2.11 |
|
|
|
| 24 |
A |
604 |
594 |
32.46 |
|
|
|
| 25 |
A |
580 |
571 |
14.71 |
|
|
|
| 26 |
A |
465 |
458 |
1.42 |
|
|
|
| 27 |
A |
460 |
453 |
1.43 |
|
|
|
| 28 |
A |
381 |
375 |
3.65 |
|
|
|
| 29 |
A |
227 |
223 |
1.39 |
|
|
|
| 30 |
A |
127 |
125 |
0.49 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20419.1 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 20082.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.353 |
|
|
|
| 2 |
H |
0.197 |
|
|
|
| 3 |
H |
0.195 |
|
|
|
| 4 |
C |
-0.319 |
|
|
|
| 5 |
H |
0.172 |
|
|
|
| 6 |
H |
0.183 |
|
|
|
| 7 |
C |
-0.468 |
|
|
|
| 8 |
H |
0.198 |
|
|
|
| 9 |
H |
0.201 |
|
|
|
| 10 |
C |
0.235 |
|
|
|
| 11 |
O |
-0.386 |
|
|
|
| 12 |
S |
0.144 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.509 |
1.870 |
0.383 |
3.994 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-45.997 |
-0.332 |
-0.023 |
| y |
-0.332 |
-42.229 |
0.151 |
| z |
-0.023 |
0.151 |
-41.906 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.930 |
-0.332 |
-0.023 |
| y |
-0.332 |
1.723 |
0.151 |
| z |
-0.023 |
0.151 |
2.207 |
|
| Polar |
|---|
| 3z2-r2 | 4.414 |
|---|
| x2-y2 | -3.769 |
|---|
| xy | -0.332 |
|---|
| xz | -0.023 |
|---|
| yz | 0.151 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.427 |
-0.496 |
0.134 |
| y |
-0.496 |
9.794 |
0.165 |
| z |
0.134 |
0.165 |
6.401 |
<r2> (average value of r
2) Å
2
| <r2> |
180.799 |
| (<r2>)1/2 |
13.446 |