return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5CH2CH3 (Ethylbenzene)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-309.611478
Energy at 298.15K-309.622132
HF Energy-308.774632
Nuclear repulsion energy341.741351
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3340 3085 17.99      
2 A' 3317 3064 6.35      
3 A' 3301 3049 9.63      
4 A' 3251 3003 44.90      
5 A' 3178 2936 26.38      
6 A' 3174 2931 32.76      
7 A' 1779 1644 4.76      
8 A' 1634 1509 14.67      
9 A' 1613 1490 2.96      
10 A' 1596 1474 0.42      
11 A' 1518 1402 1.94      
12 A' 1457 1345 4.17      
13 A' 1302 1203 0.15      
14 A' 1277 1180 0.18      
15 A' 1158 1069 7.33      
16 A' 1116 1031 2.16      
17 A' 1066 985 0.22      
18 A' 1037 958 1.71      
19 A' 1008 931 0.05      
20 A' 959 885 1.20      
21 A' 824 761 4.37      
22 A' 798 738 53.88      
23 A' 708 654 12.72      
24 A' 587 542 5.14      
25 A' 523 483 4.60      
26 A' 327 302 0.27      
27 A' 144 133 0.67      
28 A" 3326 3072 38.26      
29 A" 3303 3051 5.82      
30 A" 3258 3009 33.31      
31 A" 3217 2972 9.16      
32 A" 1754 1620 1.13      
33 A" 1600 1478 7.64      
34 A" 1582 1462 8.70      
35 A" 1440 1330 0.35      
36 A" 1381 1275 0.07      
37 A" 1338 1236 0.06      
38 A" 1240 1145 0.03      
39 A" 1184 1094 3.13      
40 A" 1122 1036 1.47      
41 A" 1011 933 0.00      
42 A" 909 840 0.01      
43 A" 833 770 0.60      
44 A" 665 614 0.03      
45 A" 433 400 0.00      
46 A" 376 347 0.09      
47 A" 234 217 0.02      
48 A" 40 37 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 36115.1 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 33359.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.15312 0.04926 0.04089

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.235 0.489 0.000
C2 -0.236 -0.223 1.194
C3 -0.236 -1.609 1.197
C4 -0.235 -2.307 0.000
C5 -0.236 -1.609 -1.197
C6 -0.236 -0.223 -1.194
C7 -0.190 1.996 0.000
C8 1.240 2.536 0.000
H9 -0.240 0.312 2.133
H10 -0.240 -2.144 2.135
H11 -0.238 -3.387 0.000
H12 -0.240 -2.144 -2.135
H13 -0.240 0.312 -2.133
H14 -0.720 2.375 -0.873
H15 -0.720 2.375 0.873
H16 1.782 2.193 0.878
H17 1.782 2.193 -0.878
H18 1.247 3.624 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.39062.41582.79672.41581.39061.50742.52292.14003.38983.87593.38982.14002.13342.13342.78292.78293.4677
C21.39061.38572.40192.76362.38812.52063.34901.08082.13853.38113.84323.36973.35502.66233.16373.76894.2927
C32.41581.38571.38582.39412.76363.79884.55952.13731.07962.14303.37463.84444.51544.02624.31594.77845.5693
C42.79672.40191.38581.38582.40194.30365.06273.37822.14131.07922.14133.37824.78744.78745.00925.00926.1138
C52.41582.76362.39411.38581.38573.79884.55953.84443.37462.14301.07962.13734.02624.51544.77844.31595.5693
C61.39062.38812.76362.40191.38572.52063.34903.36973.84323.38112.13851.08082.66233.35503.76893.16374.2927
C71.50742.52063.79884.30363.79882.52061.52822.71754.65805.38284.65802.71751.08911.08912.16772.16772.1714
C82.52293.34904.55955.06274.55953.34901.52823.41805.35206.10395.35203.41802.15162.15161.08781.08781.0884
H92.14001.08082.13733.37823.84443.36972.71753.41802.45624.26994.92404.26543.67622.46383.03314.08584.2104
H103.38982.13851.07962.14133.37463.84324.65805.35202.45622.47044.27004.92405.44914.71604.94695.65456.3273
H113.87593.38112.14301.07922.14303.38115.38286.10394.26992.47042.47044.26995.84705.84705.99855.99857.1662
H123.38983.84323.37462.14131.07962.13854.65805.35204.92404.27002.47042.45624.71605.44915.65454.94696.3273
H132.14003.36973.84443.37822.13731.08082.71753.41804.26544.92404.26992.45622.46383.67624.08583.03314.2104
H142.13343.35504.51544.78744.02622.66231.08912.15163.67625.44915.84704.71602.46381.74533.05952.50892.4884
H152.13342.66234.02624.78744.51543.35501.08912.15162.46384.71605.84705.44913.67621.74532.50893.05952.4884
H162.78293.16374.31595.00924.77843.76892.16771.08783.03314.94695.99855.65454.08583.05952.50891.75701.7625
H172.78293.76894.77845.00924.31593.16372.16771.08784.08585.65455.99854.94693.03312.50893.05951.75701.7625
H183.46774.29275.56936.11385.56934.29272.17141.08844.21046.32737.16626.32734.21042.48842.48841.76251.7625

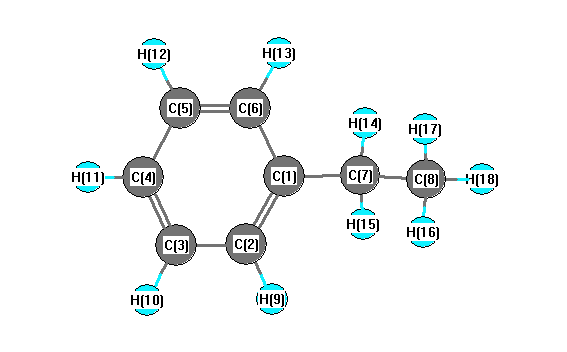
picture of Ethylbenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.952 C1 C2 H9 119.447
C1 C6 C5 120.952 C1 C6 H13 119.447
C1 C7 C8 112.423 C1 C7 H14 109.440
C1 C7 H15 109.440 C2 C1 C6 118.341
C2 C1 C7 120.814 C2 C3 C4 120.137
C2 C3 H10 119.805 C3 C2 H9 119.601
C3 C4 C5 119.482 C3 C4 H11 120.259
C4 C3 H10 120.058 C4 C5 C6 120.137
C4 C5 H12 120.058 C5 C4 H11 120.259
C5 C6 H13 119.601 C6 C1 C7 120.814
C6 C5 H12 119.805 C7 C8 H16 110.792
C7 C8 H17 110.792 C7 C8 H18 111.056
C8 C7 H14 109.435 C8 C7 H15 109.435
H14 C7 H15 106.492 H16 C8 H17 107.723
H16 C8 H18 108.178 H17 C8 H18 108.178
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability